return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (1,3-Dioxane)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-306.753851
Energy at 298.15K-306.765225
Nuclear repulsion energy264.547741
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3175 3024 33.65      
2 A' 3142 2992 29.50      
3 A' 3129 2979 21.26      
4 A' 3074 2928 19.51      
5 A' 3013 2870 98.27      
6 A' 2974 2832 60.04      
7 A' 1569 1494 5.39      
8 A' 1552 1478 0.12      
9 A' 1523 1450 5.16      
10 A' 1447 1378 24.72      
11 A' 1345 1281 4.79      
12 A' 1244 1185 67.26      
13 A' 1218 1160 114.03      
14 A' 1142 1087 14.21      
15 A' 1031 982 38.53      
16 A' 945 900 8.04      
17 A' 869 828 12.19      
18 A' 663 631 5.82      
19 A' 495 472 0.92      
20 A' 450 429 11.79      
21 A' 273 260 2.28      
22 A" 3139 2989 42.23      
23 A" 3005 2862 17.84      
24 A" 1551 1477 2.09      
25 A" 1476 1405 19.05      
26 A" 1426 1358 0.26      
27 A" 1400 1333 0.05      
28 A" 1361 1296 1.74      
29 A" 1289 1227 30.60      
30 A" 1255 1195 1.64      
31 A" 1133 1079 71.10      
32 A" 1066 1015 57.36      
33 A" 942 897 16.64      
34 A" 925 880 0.78      
35 A" 467 445 8.13      
36 A" 274 261 2.37      

Unscaled Zero Point Vibrational Energy (zpe) 27490.8 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 26179.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.16658 0.15992 0.09185

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.629 -1.202 0.000
O2 0.016 -0.765 1.174
O3 0.016 -0.765 -1.174
C4 0.016 0.663 1.240
C5 0.016 0.663 -1.240
C6 0.693 1.242 0.000
H7 -0.587 -2.294 0.000
H8 -1.680 -0.852 0.000
H9 0.543 0.927 2.163
H10 -1.023 1.030 1.315
H11 0.543 0.927 -2.163
H12 -1.023 1.030 -1.315
H13 1.751 0.951 0.000
H14 0.634 2.340 0.000

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.40881.40882.33062.33062.77901.09191.10803.25342.62123.25342.62123.20963.7607
O21.40882.34731.42952.80432.42172.01922.06452.02972.07993.77803.24042.70803.3766
O31.40882.34732.80431.42952.42172.01922.06453.77803.24042.02972.07992.70803.3766
C42.33061.42952.80432.47981.52693.26192.58991.09481.10533.45322.78292.15182.1754
C52.33062.80431.42952.47981.52693.26192.58993.45322.78291.09481.10532.15182.1754
C62.77902.42172.42171.52691.52693.76023.16482.19072.17272.19072.17271.09711.0995
H71.09192.01922.01923.26193.26193.76021.80904.04063.60124.04063.60123.99944.7917
H81.10802.06452.06452.58992.58993.16481.80903.57522.38833.57522.38833.87593.9424
H93.25342.02973.77801.09483.45322.19074.04063.57521.78354.32553.81562.47762.5849
H102.62122.07993.24041.10532.78292.17273.60122.38831.78353.81562.63043.07132.4883
H113.25343.77802.02973.45321.09482.19074.04063.57524.32553.81561.78352.47762.5849
H122.62123.24042.07992.78291.10532.17273.60122.38833.81562.63041.78353.07132.4883
H133.20962.70802.70802.15182.15181.09713.99943.87592.47763.07132.47763.07131.7821
H143.76073.37663.37662.17542.17541.09954.79173.94242.58492.48832.58492.48831.7821

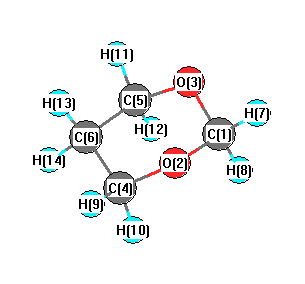
picture of 1,3-Dioxane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C4 110.392 C1 O3 C5 110.392
O2 C1 O3 112.833 O2 C1 H7 107.010
O2 C1 H8 109.652 O2 C4 C6 109.960
O2 C4 H9 106.285 O2 C4 H10 109.617
O3 C1 H7 107.010 O3 C1 H8 109.652
O3 C5 C6 109.960 O3 C5 H11 106.285
O3 C5 H12 109.617 C4 C6 C5 108.595
C4 C6 H13 109.084 C4 C6 H14 110.800
C5 C6 H13 109.084 C5 C6 H14 110.800
C6 C4 H9 112.310 C6 C4 H10 110.245
C6 C5 H11 112.310 C6 C5 H12 110.245
H7 C1 H8 110.628 H9 C4 H10 108.320
H11 C5 H12 108.320 H13 C6 H14 108.445
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability