Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.245692 |
| Energy at 298.15K | -133.249854 |
| HF Energy | -133.245692 |
| Nuclear repulsion energy | 64.821930 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3451 |
3302 |
2.27 |
|
|
|
| 2 |
A' |
3297 |
3155 |
5.17 |
|
|
|
| 3 |
A' |
3190 |
3052 |
3.89 |
|
|
|
| 4 |
A' |
3069 |
2936 |
53.39 |
|
|
|
| 5 |
A' |
1535 |
1468 |
8.01 |
|
|
|
| 6 |
A' |
1477 |
1413 |
2.25 |
|
|
|
| 7 |
A' |
1375 |
1315 |
37.18 |
|
|
|
| 8 |
A' |
1272 |
1217 |
7.58 |
|
|
|
| 9 |
A' |
1127 |
1079 |
22.65 |
|
|
|
| 10 |
A' |
1001 |
957 |
9.78 |
|
|
|
| 11 |
A' |
487 |
466 |
11.62 |
|
|
|
| 12 |
A" |
1051 |
1005 |
0.25 |
|
|
|
| 13 |
A" |
806 |
771 |
10.93 |
|
|
|
| 14 |
A" |
679 |
650 |
115.79 |
|
|
|
| 15 |
A" |
531 |
508 |
5.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12174.1 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 11647.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.418 |
0.000 |
| C2 |
1.152 |
-0.377 |
0.000 |
| N3 |
-1.193 |
-0.136 |
0.000 |
| H4 |
0.143 |
1.507 |
0.000 |
| H5 |
2.145 |
0.062 |
0.000 |
| H6 |
1.055 |
-1.458 |
0.000 |
| H7 |
-1.909 |
0.597 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3997 | 1.3149 | 1.0983 | 2.1742 | 2.1519 | 1.9172 |
C2 | 1.3997 | | 2.3574 | 2.1367 | 1.0850 | 1.0855 | 3.2123 | N3 | 1.3149 | 2.3574 | | 2.1173 | 3.3433 | 2.6075 | 1.0247 | H4 | 1.0983 | 2.1367 | 2.1173 | | 2.4685 | 3.1016 | 2.2447 | H5 | 2.1742 | 1.0850 | 3.3433 | 2.4685 | | 1.8701 | 4.0888 | H6 | 2.1519 | 1.0855 | 2.6075 | 3.1016 | 1.8701 | | 3.6063 | H7 | 1.9172 | 3.2123 | 1.0247 | 2.2447 | 4.0888 | 3.6063 | |
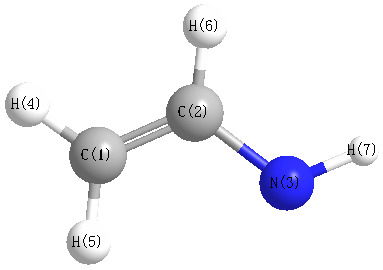 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.578 |
|
C1 |
C2 |
H6 |
119.431 |
| C1 |
N3 |
H7 |
109.433 |
|
C2 |
C1 |
N3 |
120.514 |
| C2 |
C1 |
H4 |
117.084 |
|
N3 |
C1 |
H4 |
122.402 |
| H5 |
C2 |
H6 |
118.992 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.032 |
|
|
|
| 2 |
C |
-0.330 |
|
|
|
| 3 |
N |
-0.531 |
|
|
|
| 4 |
H |
0.149 |
|
|
|
| 5 |
H |
0.178 |
|
|
|
| 6 |
H |
0.195 |
|
|
|
| 7 |
H |
0.306 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.055 |
1.611 |
0.000 |
1.926 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.477 |
-2.856 |
0.000 |
| y |
-2.856 |
-17.369 |
0.000 |
| z |
0.000 |
0.000 |
-19.815 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.115 |
-2.856 |
0.000 |
| y |
-2.856 |
0.778 |
0.000 |
| z |
0.000 |
0.000 |
-2.892 |
|
| Polar |
|---|
| 3z2-r2 | -5.785 |
|---|
| x2-y2 | 0.891 |
|---|
| xy | -2.856 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.336 |
-0.426 |
0.000 |
| y |
-0.426 |
4.028 |
0.000 |
| z |
0.000 |
0.000 |
1.751 |
<r2> (average value of r
2) Å
2
| <r2> |
45.258 |
| (<r2>)1/2 |
6.727 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.245361 |
| Energy at 298.15K | -133.249504 |
| HF Energy | -133.245361 |
| Nuclear repulsion energy | 64.806145 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3399 |
3252 |
12.63 |
|
|
|
| 2 |
A' |
3276 |
3134 |
9.52 |
|
|
|
| 3 |
A' |
3172 |
3035 |
4.75 |
|
|
|
| 4 |
A' |
3139 |
3003 |
25.45 |
|
|
|
| 5 |
A' |
1518 |
1453 |
4.66 |
|
|
|
| 6 |
A' |
1472 |
1408 |
3.83 |
|
|
|
| 7 |
A' |
1399 |
1338 |
13.44 |
|
|
|
| 8 |
A' |
1260 |
1205 |
33.98 |
|
|
|
| 9 |
A' |
1154 |
1104 |
31.85 |
|
|
|
| 10 |
A' |
993 |
950 |
0.72 |
|
|
|
| 11 |
A' |
492 |
471 |
8.20 |
|
|
|
| 12 |
A" |
1084 |
1037 |
49.01 |
|
|
|
| 13 |
A" |
815 |
780 |
72.86 |
|
|
|
| 14 |
A" |
681 |
652 |
6.91 |
|
|
|
| 15 |
A" |
487 |
466 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12170.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 11643.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.446 |
0.000 |
| C2 |
1.130 |
-0.382 |
0.000 |
| N3 |
-1.259 |
0.069 |
0.000 |
| H4 |
0.166 |
1.527 |
0.000 |
| H5 |
2.135 |
0.030 |
0.000 |
| H6 |
1.028 |
-1.465 |
0.000 |
| H7 |
-1.299 |
-0.959 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4014 | 1.3143 | 1.0936 | 2.1752 | 2.1706 | 1.9139 |
C2 | 1.4014 | | 2.4313 | 2.1389 | 1.0856 | 1.0879 | 2.4971 | N3 | 1.3143 | 2.4313 | | 2.0392 | 3.3940 | 2.7541 | 1.0285 | H4 | 1.0936 | 2.1389 | 2.0392 | | 2.4736 | 3.1142 | 2.8860 | H5 | 2.1752 | 1.0856 | 3.3940 | 2.4736 | | 1.8597 | 3.5736 | H6 | 2.1706 | 1.0879 | 2.7541 | 3.1142 | 1.8597 | | 2.3821 | H7 | 1.9139 | 2.4971 | 1.0285 | 2.8860 | 3.5736 | 2.3821 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.482 |
|
C1 |
C2 |
H6 |
120.866 |
| C1 |
N3 |
H7 |
108.945 |
|
C2 |
C1 |
N3 |
127.066 |
| C2 |
C1 |
H4 |
117.490 |
|
N3 |
C1 |
H4 |
115.444 |
| H5 |
C2 |
H6 |
117.652 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.044 |
|
|
|
| 2 |
C |
-0.358 |
|
|
|
| 3 |
N |
-0.521 |
|
|
|
| 4 |
H |
0.173 |
|
|
|
| 5 |
H |
0.185 |
|
|
|
| 6 |
H |
0.179 |
|
|
|
| 7 |
H |
0.297 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.153 |
-1.118 |
0.000 |
2.426 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.466 |
2.138 |
0.000 |
| y |
2.138 |
-15.977 |
0.000 |
| z |
0.000 |
0.000 |
-19.829 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.563 |
2.138 |
0.000 |
| y |
2.138 |
4.170 |
0.000 |
| z |
0.000 |
0.000 |
-1.607 |
|
| Polar |
|---|
| 3z2-r2 | -3.215 |
|---|
| x2-y2 | -4.489 |
|---|
| xy | 2.138 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.014 |
-0.082 |
0.000 |
| y |
-0.082 |
4.215 |
0.000 |
| z |
0.000 |
0.000 |
1.754 |
<r2> (average value of r
2) Å
2
| <r2> |
45.309 |
| (<r2>)1/2 |
6.731 |
