Jump to
S1C2
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -132.863619 |
| Energy at 298.15K | -132.867846 |
| HF Energy | -132.464668 |
| Nuclear repulsion energy | 65.040324 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3492 |
3277 |
1.70 |
|
|
|
| 2 |
A' |
3345 |
3139 |
3.36 |
|
|
|
| 3 |
A' |
3236 |
3037 |
3.95 |
|
|
|
| 4 |
A' |
3138 |
2945 |
38.00 |
|
|
|
| 5 |
A' |
1560 |
1464 |
17.81 |
|
|
|
| 6 |
A' |
1499 |
1407 |
15.44 |
|
|
|
| 7 |
A' |
1404 |
1318 |
16.04 |
|
|
|
| 8 |
A' |
1253 |
1176 |
29.21 |
|
|
|
| 9 |
A' |
1096 |
1029 |
4.86 |
|
|
|
| 10 |
A' |
1032 |
968 |
8.63 |
|
|
|
| 11 |
A' |
505 |
474 |
13.63 |
|
|
|
| 12 |
A" |
1109 |
1041 |
0.45 |
|
|
|
| 13 |
A" |
814 |
764 |
5.02 |
|
|
|
| 14 |
A" |
729 |
684 |
129.37 |
|
|
|
| 15 |
A" |
552 |
518 |
3.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12381.2 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 11621.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.420 |
0.000 |
| C2 |
1.146 |
-0.382 |
0.000 |
| N3 |
-1.185 |
-0.131 |
0.000 |
| H4 |
0.142 |
1.506 |
0.000 |
| H5 |
2.141 |
0.047 |
0.000 |
| H6 |
1.039 |
-1.460 |
0.000 |
| H7 |
-1.905 |
0.597 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3985 | 1.3068 | 1.0954 | 2.1732 | 2.1479 | 1.9136 |
C2 | 1.3985 | | 2.3443 | 2.1383 | 1.0833 | 1.0836 | 3.2045 | N3 | 1.3068 | 2.3443 | | 2.1074 | 3.3306 | 2.5906 | 1.0244 | H4 | 1.0954 | 2.1383 | 2.1074 | | 2.4751 | 3.0988 | 2.2400 | H5 | 2.1732 | 1.0833 | 3.3306 | 2.4751 | | 1.8668 | 4.0836 | H6 | 2.1479 | 1.0836 | 2.5906 | 3.0988 | 1.8668 | | 3.5917 | H7 | 1.9136 | 3.2045 | 1.0244 | 2.2400 | 4.0836 | 3.5917 | |
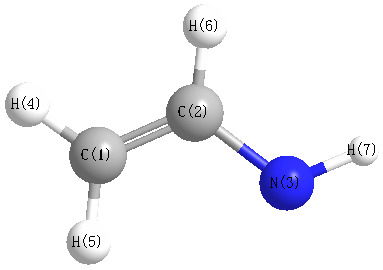 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.724 |
|
C1 |
C2 |
H6 |
119.306 |
| C1 |
N3 |
H7 |
109.753 |
|
C2 |
C1 |
N3 |
120.083 |
| C2 |
C1 |
H4 |
117.533 |
|
N3 |
C1 |
H4 |
122.384 |
| H5 |
C2 |
H6 |
118.970 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3/6-31G*
| | hartrees |
|---|
| Energy at 0K | -132.862899 |
| Energy at 298.15K | -132.867119 |
| HF Energy | -132.464094 |
| Nuclear repulsion energy | 65.019122 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3441 |
3230 |
9.36 |
|
|
|
| 2 |
A' |
3327 |
3122 |
6.36 |
|
|
|
| 3 |
A' |
3222 |
3024 |
2.66 |
|
|
|
| 4 |
A' |
3201 |
3005 |
19.83 |
|
|
|
| 5 |
A' |
1551 |
1456 |
6.44 |
|
|
|
| 6 |
A' |
1489 |
1398 |
2.06 |
|
|
|
| 7 |
A' |
1405 |
1319 |
33.93 |
|
|
|
| 8 |
A' |
1246 |
1169 |
34.60 |
|
|
|
| 9 |
A' |
1118 |
1050 |
27.79 |
|
|
|
| 10 |
A' |
1023 |
960 |
0.62 |
|
|
|
| 11 |
A' |
509 |
478 |
9.32 |
|
|
|
| 12 |
A" |
1135 |
1065 |
55.44 |
|
|
|
| 13 |
A" |
826 |
776 |
80.21 |
|
|
|
| 14 |
A" |
726 |
681 |
1.81 |
|
|
|
| 15 |
A" |
527 |
495 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12373.9 cm
-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 11614.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.450 |
0.000 |
| C2 |
1.123 |
-0.385 |
0.000 |
| N3 |
-1.251 |
0.069 |
0.000 |
| H4 |
0.167 |
1.528 |
0.000 |
| H5 |
2.128 |
0.020 |
0.000 |
| H6 |
1.015 |
-1.465 |
0.000 |
| H7 |
-1.290 |
-0.958 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3994 | 1.3080 | 1.0909 | 2.1714 | 2.1675 | 1.9102 |
C2 | 1.3994 | | 2.4170 | 2.1388 | 1.0836 | 1.0859 | 2.4805 | N3 | 1.3080 | 2.4170 | | 2.0348 | 3.3798 | 2.7361 | 1.0280 | H4 | 1.0909 | 2.1388 | 2.0348 | | 2.4747 | 3.1113 | 2.8821 | H5 | 2.1714 | 1.0836 | 3.3798 | 2.4747 | | 1.8561 | 3.5557 | H6 | 2.1675 | 1.0859 | 2.7361 | 3.1113 | 1.8561 | | 2.3598 | H7 | 1.9102 | 2.4805 | 1.0280 | 2.8821 | 3.5557 | 2.3598 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.458 |
|
C1 |
C2 |
H6 |
120.898 |
| C1 |
N3 |
H7 |
109.133 |
|
C2 |
C1 |
N3 |
126.416 |
| C2 |
C1 |
H4 |
117.841 |
|
N3 |
C1 |
H4 |
115.743 |
| H5 |
C2 |
H6 |
117.643 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
