Jump to
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.121118 |
| Energy at 298.15K | -133.125211 |
| HF Energy | -133.121118 |
| Nuclear repulsion energy | 64.398522 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3331 |
3276 |
5.02 |
|
|
|
| 2 |
A' |
3217 |
3164 |
5.58 |
|
|
|
| 3 |
A' |
3111 |
3060 |
3.32 |
|
|
|
| 4 |
A' |
2972 |
2923 |
58.71 |
|
|
|
| 5 |
A' |
1500 |
1475 |
4.30 |
|
|
|
| 6 |
A' |
1435 |
1412 |
2.13 |
|
|
|
| 7 |
A' |
1334 |
1312 |
38.53 |
|
|
|
| 8 |
A' |
1250 |
1230 |
5.20 |
|
|
|
| 9 |
A' |
1109 |
1091 |
22.57 |
|
|
|
| 10 |
A' |
969 |
953 |
11.00 |
|
|
|
| 11 |
A' |
473 |
465 |
10.49 |
|
|
|
| 12 |
A" |
1013 |
997 |
0.09 |
|
|
|
| 13 |
A" |
781 |
768 |
11.04 |
|
|
|
| 14 |
A" |
642 |
632 |
103.82 |
|
|
|
| 15 |
A" |
518 |
509 |
6.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11828.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 11633.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| C2 |
1.159 |
-0.376 |
0.000 |
| N3 |
-1.200 |
-0.142 |
0.000 |
| H4 |
0.147 |
1.519 |
0.000 |
| H5 |
2.158 |
0.067 |
0.000 |
| H6 |
1.061 |
-1.465 |
0.000 |
| H7 |
-1.917 |
0.604 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4062 | 1.3253 | 1.1078 | 2.1865 | 2.1640 | 1.9262 |
C2 | 1.4062 | | 2.3702 | 2.1478 | 1.0930 | 1.0937 | 3.2284 | N3 | 1.3253 | 2.3702 | | 2.1383 | 3.3642 | 2.6198 | 1.0348 | H4 | 1.1078 | 2.1478 | 2.1383 | | 2.4798 | 3.1208 | 2.2583 | H5 | 2.1865 | 1.0930 | 3.3642 | 2.4798 | | 1.8844 | 4.1104 | H6 | 2.1640 | 1.0937 | 2.6198 | 3.1208 | 1.8844 | | 3.6266 | H7 | 1.9262 | 3.2284 | 1.0348 | 2.2583 | 4.1104 | 3.6266 | |
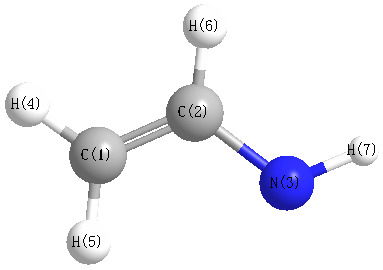 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.567 |
|
C1 |
C2 |
H6 |
119.391 |
| C1 |
N3 |
H7 |
108.778 |
|
C2 |
C1 |
N3 |
120.364 |
| C2 |
C1 |
H4 |
116.880 |
|
N3 |
C1 |
H4 |
122.756 |
| H5 |
C2 |
H6 |
119.042 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.039 |
|
|
|
| 2 |
C |
-0.298 |
|
|
|
| 3 |
N |
-0.500 |
|
|
|
| 4 |
H |
0.130 |
|
|
|
| 5 |
H |
0.163 |
|
|
|
| 6 |
H |
0.178 |
|
|
|
| 7 |
H |
0.288 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.029 |
1.533 |
0.000 |
1.846 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.612 |
-2.761 |
0.000 |
| y |
-2.761 |
-17.535 |
0.000 |
| z |
0.000 |
0.000 |
-19.847 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.080 |
-2.761 |
0.000 |
| y |
-2.761 |
0.694 |
0.000 |
| z |
0.000 |
0.000 |
-2.774 |
|
| Polar |
|---|
| 3z2-r2 | -5.547 |
|---|
| x2-y2 | 0.924 |
|---|
| xy | -2.761 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.519 |
-0.467 |
0.000 |
| y |
-0.467 |
4.183 |
0.000 |
| z |
0.000 |
0.000 |
1.769 |
<r2> (average value of r
2) Å
2
| <r2> |
45.729 |
| (<r2>)1/2 |
6.762 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -133.121093 |
| Energy at 298.15K | -133.125154 |
| HF Energy | -133.121093 |
| Nuclear repulsion energy | 64.359442 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3266 |
3212 |
20.44 |
|
|
|
| 2 |
A' |
3193 |
3140 |
10.59 |
|
|
|
| 3 |
A' |
3092 |
3041 |
5.29 |
|
|
|
| 4 |
A' |
3049 |
2999 |
25.67 |
|
|
|
| 5 |
A' |
1477 |
1452 |
4.48 |
|
|
|
| 6 |
A' |
1435 |
1411 |
4.65 |
|
|
|
| 7 |
A' |
1371 |
1349 |
10.10 |
|
|
|
| 8 |
A' |
1237 |
1217 |
30.25 |
|
|
|
| 9 |
A' |
1131 |
1113 |
32.03 |
|
|
|
| 10 |
A' |
964 |
948 |
0.50 |
|
|
|
| 11 |
A' |
476 |
468 |
7.37 |
|
|
|
| 12 |
A" |
1050 |
1033 |
45.03 |
|
|
|
| 13 |
A" |
788 |
775 |
64.07 |
|
|
|
| 14 |
A" |
650 |
640 |
6.89 |
|
|
|
| 15 |
A" |
455 |
448 |
0.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11817.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 11622.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.446 |
0.000 |
| C2 |
1.139 |
-0.383 |
0.000 |
| N3 |
-1.270 |
0.070 |
0.000 |
| H4 |
0.168 |
1.536 |
0.000 |
| H5 |
2.150 |
0.033 |
0.000 |
| H6 |
1.038 |
-1.474 |
0.000 |
| H7 |
-1.302 |
-0.969 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4084 | 1.3239 | 1.1027 | 2.1896 | 2.1825 | 1.9226 |
C2 | 1.4084 | | 2.4505 | 2.1503 | 1.0939 | 1.0961 | 2.5098 | N3 | 1.3239 | 2.4505 | | 2.0527 | 3.4201 | 2.7763 | 1.0398 | H4 | 1.1027 | 2.1503 | 2.0527 | | 2.4876 | 3.1331 | 2.9040 | H5 | 2.1896 | 1.0939 | 3.4201 | 2.4876 | | 1.8738 | 3.5946 | H6 | 2.1825 | 1.0961 | 2.7763 | 3.1331 | 1.8738 | | 2.3931 | H7 | 1.9226 | 2.5098 | 1.0398 | 2.9040 | 3.5946 | 2.3931 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.596 |
|
C1 |
C2 |
H6 |
120.747 |
| C1 |
N3 |
H7 |
108.254 |
|
C2 |
C1 |
N3 |
127.471 |
| C2 |
C1 |
H4 |
117.294 |
|
N3 |
C1 |
H4 |
115.235 |
| H5 |
C2 |
H6 |
117.658 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.046 |
|
|
|
| 2 |
C |
-0.323 |
|
|
|
| 3 |
N |
-0.487 |
|
|
|
| 4 |
H |
0.155 |
|
|
|
| 5 |
H |
0.168 |
|
|
|
| 6 |
H |
0.164 |
|
|
|
| 7 |
H |
0.278 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.085 |
-1.071 |
0.000 |
2.344 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.495 |
2.046 |
0.000 |
| y |
2.046 |
-16.167 |
0.000 |
| z |
0.000 |
0.000 |
-19.863 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.480 |
2.046 |
0.000 |
| y |
2.046 |
4.012 |
0.000 |
| z |
0.000 |
0.000 |
-1.532 |
|
| Polar |
|---|
| 3z2-r2 | -3.064 |
|---|
| x2-y2 | -4.328 |
|---|
| xy | 2.046 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.197 |
-0.096 |
0.000 |
| y |
-0.096 |
4.371 |
0.000 |
| z |
0.000 |
0.000 |
1.769 |
<r2> (average value of r
2) Å
2
| <r2> |
45.832 |
| (<r2>)1/2 |
6.770 |
