return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Butene, 2-methyl-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-195.856369
Energy at 298.15K-195.866107
HF Energy-195.144252
Nuclear repulsion energy173.790842
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3178 3026 38.82      
2 A 3161 3010 34.87      
3 A 3149 2998 14.27      
4 A 3142 2992 12.12      
5 A 3058 2913 12.21      
6 A 3054 2908 55.43      
7 A 3047 2901 23.55      
8 A 1788 1703 0.61      
9 A 1552 1478 5.71      
10 A 1550 1476 6.34      
11 A 1537 1464 1.34      
12 A 1478 1408 1.52      
13 A 1468 1398 1.64      
14 A 1466 1396 5.74      
15 A 1413 1346 2.26      
16 A 1273 1212 3.02      
17 A 1165 1110 5.71      
18 A 1096 1044 5.15      
19 A 999 951 1.68      
20 A 989 942 0.90      
21 A 787 749 1.35      
22 A 529 503 1.89      
23 A 391 372 0.27      
24 A 306 291 0.19      
25 A 3110 2962 30.93      
26 A 3107 2959 41.64      
27 A 3103 2955 7.45      
28 A 1542 1469 17.23      
29 A 1535 1461 0.57      
30 A 1523 1451 0.84      
31 A 1134 1080 0.00      
32 A 1089 1037 0.85      
33 A 1040 991 1.83      
34 A 835 795 12.36      
35 A 442 421 2.16      
36 A 262 250 1.37      
37 A 177 169 0.84      
38 A 120 114 0.34      
39 A 93 88 0.17      

Unscaled Zero Point Vibrational Energy (zpe) 30342.8 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 28895.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.26693 0.11635 0.08487

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.509 0.503 0.000
H2 -1.976 -0.486 0.000
H3 -1.872 1.049 0.883
H4 -1.872 1.049 -0.883
C5 0.264 -2.103 0.000
H6 0.645 -2.636 -0.883
H7 0.645 -2.636 0.883
H8 -0.826 -2.193 0.000
C9 0.737 -0.674 0.000
H10 1.825 -0.559 0.000
C11 0.000 0.451 0.000
C12 0.665 1.808 0.000
H13 1.758 1.723 0.000
H14 0.366 2.391 0.883
H15 0.366 2.391 -0.883

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 C11 C12 H13 H14 H15
C11.09341.09951.09953.15263.90803.90802.78132.53633.49891.51032.53573.48732.80362.8036
H21.09341.77371.77372.76373.50343.50342.05882.72043.80172.18693.49804.33833.81313.8131
H31.09951.77371.76533.90924.79924.46263.51933.24924.12662.15452.79103.79562.60903.1504
H41.09951.77371.76533.90924.46264.79923.51933.24924.12662.15452.79103.79563.15042.6090
C53.15262.76373.90923.90921.09901.09901.09411.50562.19542.56753.93154.10724.58124.5812
H63.90803.50344.79924.46261.09901.76561.77152.15372.54713.27454.53084.58465.33555.0348
H73.90803.50344.46264.79921.09901.76561.77152.15372.54713.27454.53084.58465.03485.3355
H82.78132.05883.51933.51931.09411.77151.77152.17993.11392.76964.26954.69144.81794.8179
C92.53632.72043.24923.24921.50562.15372.15372.17991.09331.34472.48272.60493.21113.2111
H103.49893.80174.12664.12662.19542.54712.54713.11391.09332.08512.63542.28253.40713.4071
C111.51032.18692.15452.15452.56753.27453.27452.76961.34472.08511.51132.16982.16302.1630
C122.53573.49802.79102.79103.93154.53084.53084.26952.48272.63541.51131.09611.09981.0998
H133.48734.33833.79563.79564.10724.58464.58464.69142.60492.28252.16981.09611.77871.7787
H142.80363.81312.60903.15044.58125.33555.03484.81793.21113.40712.16301.09981.77871.7663
H152.80363.81313.15042.60904.58125.03485.33554.81793.21113.40712.16301.09981.77871.7663

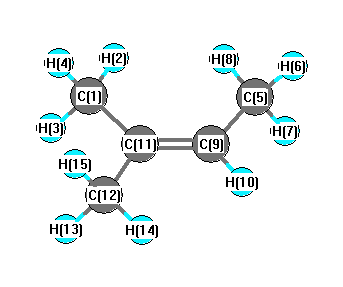
picture of 2-Butene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C11 C9 125.245 C1 C11 C12 114.108
H2 C1 H3 107.962 H2 C1 H4 107.962
H2 C1 C11 113.299 H3 C1 H4 106.784
H3 C1 C11 110.293 H4 C1 C11 110.293
C5 C9 H10 114.360 C5 C9 C11 128.434
H6 C5 H7 106.887 H6 C5 H8 107.758
H6 C5 C9 110.587 H7 C5 H8 107.758
H7 C5 C9 110.587 H8 C5 C9 113.016
C9 C11 C12 120.646 H10 C9 C11 117.206
C11 C12 H13 111.652 C11 C12 H14 110.887
C11 C12 H15 110.887 H13 C12 H14 108.197
H13 C12 H15 108.197 H14 C12 H15 106.841
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability