return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2(SH)CH(CH3)CH3 (1-Propanethiol, 2-methyl-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-555.394035
Energy at 298.15K-555.404960
HF Energy-554.803981
Nuclear repulsion energy234.936346
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3248 3000 26.94      
2 A 3241 2994 33.69      
3 A 3236 2989 44.73      
4 A 3229 2983 25.54      
5 A 3225 2979 6.35      
6 A 3183 2940 29.58      
7 A 3158 2917 21.28      
8 A 3155 2915 4.15      
9 A 3151 2911 20.64      
10 A 2844 2627 22.78      
11 A 1605 1483 13.95      
12 A 1600 1478 3.28      
13 A 1594 1472 3.58      
14 A 1585 1464 1.66      
15 A 1578 1458 3.94      
16 A 1523 1407 8.90      
17 A 1508 1393 6.06      
18 A 1471 1359 4.36      
19 A 1456 1345 2.91      
20 A 1378 1273 25.12      
21 A 1321 1221 3.66      
22 A 1271 1174 5.07      
23 A 1222 1129 3.07      
24 A 1166 1077 2.85      
25 A 1041 962 1.30      
26 A 1029 950 0.38      
27 A 989 913 1.56      
28 A 971 897 3.70      
29 A 934 863 1.52      
30 A 859 794 2.29      
31 A 781 721 1.42      
32 A 451 416 0.07      
33 A 417 385 1.08      
34 A 359 331 0.12      
35 A 267 247 0.05      
36 A 239 221 0.36      
37 A 221 204 1.78      
38 A 174 161 16.37      
39 A 87 80 10.84      

Unscaled Zero Point Vibrational Energy (zpe) 30382.1 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 28064.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.24836 0.07168 0.05988

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.014 1.445 0.165
H2 0.133 2.043 -0.055
H3 1.173 1.468 1.244
H4 1.870 1.923 -0.310
S5 -1.899 0.066 -0.126
H6 -2.691 -0.843 0.448
C7 2.148 -0.785 -0.061
H8 2.073 -1.803 -0.442
H9 3.001 -0.309 -0.542
H10 2.356 -0.839 1.009
C11 -0.313 -0.712 0.329
H12 -0.330 -1.749 0.004
H13 -0.197 -0.695 1.412
C14 0.868 0.006 -0.324
H15 0.693 0.022 -1.402

Atom - Atom Distances (Å)
  C1 H2 H3 H4 S5 H6 C7 H8 H9 H10 C11 H12 H13 C14 H15
C11.08801.09111.08953.23644.36372.51103.46922.74262.78032.53753.46932.75751.52642.1403
H21.08801.76121.76042.83614.06903.47234.32503.74123.79282.81713.82133.12442.18232.4924
H31.09111.76121.76273.64444.57232.77943.78793.11232.60342.79223.76152.56652.16513.0530
H41.08951.76041.76274.20635.38802.73323.73372.51263.09913.48104.29283.75452.16342.4884
S53.23642.83613.64444.20631.33504.13574.40104.93224.49631.82432.40292.41672.77482.8897
H64.36374.06904.57235.38801.33504.86524.94005.80195.07822.38442.56712.67723.73903.9523
C72.51103.47232.77942.73324.13574.86521.08971.08941.09092.49212.65972.77041.52712.1367
H83.46924.32503.78793.73374.40104.94001.08971.76191.76432.73422.44463.13312.17672.4813
H92.74263.74123.11232.51264.93225.80191.08941.76191.76123.45023.67033.76802.16742.4852
H102.78033.79282.60343.09914.49635.07821.09091.76431.76122.75743.00932.58952.16923.0529
C112.53752.81712.79223.48101.82432.38442.49212.73423.45022.75741.08761.08911.52822.1324
H123.46933.82133.76154.29282.40292.56712.65972.44463.67033.00931.08761.76382.15052.4825
H132.75753.12442.56653.75452.41672.67722.77043.13313.76802.58951.08911.76382.15393.0374
C141.52642.18232.16512.16342.77483.73901.52712.17672.16742.16921.52822.15052.15391.0925
H152.14032.49243.05302.48842.88973.95232.13672.48132.48523.05292.13242.48253.03741.0925

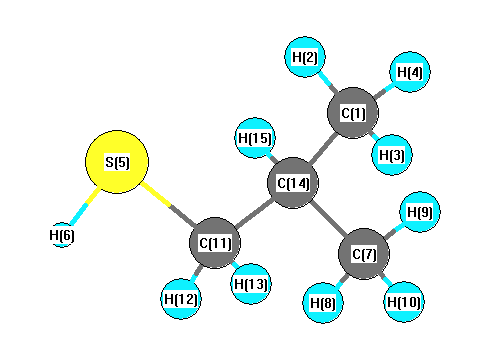
picture of 1-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C14 C7 110.645 C1 C14 C11 112.343
C1 C14 H15 108.493 H2 C1 H3 107.845
H2 C1 H4 107.893 H2 C1 C14 112.089
H3 C1 H4 107.874 H3 C1 C14 110.514
H4 C1 C14 110.475 S5 C11 H12 108.561
S5 C11 H13 109.494 S5 C11 C14 111.413
H6 S5 C11 96.779 C7 C14 C11 109.307
C7 C14 H15 108.165 H8 C7 H9 107.903
H8 C7 H10 108.009 H8 C7 C14 111.478
H9 C7 H10 107.751 H9 C7 C14 110.750
H10 C7 C14 110.804 C11 C14 H15 107.755
H12 C11 H13 108.255 H12 C11 C14 109.437
H13 C11 C14 109.617
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability