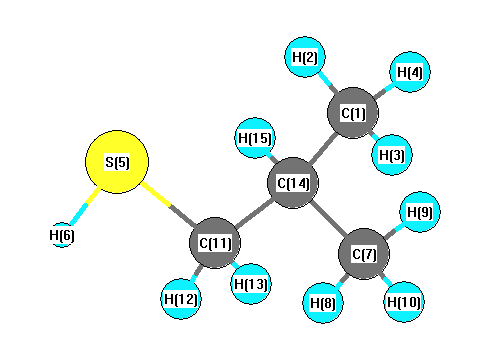
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3155 |
3000 |
27.13 |
|
|
|
| 2 |
A |
3148 |
2994 |
30.59 |
|
|
|
| 3 |
A |
3144 |
2990 |
55.48 |
|
|
|
| 4 |
A |
3139 |
2985 |
9.53 |
|
|
|
| 5 |
A |
3136 |
2982 |
5.34 |
|
|
|
| 6 |
A |
3082 |
2931 |
32.71 |
|
|
|
| 7 |
A |
3066 |
2915 |
19.94 |
|
|
|
| 8 |
A |
3061 |
2911 |
21.67 |
|
|
|
| 9 |
A |
3058 |
2909 |
3.84 |
|
|
|
| 10 |
A |
2736 |
2602 |
22.33 |
|
|
|
| 11 |
A |
1535 |
1460 |
12.08 |
|
|
|
| 12 |
A |
1532 |
1457 |
7.32 |
|
|
|
| 13 |
A |
1522 |
1447 |
3.63 |
|
|
|
| 14 |
A |
1513 |
1439 |
0.72 |
|
|
|
| 15 |
A |
1503 |
1429 |
4.56 |
|
|
|
| 16 |
A |
1447 |
1376 |
9.36 |
|
|
|
| 17 |
A |
1430 |
1360 |
7.92 |
|
|
|
| 18 |
A |
1394 |
1326 |
2.97 |
|
|
|
| 19 |
A |
1379 |
1311 |
2.67 |
|
|
|
| 20 |
A |
1302 |
1238 |
25.63 |
|
|
|
| 21 |
A |
1258 |
1196 |
5.09 |
|
|
|
| 22 |
A |
1215 |
1155 |
5.38 |
|
|
|
| 23 |
A |
1168 |
1111 |
3.28 |
|
|
|
| 24 |
A |
1107 |
1053 |
3.50 |
|
|
|
| 25 |
A |
997 |
948 |
1.99 |
|
|
|
| 26 |
A |
984 |
936 |
0.83 |
|
|
|
| 27 |
A |
945 |
899 |
1.97 |
|
|
|
| 28 |
A |
925 |
879 |
2.90 |
|
|
|
| 29 |
A |
890 |
847 |
2.64 |
|
|
|
| 30 |
A |
821 |
781 |
3.00 |
|
|
|
| 31 |
A |
747 |
710 |
1.76 |
|
|
|
| 32 |
A |
427 |
406 |
0.11 |
|
|
|
| 33 |
A |
395 |
376 |
1.22 |
|
|
|
| 34 |
A |
340 |
324 |
0.15 |
|
|
|
| 35 |
A |
255 |
242 |
0.07 |
|
|
|
| 36 |
A |
228 |
217 |
0.29 |
|
|
|
| 37 |
A |
208 |
197 |
2.15 |
|
|
|
| 38 |
A |
166 |
158 |
14.52 |
|
|
|
| 39 |
A |
84 |
80 |
12.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29218.3 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 27786.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.512 |
|
|
|
| 2 |
H |
0.195 |
|
|
|
| 3 |
H |
0.166 |
|
|
|
| 4 |
H |
0.170 |
|
|
|
| 5 |
S |
-0.094 |
|
|
|
| 6 |
H |
0.106 |
|
|
|
| 7 |
C |
-0.509 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
C |
-0.460 |
|
|
|
| 12 |
H |
0.197 |
|
|
|
| 13 |
H |
0.194 |
|
|
|
| 14 |
C |
-0.143 |
|
|
|
| 15 |
H |
0.178 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.091 |
-1.218 |
0.650 |
1.760 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.601 |
2.684 |
-1.807 |
| y |
2.684 |
-39.919 |
-1.100 |
| z |
-1.807 |
-1.100 |
-41.334 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.026 |
2.684 |
-1.807 |
| y |
2.684 |
1.048 |
-1.100 |
| z |
-1.807 |
-1.100 |
-1.074 |
|
| Polar |
|---|
| 3z2-r2 | -2.148 |
|---|
| x2-y2 | -0.681 |
|---|
| xy | 2.684 |
|---|
| xz | -1.807 |
|---|
| yz | -1.100 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.548 |
-0.026 |
-0.029 |
| y |
-0.026 |
8.246 |
-0.227 |
| z |
-0.029 |
-0.227 |
7.284 |
<r2> (average value of r
2) Å
2
| <r2> |
200.533 |
| (<r2>)1/2 |
14.161 |