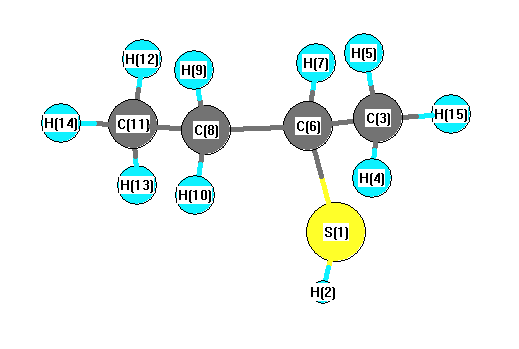
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3169 |
3014 |
17.25 |
|
|
|
| 2 |
A |
3162 |
3007 |
19.48 |
|
|
|
| 3 |
A |
3146 |
2992 |
35.76 |
|
|
|
| 4 |
A |
3135 |
2981 |
28.70 |
|
|
|
| 5 |
A |
3091 |
2940 |
23.17 |
|
|
|
| 6 |
A |
3075 |
2924 |
5.96 |
|
|
|
| 7 |
A |
3070 |
2920 |
22.39 |
|
|
|
| 8 |
A |
3063 |
2913 |
28.64 |
|
|
|
| 9 |
A |
3045 |
2896 |
22.32 |
|
|
|
| 10 |
A |
2711 |
2578 |
17.52 |
|
|
|
| 11 |
A |
1535 |
1460 |
5.82 |
|
|
|
| 12 |
A |
1523 |
1449 |
9.37 |
|
|
|
| 13 |
A |
1523 |
1448 |
4.42 |
|
|
|
| 14 |
A |
1519 |
1444 |
7.99 |
|
|
|
| 15 |
A |
1506 |
1432 |
1.21 |
|
|
|
| 16 |
A |
1438 |
1368 |
11.88 |
|
|
|
| 17 |
A |
1435 |
1365 |
1.60 |
|
|
|
| 18 |
A |
1401 |
1332 |
1.15 |
|
|
|
| 19 |
A |
1346 |
1280 |
12.17 |
|
|
|
| 20 |
A |
1322 |
1258 |
0.70 |
|
|
|
| 21 |
A |
1290 |
1227 |
3.47 |
|
|
|
| 22 |
A |
1200 |
1141 |
1.34 |
|
|
|
| 23 |
A |
1145 |
1089 |
3.87 |
|
|
|
| 24 |
A |
1137 |
1081 |
10.62 |
|
|
|
| 25 |
A |
1062 |
1010 |
0.31 |
|
|
|
| 26 |
A |
1021 |
971 |
8.89 |
|
|
|
| 27 |
A |
985 |
937 |
4.04 |
|
|
|
| 28 |
A |
888 |
845 |
6.92 |
|
|
|
| 29 |
A |
870 |
827 |
5.00 |
|
|
|
| 30 |
A |
799 |
760 |
8.37 |
|
|
|
| 31 |
A |
625 |
594 |
4.70 |
|
|
|
| 32 |
A |
461 |
438 |
0.47 |
|
|
|
| 33 |
A |
382 |
364 |
1.22 |
|
|
|
| 34 |
A |
335 |
319 |
1.59 |
|
|
|
| 35 |
A |
262 |
249 |
0.21 |
|
|
|
| 36 |
A |
237 |
225 |
4.82 |
|
|
|
| 37 |
A |
225 |
214 |
9.59 |
|
|
|
| 38 |
A |
215 |
204 |
7.67 |
|
|
|
| 39 |
A |
114 |
109 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29234.2 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 27801.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.091 |
|
|
|
| 2 |
H |
0.105 |
|
|
|
| 3 |
C |
-0.502 |
|
|
|
| 4 |
H |
0.170 |
|
|
|
| 5 |
H |
0.174 |
|
|
|
| 6 |
C |
-0.300 |
|
|
|
| 7 |
H |
0.200 |
|
|
|
| 8 |
C |
-0.296 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
H |
0.163 |
|
|
|
| 11 |
C |
-0.517 |
|
|
|
| 12 |
H |
0.166 |
|
|
|
| 13 |
H |
0.198 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.193 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.607 |
1.654 |
0.558 |
1.848 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.058 |
0.812 |
1.364 |
| y |
0.812 |
-42.503 |
-1.417 |
| z |
1.364 |
-1.417 |
-38.354 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.630 |
0.812 |
1.364 |
| y |
0.812 |
-2.297 |
-1.417 |
| z |
1.364 |
-1.417 |
3.927 |
|
| Polar |
|---|
| 3z2-r2 | 7.853 |
|---|
| x2-y2 | 0.445 |
|---|
| xy | 0.812 |
|---|
| xz | 1.364 |
|---|
| yz | -1.417 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.104 |
-0.452 |
0.197 |
| y |
-0.452 |
9.319 |
-0.154 |
| z |
0.197 |
-0.154 |
7.823 |
<r2> (average value of r
2) Å
2
| <r2> |
187.452 |
| (<r2>)1/2 |
13.691 |