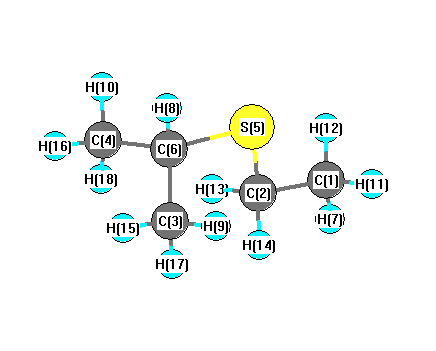
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3078 |
3028 |
23.13 |
|
|
|
| 2 |
A |
3065 |
3015 |
20.42 |
|
|
|
| 3 |
A |
3050 |
3000 |
57.85 |
|
|
|
| 4 |
A |
2991 |
2942 |
28.34 |
|
|
|
| 5 |
A |
2989 |
2940 |
30.59 |
|
|
|
| 6 |
A |
2985 |
2936 |
8.21 |
|
|
|
| 7 |
A |
2981 |
2932 |
21.80 |
|
|
|
| 8 |
A |
1490 |
1465 |
4.14 |
|
|
|
| 9 |
A |
1487 |
1462 |
5.59 |
|
|
|
| 10 |
A |
1481 |
1457 |
13.00 |
|
|
|
| 11 |
A |
1466 |
1442 |
2.47 |
|
|
|
| 12 |
A |
1394 |
1371 |
3.81 |
|
|
|
| 13 |
A |
1387 |
1365 |
2.14 |
|
|
|
| 14 |
A |
1266 |
1246 |
28.86 |
|
|
|
| 15 |
A |
1263 |
1242 |
13.43 |
|
|
|
| 16 |
A |
1159 |
1140 |
8.39 |
|
|
|
| 17 |
A |
1077 |
1059 |
24.58 |
|
|
|
| 18 |
A |
1032 |
1015 |
1.78 |
|
|
|
| 19 |
A |
982 |
966 |
4.90 |
|
|
|
| 20 |
A |
883 |
868 |
1.22 |
|
|
|
| 21 |
A |
676 |
665 |
0.58 |
|
|
|
| 22 |
A |
597 |
587 |
2.18 |
|
|
|
| 23 |
A |
454 |
447 |
2.02 |
|
|
|
| 24 |
A |
359 |
353 |
0.10 |
|
|
|
| 25 |
A |
281 |
276 |
1.05 |
|
|
|
| 26 |
A |
251 |
247 |
0.27 |
|
|
|
| 27 |
A |
155 |
152 |
0.27 |
|
|
|
| 28 |
A |
3077 |
3026 |
6.05 |
|
|
|
| 29 |
A |
3075 |
3024 |
32.48 |
|
|
|
| 30 |
A |
3045 |
2994 |
1.14 |
|
|
|
| 31 |
A |
3035 |
2985 |
8.34 |
|
|
|
| 32 |
A |
2977 |
2928 |
17.04 |
|
|
|
| 33 |
A |
1476 |
1451 |
2.88 |
|
|
|
| 34 |
A |
1472 |
1448 |
6.73 |
|
|
|
| 35 |
A |
1464 |
1439 |
0.65 |
|
|
|
| 36 |
A |
1374 |
1352 |
8.24 |
|
|
|
| 37 |
A |
1303 |
1282 |
1.24 |
|
|
|
| 38 |
A |
1241 |
1220 |
0.27 |
|
|
|
| 39 |
A |
1114 |
1096 |
1.43 |
|
|
|
| 40 |
A |
1024 |
1007 |
0.14 |
|
|
|
| 41 |
A |
936 |
920 |
0.15 |
|
|
|
| 42 |
A |
911 |
896 |
1.63 |
|
|
|
| 43 |
A |
781 |
768 |
5.22 |
|
|
|
| 44 |
A |
312 |
307 |
1.73 |
|
|
|
| 45 |
A |
248 |
244 |
0.02 |
|
|
|
| 46 |
A |
234 |
230 |
0.04 |
|
|
|
| 47 |
A |
70 |
69 |
1.68 |
|
|
|
| 48 |
A |
38 |
38 |
0.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34741.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 34167.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.494 |
|
|
|
| 2 |
C |
-0.435 |
|
|
|
| 3 |
C |
-0.475 |
|
|
|
| 4 |
C |
-0.475 |
|
|
|
| 5 |
S |
0.083 |
|
|
|
| 6 |
C |
-0.268 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.184 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
| 11 |
H |
0.179 |
|
|
|
| 12 |
H |
0.179 |
|
|
|
| 13 |
H |
0.182 |
|
|
|
| 14 |
H |
0.182 |
|
|
|
| 15 |
H |
0.160 |
|
|
|
| 16 |
H |
0.160 |
|
|
|
| 17 |
H |
0.161 |
|
|
|
| 18 |
H |
0.161 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.155 |
-1.706 |
0.000 |
1.713 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.475 |
0.620 |
0.000 |
| y |
0.620 |
-48.847 |
0.000 |
| z |
0.000 |
0.000 |
-47.765 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.831 |
0.620 |
0.000 |
| y |
0.620 |
-3.727 |
0.000 |
| z |
0.000 |
0.000 |
-2.104 |
|
| Polar |
|---|
| 3z2-r2 | -4.208 |
|---|
| x2-y2 | 6.372 |
|---|
| xy | 0.620 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.847 |
1.372 |
0.000 |
| y |
1.372 |
10.353 |
0.000 |
| z |
0.000 |
0.000 |
9.378 |
<r2> (average value of r
2) Å
2
| <r2> |
269.005 |
| (<r2>)1/2 |
16.401 |