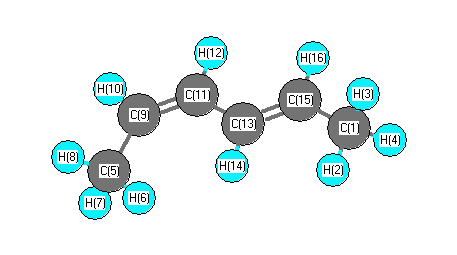
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3178 |
3040 |
74.24 |
|
|
|
| 2 |
A' |
3171 |
3034 |
2.85 |
|
|
|
| 3 |
A' |
3156 |
3020 |
12.19 |
|
|
|
| 4 |
A' |
3153 |
3016 |
20.96 |
|
|
|
| 5 |
A' |
3145 |
3009 |
1.23 |
|
|
|
| 6 |
A' |
3131 |
2995 |
7.30 |
|
|
|
| 7 |
A' |
3042 |
2910 |
32.79 |
|
|
|
| 8 |
A' |
3036 |
2905 |
45.80 |
|
|
|
| 9 |
A' |
1751 |
1675 |
2.74 |
|
|
|
| 10 |
A' |
1707 |
1633 |
5.37 |
|
|
|
| 11 |
A' |
1517 |
1452 |
8.96 |
|
|
|
| 12 |
A' |
1513 |
1447 |
14.55 |
|
|
|
| 13 |
A' |
1459 |
1396 |
5.87 |
|
|
|
| 14 |
A' |
1435 |
1373 |
3.14 |
|
|
|
| 15 |
A' |
1428 |
1366 |
4.36 |
|
|
|
| 16 |
A' |
1347 |
1288 |
1.26 |
|
|
|
| 17 |
A' |
1320 |
1263 |
1.98 |
|
|
|
| 18 |
A' |
1286 |
1231 |
0.98 |
|
|
|
| 19 |
A' |
1193 |
1142 |
0.47 |
|
|
|
| 20 |
A' |
1119 |
1071 |
0.18 |
|
|
|
| 21 |
A' |
1063 |
1017 |
0.37 |
|
|
|
| 22 |
A' |
946 |
905 |
33.11 |
|
|
|
| 23 |
A' |
939 |
899 |
3.47 |
|
|
|
| 24 |
A' |
601 |
575 |
6.08 |
|
|
|
| 25 |
A' |
466 |
445 |
0.21 |
|
|
|
| 26 |
A' |
318 |
304 |
1.92 |
|
|
|
| 27 |
A' |
150 |
143 |
0.56 |
|
|
|
| 28 |
A" |
3092 |
2958 |
23.03 |
|
|
|
| 29 |
A" |
3089 |
2955 |
22.07 |
|
|
|
| 30 |
A" |
1505 |
1440 |
8.05 |
|
|
|
| 31 |
A" |
1502 |
1437 |
5.63 |
|
|
|
| 32 |
A" |
1069 |
1023 |
0.02 |
|
|
|
| 33 |
A" |
1064 |
1018 |
0.92 |
|
|
|
| 34 |
A" |
1030 |
985 |
16.83 |
|
|
|
| 35 |
A" |
981 |
938 |
14.04 |
|
|
|
| 36 |
A" |
847 |
811 |
12.12 |
|
|
|
| 37 |
A" |
729 |
697 |
14.95 |
|
|
|
| 38 |
A" |
443 |
424 |
4.59 |
|
|
|
| 39 |
A" |
216 |
206 |
0.03 |
|
|
|
| 40 |
A" |
201 |
192 |
0.90 |
|
|
|
| 41 |
A" |
133 |
127 |
0.03 |
|
|
|
| 42 |
A" |
109 |
105 |
1.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31288.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29934.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.559 |
|
|
-0.328 |
| 2 |
H |
0.176 |
|
|
0.094 |
| 3 |
H |
0.180 |
|
|
0.110 |
| 4 |
H |
0.180 |
|
|
0.110 |
| 5 |
C |
-0.555 |
|
|
-0.201 |
| 6 |
H |
0.178 |
|
|
0.066 |
| 7 |
H |
0.181 |
|
|
0.076 |
| 8 |
H |
0.181 |
|
|
0.076 |
| 9 |
C |
-0.150 |
|
|
-0.035 |
| 10 |
H |
0.153 |
|
|
0.087 |
| 11 |
C |
-0.145 |
|
|
-0.270 |
| 12 |
H |
0.147 |
|
|
0.135 |
| 13 |
C |
-0.130 |
|
|
-0.145 |
| 14 |
H |
0.145 |
|
|
0.174 |
| 15 |
C |
-0.136 |
|
|
-0.069 |
| 16 |
H |
0.152 |
|
|
0.121 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.222 |
0.132 |
0.000 |
0.258 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.237 |
0.159 |
0.000 |
0.285 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.712 |
-0.608 |
0.000 |
| y |
-0.608 |
-33.654 |
0.000 |
| z |
0.000 |
0.000 |
-40.420 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.325 |
-0.608 |
0.000 |
| y |
-0.608 |
4.412 |
0.000 |
| z |
0.000 |
0.000 |
-5.737 |
|
| Polar |
|---|
| 3z2-r2 | -11.475 |
|---|
| x2-y2 | -2.058 |
|---|
| xy | -0.608 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
260.264 |
| (<r2>)1/2 |
16.133 |