return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (2,4-Hexadiene, (E,Z)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-234.323408
Energy at 298.15K-234.332879
HF Energy-234.067770
Nuclear repulsion energy210.886783
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3196 3034 72.87      
2 A' 3189 3027 1.61      
3 A' 3174 3013 18.10      
4 A' 3170 3009 15.90      
5 A' 3163 3003 0.75      
6 A' 3150 2990 8.53      
7 A' 3069 2913 30.86      
8 A' 3063 2907 41.19      
9 A' 1750 1661 2.18      
10 A' 1702 1615 4.07      
11 A' 1544 1465 7.30      
12 A' 1540 1462 10.74      
13 A' 1478 1403 5.66      
14 A' 1460 1386 4.38      
15 A' 1452 1378 4.10      
16 A' 1362 1293 1.22      
17 A' 1339 1271 2.17      
18 A' 1301 1235 0.73      
19 A' 1204 1143 0.54      
20 A' 1126 1069 0.26      
21 A' 1074 1019 0.51      
22 A' 956 907 21.83      
23 A' 943 895 5.77      
24 A' 605 575 5.39      
25 A' 469 445 0.20      
26 A' 324 308 1.63      
27 A' 153 145 0.52      
28 A" 3118 2960 25.17      
29 A" 3116 2958 23.43      
30 A" 1532 1454 6.26      
31 A" 1529 1451 4.76      
32 A" 1087 1032 0.00      
33 A" 1084 1029 1.22      
34 A" 1030 978 17.55      
35 A" 983 933 10.76      
36 A" 844 801 10.32      
37 A" 733 695 13.62      
38 A" 441 418 3.87      
39 A" 213 202 0.01      
40 A" 201 191 0.86      
41 A" 125 119 0.01      
42 A" 106 101 0.91      

Unscaled Zero Point Vibrational Energy (zpe) 31547.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29945.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.37090 0.04875 0.04378

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.262 2.859 0.000
H2 1.321 2.590 0.000
H3 0.055 3.463 0.881
H4 0.055 3.463 -0.881
C5 1.044 -2.408 0.000
H6 1.725 -1.563 0.000
H7 1.249 -3.003 -0.882
H8 1.249 -3.003 0.882
C9 -0.345 -1.971 0.000
H10 -1.057 -2.716 0.000
C11 -0.786 -0.690 0.000
H12 -1.801 -0.563 0.000
C13 0.000 0.426 0.000
H14 1.013 0.361 0.000
C15 -0.547 1.654 0.000
H16 -1.574 1.749 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 C11 H12 C13 H14 C15 H16
C11.09261.08841.08845.32464.65746.00956.00954.86785.72863.70053.99562.44652.60881.45072.1456
H21.09261.77241.77245.00534.17205.66225.66224.85535.81403.89824.43682.53452.25012.08853.0145
H31.08841.77241.76216.01905.36916.80806.57585.51996.34014.32864.52043.16273.36482.10032.5242
H41.08841.77241.76216.01905.36916.57586.80805.51996.34014.32864.52043.16273.36482.10032.5242
C55.32465.00536.01906.01901.08561.08351.08351.45582.12342.50983.39103.02052.76884.36264.9122
H64.65744.17205.36915.36911.08561.75461.75462.10973.01162.65833.66522.63292.05083.93834.6741
H76.00955.66226.80806.57581.08351.75461.76372.09372.48593.20454.00463.75493.48535.06895.5968
H86.00955.66226.57586.80801.08351.75461.76372.09372.48593.20454.00463.75493.48535.06895.5968
C94.86784.85535.51995.51991.45582.10972.09372.09371.03061.35462.02582.42212.69833.63103.9173
H105.72865.81406.34016.34012.12343.01162.48592.48591.03062.04362.27783.31533.70824.39984.4941
C113.70053.89824.32864.32862.50982.65833.20453.20451.35462.04361.02311.36562.08372.35672.5628
H123.99564.43684.52044.52043.39103.66524.00464.00462.02582.27781.02312.05512.96212.54762.3225
C132.44652.53453.16273.16273.02052.63293.75493.75492.42213.31531.36562.05511.01531.34422.0556
H142.60882.25013.36483.36482.76882.05083.48533.48532.69833.70822.08372.96211.01532.02662.9360
C151.45072.08852.10032.10034.36263.93835.06895.06893.63104.39982.35672.54761.34422.02661.0316
H162.14563.01452.52422.52424.91224.67415.59685.59683.91734.49412.56282.32252.05562.93601.0316

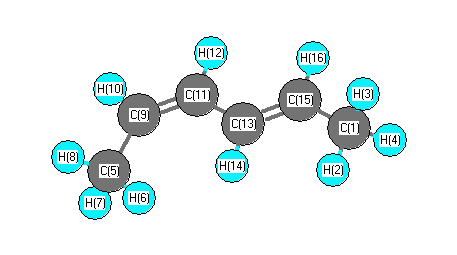
picture of 2,4-Hexadiene, (E,Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C15 C13 122.126 C1 C15 H16 118.638
H2 C1 H3 108.721 H2 C1 H4 108.721
H2 C1 C15 109.607 H3 C1 H4 108.099
H3 C1 C15 110.816 H4 C1 C15 110.816
C5 C9 H10 116.254 C5 C9 C11 126.475
H6 C5 H7 107.979 H6 C5 H8 107.979
H6 C5 C9 111.393 H7 C5 H8 108.965
H7 C5 C9 110.222 H8 C5 C9 110.222
C9 C11 H12 116.154 C9 C11 C13 125.849
H10 C9 C11 117.271 C11 C13 H14 121.430
C11 C13 C15 120.853 H12 C11 C13 117.997
C13 C15 H16 119.235 H14 C13 C15 117.717
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.502      
2 H 0.161      
3 H 0.163      
4 H 0.163      
5 C -0.501      
6 H 0.163      
7 H 0.164      
8 H 0.164      
9 C -0.146      
10 H 0.146      
11 C -0.142      
12 H 0.144      
13 C -0.134      
14 H 0.144      
15 C -0.135      
16 H 0.146      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.620 -1.216 0.000
y -1.216 16.671 0.000
z 0.000 0.000 5.266


<r2> (average value of r2) Å2
<r2> 260.508
(<r2>)1/2 16.140