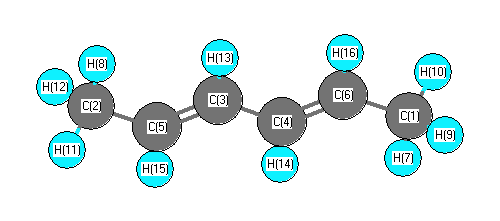
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3155 |
3018 |
0.00 |
|
|
|
| 2 |
Ag |
3146 |
3010 |
0.00 |
|
|
|
| 3 |
Ag |
3131 |
2995 |
0.00 |
|
|
|
| 4 |
Ag |
3036 |
2904 |
0.00 |
|
|
|
| 5 |
Ag |
1755 |
1679 |
0.00 |
|
|
|
| 6 |
Ag |
1517 |
1452 |
0.00 |
|
|
|
| 7 |
Ag |
1437 |
1374 |
0.00 |
|
|
|
| 8 |
Ag |
1370 |
1311 |
0.00 |
|
|
|
| 9 |
Ag |
1335 |
1277 |
0.00 |
|
|
|
| 10 |
Ag |
1212 |
1160 |
0.00 |
|
|
|
| 11 |
Ag |
1133 |
1084 |
0.00 |
|
|
|
| 12 |
Ag |
943 |
902 |
0.00 |
|
|
|
| 13 |
Ag |
435 |
416 |
0.00 |
|
|
|
| 14 |
Ag |
331 |
317 |
0.00 |
|
|
|
| 15 |
Au |
3088 |
2955 |
47.59 |
|
|
|
| 16 |
Au |
1502 |
1437 |
14.23 |
|
|
|
| 17 |
Au |
1067 |
1020 |
0.02 |
|
|
|
| 18 |
Au |
1036 |
991 |
57.97 |
|
|
|
| 19 |
Au |
767 |
734 |
5.69 |
|
|
|
| 20 |
Au |
250 |
239 |
1.43 |
|
|
|
| 21 |
Au |
167 |
160 |
0.07 |
|
|
|
| 22 |
Au |
105 |
100 |
1.73 |
|
|
|
| 23 |
Bg |
3088 |
2954 |
0.00 |
|
|
|
| 24 |
Bg |
1502 |
1437 |
0.00 |
|
|
|
| 25 |
Bg |
1069 |
1023 |
0.00 |
|
|
|
| 26 |
Bg |
975 |
933 |
0.00 |
|
|
|
| 27 |
Bg |
863 |
826 |
0.00 |
|
|
|
| 28 |
Bg |
339 |
324 |
0.00 |
|
|
|
| 29 |
Bg |
195 |
186 |
0.00 |
|
|
|
| 30 |
Bu |
3159 |
3022 |
106.79 |
|
|
|
| 31 |
Bu |
3149 |
3013 |
3.63 |
|
|
|
| 32 |
Bu |
3131 |
2995 |
15.96 |
|
|
|
| 33 |
Bu |
3035 |
2904 |
92.66 |
|
|
|
| 34 |
Bu |
1718 |
1643 |
5.96 |
|
|
|
| 35 |
Bu |
1512 |
1447 |
20.22 |
|
|
|
| 36 |
Bu |
1431 |
1369 |
8.15 |
|
|
|
| 37 |
Bu |
1346 |
1287 |
2.35 |
|
|
|
| 38 |
Bu |
1275 |
1220 |
3.13 |
|
|
|
| 39 |
Bu |
1105 |
1058 |
1.35 |
|
|
|
| 40 |
Bu |
951 |
910 |
33.47 |
|
|
|
| 41 |
Bu |
542 |
519 |
0.04 |
|
|
|
| 42 |
Bu |
137 |
131 |
1.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31218.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29866.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.558 |
|
|
-0.357 |
| 2 |
C |
-0.558 |
|
|
-0.362 |
| 3 |
C |
-0.131 |
|
|
-0.261 |
| 4 |
C |
-0.131 |
|
|
-0.274 |
| 5 |
C |
-0.141 |
|
|
0.035 |
| 6 |
C |
-0.141 |
|
|
0.039 |
| 7 |
H |
0.176 |
|
|
0.109 |
| 8 |
H |
0.176 |
|
|
0.111 |
| 9 |
H |
0.180 |
|
|
0.109 |
| 10 |
H |
0.180 |
|
|
0.109 |
| 11 |
H |
0.180 |
|
|
0.110 |
| 12 |
H |
0.180 |
|
|
0.110 |
| 13 |
H |
0.144 |
|
|
0.169 |
| 14 |
H |
0.144 |
|
|
0.173 |
| 15 |
H |
0.150 |
|
|
0.091 |
| 16 |
H |
0.150 |
|
|
0.090 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.003 |
0.000 |
0.000 |
0.003 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.830 |
-0.989 |
0.000 |
| y |
-0.989 |
-34.291 |
0.000 |
| z |
0.000 |
0.000 |
-40.425 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.528 |
-0.989 |
0.000 |
| y |
-0.989 |
3.337 |
0.000 |
| z |
0.000 |
0.000 |
-5.865 |
|
| Polar |
|---|
| 3z2-r2 | -11.730 |
|---|
| x2-y2 | -0.539 |
|---|
| xy | -0.989 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
290.415 |
| (<r2>)1/2 |
17.042 |