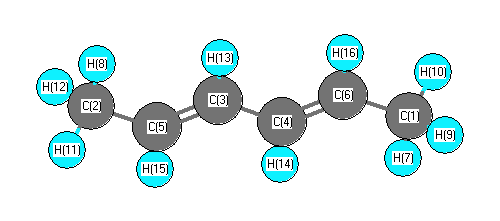
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3073 |
3023 |
0.00 |
|
|
|
| 2 |
Ag |
3060 |
3010 |
0.00 |
|
|
|
| 3 |
Ag |
3054 |
3003 |
0.00 |
|
|
|
| 4 |
Ag |
2960 |
2911 |
0.00 |
|
|
|
| 5 |
Ag |
1690 |
1662 |
0.00 |
|
|
|
| 6 |
Ag |
1475 |
1450 |
0.00 |
|
|
|
| 7 |
Ag |
1391 |
1368 |
0.00 |
|
|
|
| 8 |
Ag |
1329 |
1307 |
0.00 |
|
|
|
| 9 |
Ag |
1295 |
1273 |
0.00 |
|
|
|
| 10 |
Ag |
1185 |
1166 |
0.00 |
|
|
|
| 11 |
Ag |
1109 |
1091 |
0.00 |
|
|
|
| 12 |
Ag |
918 |
903 |
0.00 |
|
|
|
| 13 |
Ag |
426 |
419 |
0.00 |
|
|
|
| 14 |
Ag |
325 |
320 |
0.00 |
|
|
|
| 15 |
Au |
3008 |
2959 |
44.25 |
|
|
|
| 16 |
Au |
1458 |
1434 |
13.58 |
|
|
|
| 17 |
Au |
1028 |
1011 |
0.57 |
|
|
|
| 18 |
Au |
1003 |
987 |
48.78 |
|
|
|
| 19 |
Au |
741 |
729 |
6.51 |
|
|
|
| 20 |
Au |
251 |
247 |
1.45 |
|
|
|
| 21 |
Au |
168 |
166 |
0.01 |
|
|
|
| 22 |
Au |
104 |
102 |
1.85 |
|
|
|
| 23 |
Bg |
3008 |
2959 |
0.00 |
|
|
|
| 24 |
Bg |
1458 |
1434 |
0.00 |
|
|
|
| 25 |
Bg |
1032 |
1014 |
0.00 |
|
|
|
| 26 |
Bg |
938 |
922 |
0.00 |
|
|
|
| 27 |
Bg |
829 |
815 |
0.00 |
|
|
|
| 28 |
Bg |
330 |
324 |
0.00 |
|
|
|
| 29 |
Bg |
196 |
192 |
0.00 |
|
|
|
| 30 |
Bu |
3076 |
3025 |
113.62 |
|
|
|
| 31 |
Bu |
3065 |
3015 |
4.08 |
|
|
|
| 32 |
Bu |
3054 |
3003 |
9.92 |
|
|
|
| 33 |
Bu |
2959 |
2910 |
94.29 |
|
|
|
| 34 |
Bu |
1659 |
1632 |
7.09 |
|
|
|
| 35 |
Bu |
1469 |
1445 |
21.14 |
|
|
|
| 36 |
Bu |
1387 |
1364 |
7.21 |
|
|
|
| 37 |
Bu |
1304 |
1283 |
2.32 |
|
|
|
| 38 |
Bu |
1237 |
1216 |
3.99 |
|
|
|
| 39 |
Bu |
1078 |
1060 |
1.13 |
|
|
|
| 40 |
Bu |
928 |
913 |
38.69 |
|
|
|
| 41 |
Bu |
529 |
520 |
0.05 |
|
|
|
| 42 |
Bu |
136 |
133 |
1.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30360.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29859.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.536 |
|
|
|
| 2 |
C |
-0.536 |
|
|
|
| 3 |
C |
-0.108 |
|
|
|
| 4 |
C |
-0.108 |
|
|
|
| 5 |
C |
-0.116 |
|
|
|
| 6 |
C |
-0.116 |
|
|
|
| 7 |
H |
0.166 |
|
|
|
| 8 |
H |
0.166 |
|
|
|
| 9 |
H |
0.171 |
|
|
|
| 10 |
H |
0.171 |
|
|
|
| 11 |
H |
0.171 |
|
|
|
| 12 |
H |
0.171 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.121 |
|
|
|
| 15 |
H |
0.130 |
|
|
|
| 16 |
H |
0.130 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.863 |
-0.318 |
0.000 |
| y |
-0.318 |
-33.560 |
0.000 |
| z |
0.000 |
0.000 |
-40.482 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.158 |
-0.318 |
0.000 |
| y |
-0.318 |
4.612 |
0.000 |
| z |
0.000 |
0.000 |
-5.770 |
|
| Polar |
|---|
| 3z2-r2 | -11.540 |
|---|
| x2-y2 | -2.302 |
|---|
| xy | -0.318 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.777 |
-0.936 |
0.000 |
| y |
-0.936 |
19.286 |
0.000 |
| z |
0.000 |
0.000 |
5.441 |
<r2> (average value of r
2) Å
2
| <r2> |
293.115 |
| (<r2>)1/2 |
17.121 |