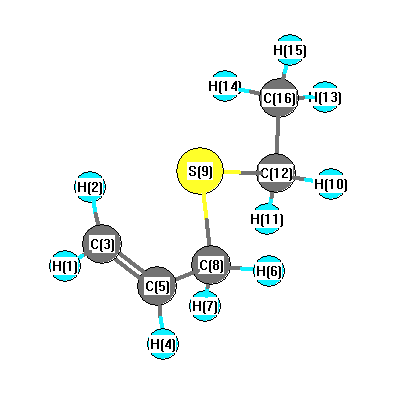
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3259 |
3118 |
19.07 |
|
|
|
| 2 |
A |
3179 |
3041 |
3.27 |
|
|
|
| 3 |
A |
3156 |
3019 |
20.37 |
|
|
|
| 4 |
A |
3151 |
3014 |
25.66 |
|
|
|
| 5 |
A |
3141 |
3005 |
22.51 |
|
|
|
| 6 |
A |
3105 |
2971 |
8.95 |
|
|
|
| 7 |
A |
3067 |
2934 |
16.96 |
|
|
|
| 8 |
A |
3065 |
2933 |
25.11 |
|
|
|
| 9 |
A |
3058 |
2926 |
27.98 |
|
|
|
| 10 |
A |
3031 |
2899 |
27.85 |
|
|
|
| 11 |
A |
1743 |
1667 |
20.95 |
|
|
|
| 12 |
A |
1529 |
1463 |
2.53 |
|
|
|
| 13 |
A |
1519 |
1453 |
8.65 |
|
|
|
| 14 |
A |
1514 |
1448 |
3.28 |
|
|
|
| 15 |
A |
1491 |
1426 |
11.86 |
|
|
|
| 16 |
A |
1466 |
1403 |
5.30 |
|
|
|
| 17 |
A |
1436 |
1374 |
3.89 |
|
|
|
| 18 |
A |
1347 |
1289 |
4.39 |
|
|
|
| 19 |
A |
1334 |
1277 |
7.22 |
|
|
|
| 20 |
A |
1302 |
1246 |
40.93 |
|
|
|
| 21 |
A |
1282 |
1226 |
0.04 |
|
|
|
| 22 |
A |
1198 |
1146 |
1.20 |
|
|
|
| 23 |
A |
1099 |
1052 |
4.55 |
|
|
|
| 24 |
A |
1079 |
1033 |
1.86 |
|
|
|
| 25 |
A |
1064 |
1018 |
0.07 |
|
|
|
| 26 |
A |
1024 |
979 |
13.51 |
|
|
|
| 27 |
A |
1011 |
968 |
4.62 |
|
|
|
| 28 |
A |
950 |
909 |
2.62 |
|
|
|
| 29 |
A |
942 |
902 |
35.49 |
|
|
|
| 30 |
A |
928 |
888 |
7.23 |
|
|
|
| 31 |
A |
801 |
766 |
5.31 |
|
|
|
| 32 |
A |
766 |
733 |
3.63 |
|
|
|
| 33 |
A |
706 |
675 |
1.00 |
|
|
|
| 34 |
A |
549 |
525 |
12.32 |
|
|
|
| 35 |
A |
504 |
482 |
0.21 |
|
|
|
| 36 |
A |
334 |
320 |
0.81 |
|
|
|
| 37 |
A |
248 |
238 |
0.09 |
|
|
|
| 38 |
A |
241 |
231 |
0.17 |
|
|
|
| 39 |
A |
144 |
138 |
0.83 |
|
|
|
| 40 |
A |
124 |
118 |
0.27 |
|
|
|
| 41 |
A |
71 |
68 |
0.19 |
|
|
|
| 42 |
A |
27 |
25 |
1.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30491.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29171.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.166 |
|
|
0.167 |
| 2 |
H |
0.191 |
|
|
0.209 |
| 3 |
C |
-0.379 |
|
|
-0.436 |
| 4 |
H |
0.164 |
|
|
0.093 |
| 5 |
C |
-0.091 |
|
|
0.016 |
| 6 |
H |
0.200 |
|
|
0.129 |
| 7 |
H |
0.200 |
|
|
0.129 |
| 8 |
C |
-0.515 |
|
|
-0.208 |
| 9 |
S |
0.114 |
|
|
-0.276 |
| 10 |
H |
0.190 |
|
|
0.054 |
| 11 |
H |
0.190 |
|
|
0.054 |
| 12 |
C |
-0.461 |
|
|
0.058 |
| 13 |
H |
0.178 |
|
|
0.059 |
| 14 |
H |
0.189 |
|
|
0.093 |
| 15 |
H |
0.189 |
|
|
0.093 |
| 16 |
C |
-0.524 |
|
|
-0.237 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.516 |
1.787 |
0.000 |
1.859 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.533 |
1.756 |
0.000 |
1.835 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.050 |
-0.031 |
-0.000 |
| y |
-0.031 |
-43.454 |
-0.000 |
| z |
-0.000 |
-0.000 |
-47.757 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.556 |
-0.031 |
-0.000 |
| y |
-0.031 |
0.449 |
-0.000 |
| z |
-0.000 |
-0.000 |
-6.005 |
|
| Polar |
|---|
| 3z2-r2 | -12.010 |
|---|
| x2-y2 | 3.404 |
|---|
| xy | -0.031 |
|---|
| xz | -0.000 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
295.973 |
| (<r2>)1/2 |
17.204 |