return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (3-Ethylthio-1-propene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-594.294351
Energy at 298.15K-594.304378
HF Energy-594.038611
Nuclear repulsion energy285.251158
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3285 3118 18.58      
2 A' 3204 3041 3.35      
3 A' 3179 3018 19.71      
4 A' 3162 3002 23.82      
5 A' 3089 2932 23.85      
6 A' 3083 2926 27.02      
7 A' 3059 2904 27.17      
8 A' 1738 1649 17.67      
9 A' 1555 1476 1.67      
10 A' 1540 1462 3.63      
11 A' 1521 1443 10.75      
12 A' 1488 1413 4.88      
13 A' 1459 1385 2.32      
14 A' 1366 1297 4.18      
15 A' 1351 1282 11.34      
16 A' 1324 1257 40.75      
17 A' 1112 1055 4.13      
18 A' 1092 1036 1.39      
19 A' 1018 966 3.62      
20 A' 954 906 2.21      
21 A' 769 730 2.68      
22 A' 709 673 1.13      
23 A' 508 482 0.31      
24 A' 339 322 0.73      
25 A' 247 234 0.18      
26 A' 126 120 0.25      
27 A" 3172 3011 26.82      
28 A" 3129 2970 9.11      
29 A" 3096 2938 17.84      
30 A" 1543 1465 7.17      
31 A" 1302 1235 0.00      
32 A" 1208 1147 1.39      
33 A" 1082 1027 0.09      
34 A" 1031 979 14.09      
35 A" 952 903 4.95      
36 A" 939 891 32.46      
37 A" 813 771 4.03      
38 A" 553 525 10.52      
39 A" 253 240 0.10      
40 A" 133 126 0.66      
41 A" 63 60 0.15      
42 A" 18 17 1.86      

Unscaled Zero Point Vibrational Energy (zpe) 30780.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29217.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.32571 0.04209 0.03807

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -3.779 -1.184 0.000
H2 -2.689 0.237 0.000
C3 -2.812 -0.796 0.000
H4 -1.960 -2.649 0.000
C5 -1.761 -1.625 0.000
H6 0.147 -1.651 0.892
H7 0.147 -1.651 -0.892
C8 -0.331 -1.230 0.000
S9 0.000 0.558 0.000
H10 2.133 -0.109 0.892
H11 2.133 -0.109 -0.892
C12 1.811 0.433 0.000
H13 3.528 1.749 0.000
H14 2.129 2.379 -0.894
H15 2.129 2.379 0.894
C16 2.441 1.832 0.000

Atom - Atom Distances (Å)
  H1 H2 C3 H4 C5 H6 H7 C8 S9 H10 H11 C12 H13 H14 H15 C16
H11.79021.04182.33542.06484.05234.05233.44744.16056.07406.07405.81847.87356.95646.95646.9121
H21.79021.03972.97692.07993.52173.52172.77682.70834.91584.91584.50426.39885.34815.34815.3725
C31.04181.03972.04001.33813.20613.20612.51813.12055.07095.07094.78306.83175.94045.94045.8733
H42.33542.97692.04001.04392.49622.49622.16063.75884.89924.89924.87057.03366.54286.54286.2812
C52.06482.07991.33811.04392.10632.10631.48342.80454.27274.27274.12246.27405.65375.65375.4413
H64.05233.52173.20612.49622.10631.78351.09602.38662.51463.08282.81224.87774.83344.49164.2651
H74.05233.52173.20612.49622.10631.78351.09602.38663.08282.51462.81224.87774.49164.83344.2651
C83.44742.77682.51812.16061.48341.09601.09601.81812.85022.85022.71194.87574.45864.45864.1306
S94.16052.70833.12053.75882.80452.38662.38661.81812.40582.40581.81503.72412.94092.94092.7536
H106.07404.91585.07094.89924.27272.51463.08282.85022.40581.78341.09202.48893.06252.48832.1579
H116.07404.91585.07094.89924.27273.08282.51462.85022.40581.78341.09202.48892.48833.06252.1579
C125.81844.50424.78304.87054.12242.81222.81222.71191.81501.09201.09202.16382.16522.16521.5343
H137.87356.39886.83177.03366.27404.87774.87774.87573.72412.48892.48892.16381.77631.77631.0905
H146.95645.34815.94046.54285.65374.83344.49164.45862.94093.06252.48832.16521.77631.78731.0936
H156.95645.34815.94046.54285.65374.49164.83344.45862.94092.48833.06252.16521.77631.78731.0936
C166.91215.37255.87336.28125.44134.26514.26514.13062.75362.15792.15791.53431.09051.09361.0936

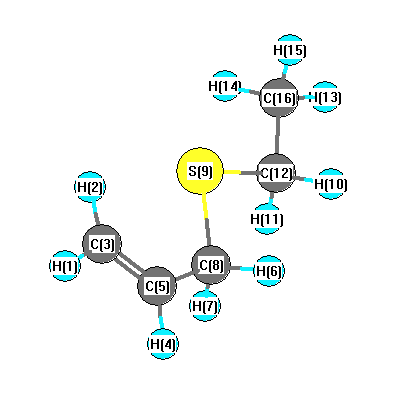
picture of 3-Ethylthio-1-propene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C3 H2 118.640 H1 C3 C5 119.845
H2 C3 C5 121.514 C3 C5 H4 117.296
C3 C5 C8 126.288 H4 C5 C8 116.416
C5 C8 H6 108.557 C5 C8 H7 108.557
C5 C8 S9 115.933 H6 C8 H7 108.906
H6 C8 S9 107.360 H7 C8 S9 107.360
C8 S9 C12 96.565 S9 C12 H10 109.161
S9 C12 H11 109.161 S9 C12 C16 110.318
H10 C12 H11 109.485 H10 C12 C16 109.350
H11 C12 C16 109.350 C12 C16 H13 109.894
C12 C16 H14 109.826 C12 C16 H15 109.826
H13 C16 H14 108.832 H13 C16 H15 108.832
H14 C16 H15 109.608
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.158      
2 H 0.183      
3 C -0.360      
4 H 0.156      
5 C -0.095      
6 H 0.183      
7 H 0.183      
8 C -0.472      
9 S 0.112      
10 H 0.174      
11 H 0.174      
12 C -0.425      
13 H 0.161      
14 H 0.172      
15 H 0.172      
16 C -0.476      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 12.938 1.156 0.000
y 1.156 10.597 0.000
z 0.000 0.000 6.665


<r2> (average value of r2) Å2
<r2> 296.590
(<r2>)1/2 17.222