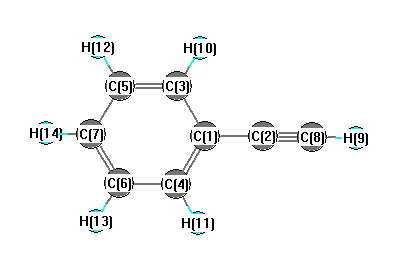
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3514 |
3341 |
79.85 |
|
|
|
| 2 |
A |
3245 |
3086 |
5.42 |
|
|
|
| 3 |
A |
3233 |
3075 |
17.41 |
|
|
|
| 4 |
A |
3213 |
3056 |
0.32 |
|
|
|
| 5 |
A |
2245 |
2135 |
1.98 |
|
|
|
| 6 |
A |
1684 |
1601 |
1.84 |
|
|
|
| 7 |
A |
1547 |
1471 |
16.63 |
|
|
|
| 8 |
A |
1245 |
1184 |
1.03 |
|
|
|
| 9 |
A |
1213 |
1153 |
0.00 |
|
|
|
| 10 |
A |
1066 |
1014 |
3.33 |
|
|
|
| 11 |
A |
1022 |
972 |
0.21 |
|
|
|
| 12 |
A |
783 |
745 |
2.81 |
|
|
|
| 13 |
A |
473 |
449 |
0.34 |
|
|
|
| 14 |
A |
977 |
930 |
0.00 |
|
|
|
| 15 |
A |
862 |
820 |
0.00 |
|
|
|
| 16 |
A |
412 |
392 |
0.00 |
|
|
|
| 17 |
A |
1005 |
956 |
0.10 |
|
|
|
| 18 |
A |
936 |
890 |
4.02 |
|
|
|
| 19 |
A |
778 |
740 |
47.95 |
|
|
|
| 20 |
A |
708 |
674 |
24.08 |
|
|
|
| 21 |
A |
595 |
566 |
34.66 |
|
|
|
| 22 |
A |
559 |
531 |
18.99 |
|
|
|
| 23 |
A |
377 |
359 |
1.57 |
|
|
|
| 24 |
A |
146 |
138 |
1.18 |
|
|
|
| 25 |
A |
3241 |
3082 |
19.55 |
|
|
|
| 26 |
A |
3223 |
3065 |
5.16 |
|
|
|
| 27 |
A |
1653 |
1572 |
1.10 |
|
|
|
| 28 |
A |
1497 |
1424 |
5.73 |
|
|
|
| 29 |
A |
1382 |
1314 |
0.29 |
|
|
|
| 30 |
A |
1344 |
1278 |
0.00 |
|
|
|
| 31 |
A |
1195 |
1137 |
0.01 |
|
|
|
| 32 |
A |
1117 |
1063 |
4.32 |
|
|
|
| 33 |
A |
648 |
616 |
43.76 |
|
|
|
| 34 |
A |
635 |
604 |
2.16 |
|
|
|
| 35 |
A |
537 |
511 |
1.61 |
|
|
|
| 36 |
A |
158 |
151 |
0.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24233.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 23046.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.014 |
|
|
|
| 2 |
C |
0.309 |
|
|
|
| 3 |
C |
-0.156 |
|
|
|
| 4 |
C |
-0.156 |
|
|
|
| 5 |
C |
-0.173 |
|
|
|
| 6 |
C |
-0.173 |
|
|
|
| 7 |
C |
-0.157 |
|
|
|
| 8 |
C |
-0.597 |
|
|
|
| 9 |
H |
0.239 |
|
|
|
| 10 |
H |
0.182 |
|
|
|
| 11 |
H |
0.182 |
|
|
|
| 12 |
H |
0.172 |
|
|
|
| 13 |
H |
0.172 |
|
|
|
| 14 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.633 |
0.633 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.674 |
0.000 |
0.000 |
| y |
0.000 |
-41.136 |
0.000 |
| z |
0.000 |
0.000 |
-36.823 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.695 |
0.000 |
0.000 |
| y |
0.000 |
2.112 |
0.000 |
| z |
0.000 |
0.000 |
8.583 |
|
| Polar |
|---|
| 3z2-r2 | 17.165 |
|---|
| x2-y2 | -8.538 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.994 |
0.000 |
0.000 |
| y |
0.000 |
11.115 |
0.000 |
| z |
0.000 |
0.000 |
19.158 |
<r2> (average value of r
2) Å
2
| <r2> |
263.751 |
| (<r2>)1/2 |
16.240 |