return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2SHCH2SH (1,2-Ethanedithiol)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-874.800002
Energy at 298.15K-874.806248
HF Energy-874.239404
Nuclear repulsion energy202.039466
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3100 2952 0.00      
2 Ag 2715 2586 0.00      
3 Ag 1548 1474 0.00      
4 Ag 1402 1335 0.00      
5 Ag 1137 1083 0.00      
6 Ag 934 890 0.00      
7 Ag 767 730 0.00      
8 Ag 291 277 0.00      
9 Au 3168 3017 13.26      
10 Au 1174 1118 4.65      
11 Au 805 767 2.95      
12 Au 151 144 26.42      
13 Au 89 85 30.24      
14 Bg 3148 2998 0.00      
15 Bg 1340 1276 0.00      
16 Bg 1020 972 0.00      
17 Bg 157 149 0.00      
18 Bu 3106 2958 27.59      
19 Bu 2716 2586 45.95      
20 Bu 1549 1476 5.95      
21 Bu 1301 1239 55.77      
22 Bu 925 881 6.78      
23 Bu 753 717 4.73      
24 Bu 208 198 8.17      

Unscaled Zero Point Vibrational Energy (zpe) 16752.8 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 15953.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.83675 0.05052 0.04852

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.763 0.000
C2 0.000 -0.763 0.000
S3 1.737 -1.345 0.000
S4 -1.737 1.345 0.000
H5 1.451 -2.662 0.000
H6 -1.451 2.662 0.000
H7 -0.516 -1.137 0.890
H8 -0.516 -1.137 -0.890
H9 0.516 1.137 0.890
H10 0.516 1.137 -0.890

Atom - Atom Distances (Å)
  C1 C2 S3 S4 H5 H6 H7 H8 H9 H10
C11.52572.73151.83223.71942.38992.16062.16061.09491.0949
C21.52571.83222.73152.38993.71941.09491.09492.16062.1606
S32.73151.83224.39411.34785.12052.43192.43192.90572.9057
S41.83222.73154.39415.12051.34782.90572.90572.43192.4319
H53.71942.38991.34785.12056.06332.64332.64334.01224.0122
H62.38993.71945.12051.34786.06334.01224.01222.64332.6433
H72.16061.09492.43192.90572.64334.01221.78032.49743.0670
H82.16061.09492.43192.90572.64334.01221.78033.06702.4974
H91.09492.16062.90572.43194.01222.64332.49743.06701.7803
H101.09492.16062.90572.43194.01222.64333.06702.49741.7803

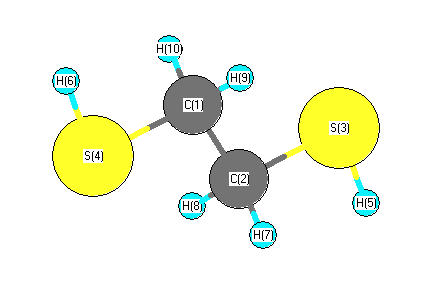
picture of 1,2-Ethanedithiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 S3 108.520 C1 C2 H7 109.982
C1 C2 H8 109.982 C1 S4 H6 96.253
C2 C1 S4 108.520 C2 C1 H9 109.982
C2 C1 H10 109.982 C2 S3 H5 96.253
S3 C2 H7 109.785 S3 C2 H8 109.785
S4 C1 H9 109.785 S4 C1 H10 109.785
H7 C2 H8 108.780 H9 C1 H10 108.780
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability