return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NHNHCH3 (dimethyl hydrazine)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-189.864569
Energy at 298.15K-189.875039
Nuclear repulsion energy136.464232
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3513 3346 0.73      
2 A 3146 2996 2.43      
3 A 3107 2959 12.64      
4 A 3037 2892 87.39      
5 A 1587 1512 1.51      
6 A 1569 1494 0.86      
7 A 1540 1467 0.01      
8 A 1494 1423 0.01      
9 A 1301 1239 19.19      
10 A 1185 1128 0.19      
11 A 1160 1105 0.36      
12 A 963 917 31.79      
13 A 773 736 133.68      
14 A 346 330 8.49      
15 A 331 316 0.71      
16 A 187 178 1.12      
17 B 3530 3362 0.83      
18 B 3146 2996 54.07      
19 B 3106 2958 69.75      
20 B 3022 2878 6.92      
21 B 1563 1489 10.71      
22 B 1535 1462 13.20      
23 B 1494 1423 0.70      
24 B 1478 1408 0.40      
25 B 1202 1144 2.28      
26 B 1165 1110 26.87      
27 B 1047 997 0.10      
28 B 828 789 35.84      
29 B 518 494 2.25      
30 B 235 224 1.04      

Unscaled Zero Point Vibrational Energy (zpe) 24554.7 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 23383.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.48921 0.18665 0.15295

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -0.158 0.698 -0.692
N2 0.158 -0.698 -0.692
C3 0.158 1.435 0.535
C4 -0.158 -1.435 0.535
H5 -1.157 0.760 -0.886
H6 1.157 -0.760 -0.886
H7 -0.107 2.486 0.368
H8 -0.358 1.088 1.444
H9 1.240 1.383 0.711
H10 0.107 -2.486 0.368
H11 0.358 -1.088 1.444
H12 -1.240 -1.383 0.711

Atom - Atom Distances (Å)
  N1 N2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
N11.43171.46562.46091.01931.97302.07902.18082.09553.36642.83202.7329
N21.43172.46091.46561.97301.01933.36642.83202.73292.07902.18082.0955
C31.46562.46092.88762.05032.79901.09631.10211.09743.92482.68933.1505
C42.46091.46562.88762.79902.05033.92482.68933.15051.09631.10211.0974
H51.01931.97302.05032.79902.76822.37792.48522.94663.70213.33772.6736
H61.97301.01932.79902.05032.76823.70213.33772.67362.37792.48522.9466
H72.07903.36641.09633.92482.37793.70211.78211.77434.97633.76104.0456
H82.18082.83201.10212.68932.48523.33771.78211.78303.76102.29042.7237
H92.09552.73291.09743.15052.94662.67361.77431.78304.04562.72373.7144
H103.36642.07903.92481.09633.70212.37794.97633.76104.04561.78211.7743
H112.83202.18082.68931.10213.33772.48523.76102.29042.72371.78211.7830
H122.73292.09553.15051.09742.67362.94664.04562.72373.71441.77431.7830

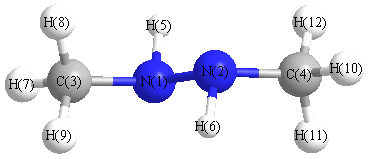
picture of dimethyl hydrazine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 116.281 N1 N2 H6 105.987
N1 C3 H7 107.614 N1 C3 H8 115.550
N1 C3 H9 108.843 N2 N1 C3 116.281
N2 N1 H5 105.987 N2 C4 H10 107.614
N2 C4 H11 115.550 N2 C4 H12 108.843
C3 N1 H5 109.892 C4 N2 H6 109.892
H7 C3 H8 108.313 H7 C3 H9 107.964
H8 C3 H9 108.324 H10 C4 H11 108.313
H10 C4 H12 107.964 H11 C4 H12 108.324
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability