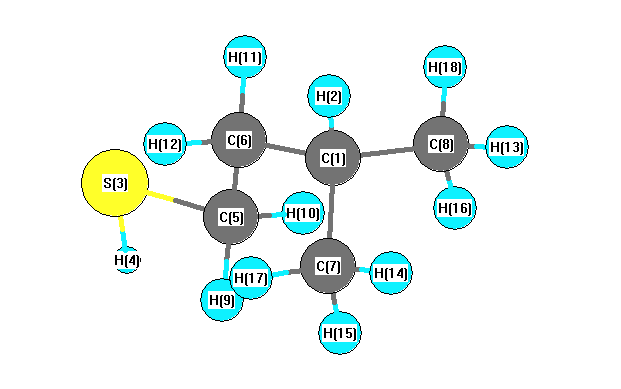
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3129 |
2993 |
55.62 |
|
|
|
| 2 |
A' |
3126 |
2991 |
67.14 |
|
|
|
| 3 |
A' |
3088 |
2955 |
26.04 |
|
|
|
| 4 |
A' |
3054 |
2922 |
1.96 |
|
|
|
| 5 |
A' |
3049 |
2917 |
37.39 |
|
|
|
| 6 |
A' |
3030 |
2899 |
12.87 |
|
|
|
| 7 |
A' |
2700 |
2583 |
26.82 |
|
|
|
| 8 |
A' |
1537 |
1471 |
10.88 |
|
|
|
| 9 |
A' |
1533 |
1467 |
7.10 |
|
|
|
| 10 |
A' |
1518 |
1452 |
0.79 |
|
|
|
| 11 |
A' |
1510 |
1445 |
8.23 |
|
|
|
| 12 |
A' |
1445 |
1383 |
6.94 |
|
|
|
| 13 |
A' |
1405 |
1344 |
1.11 |
|
|
|
| 14 |
A' |
1361 |
1302 |
24.07 |
|
|
|
| 15 |
A' |
1283 |
1227 |
13.23 |
|
|
|
| 16 |
A' |
1219 |
1166 |
2.64 |
|
|
|
| 17 |
A' |
1154 |
1104 |
3.09 |
|
|
|
| 18 |
A' |
1032 |
988 |
0.53 |
|
|
|
| 19 |
A' |
995 |
952 |
2.52 |
|
|
|
| 20 |
A' |
881 |
843 |
0.78 |
|
|
|
| 21 |
A' |
788 |
754 |
1.44 |
|
|
|
| 22 |
A' |
767 |
733 |
6.41 |
|
|
|
| 23 |
A' |
525 |
503 |
1.15 |
|
|
|
| 24 |
A' |
378 |
362 |
0.51 |
|
|
|
| 25 |
A' |
253 |
242 |
0.94 |
|
|
|
| 26 |
A' |
244 |
233 |
0.58 |
|
|
|
| 27 |
A' |
182 |
174 |
1.95 |
|
|
|
| 28 |
A" |
3146 |
3010 |
17.76 |
|
|
|
| 29 |
A" |
3127 |
2991 |
15.34 |
|
|
|
| 30 |
A" |
3120 |
2985 |
9.47 |
|
|
|
| 31 |
A" |
3089 |
2956 |
13.55 |
|
|
|
| 32 |
A" |
3048 |
2916 |
31.45 |
|
|
|
| 33 |
A" |
1524 |
1458 |
0.91 |
|
|
|
| 34 |
A" |
1510 |
1445 |
0.73 |
|
|
|
| 35 |
A" |
1426 |
1364 |
10.06 |
|
|
|
| 36 |
A" |
1386 |
1326 |
1.40 |
|
|
|
| 37 |
A" |
1331 |
1273 |
0.05 |
|
|
|
| 38 |
A" |
1229 |
1176 |
2.71 |
|
|
|
| 39 |
A" |
1115 |
1067 |
0.19 |
|
|
|
| 40 |
A" |
991 |
948 |
0.19 |
|
|
|
| 41 |
A" |
965 |
923 |
0.94 |
|
|
|
| 42 |
A" |
932 |
891 |
0.33 |
|
|
|
| 43 |
A" |
776 |
742 |
3.20 |
|
|
|
| 44 |
A" |
368 |
352 |
0.01 |
|
|
|
| 45 |
A" |
241 |
231 |
0.14 |
|
|
|
| 46 |
A" |
174 |
166 |
17.10 |
|
|
|
| 47 |
A" |
98 |
94 |
1.36 |
|
|
|
| 48 |
A" |
51 |
49 |
8.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35415.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 33882.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.121 |
|
|
0.279 |
| 2 |
H |
0.162 |
|
|
-0.019 |
| 3 |
S |
-0.089 |
|
|
-0.352 |
| 4 |
H |
0.101 |
|
|
0.203 |
| 5 |
C |
-0.469 |
|
|
-0.204 |
| 6 |
C |
-0.321 |
|
|
-0.159 |
| 7 |
C |
-0.509 |
|
|
-0.383 |
| 8 |
C |
-0.509 |
|
|
-0.383 |
| 9 |
H |
0.201 |
|
|
0.127 |
| 10 |
H |
0.201 |
|
|
0.127 |
| 11 |
H |
0.177 |
|
|
0.098 |
| 12 |
H |
0.177 |
|
|
0.098 |
| 13 |
H |
0.169 |
|
|
0.102 |
| 14 |
H |
0.169 |
|
|
0.102 |
| 15 |
H |
0.163 |
|
|
0.092 |
| 16 |
H |
0.163 |
|
|
0.092 |
| 17 |
H |
0.168 |
|
|
0.091 |
| 18 |
H |
0.168 |
|
|
0.091 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.662 |
-0.924 |
0.000 |
1.901 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.681 |
-0.964 |
0.000 |
1.938 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.674 |
3.469 |
0.000 |
| y |
3.469 |
-44.487 |
0.000 |
| z |
0.000 |
0.000 |
-48.553 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.154 |
3.469 |
0.000 |
| y |
3.469 |
5.126 |
0.000 |
| z |
0.000 |
0.000 |
-0.972 |
|
| Polar |
|---|
| 3z2-r2 | -1.944 |
|---|
| x2-y2 | -6.187 |
|---|
| xy | 3.469 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
310.652 |
| (<r2>)1/2 |
17.625 |