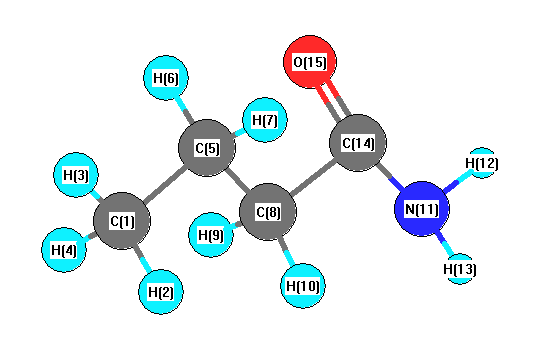
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3745 |
3583 |
32.33 |
|
|
|
| 2 |
A |
3614 |
3457 |
32.89 |
|
|
|
| 3 |
A |
3134 |
2998 |
45.89 |
|
|
|
| 4 |
A |
3131 |
2995 |
39.58 |
|
|
|
| 5 |
A |
3109 |
2974 |
4.97 |
|
|
|
| 6 |
A |
3080 |
2946 |
19.42 |
|
|
|
| 7 |
A |
3066 |
2934 |
25.33 |
|
|
|
| 8 |
A |
3049 |
2917 |
29.04 |
|
|
|
| 9 |
A |
3036 |
2905 |
13.17 |
|
|
|
| 10 |
A |
1822 |
1743 |
275.34 |
|
|
|
| 11 |
A |
1645 |
1574 |
118.63 |
|
|
|
| 12 |
A |
1533 |
1466 |
6.24 |
|
|
|
| 13 |
A |
1525 |
1459 |
7.68 |
|
|
|
| 14 |
A |
1517 |
1451 |
2.73 |
|
|
|
| 15 |
A |
1491 |
1427 |
7.21 |
|
|
|
| 16 |
A |
1438 |
1375 |
11.99 |
|
|
|
| 17 |
A |
1433 |
1371 |
65.75 |
|
|
|
| 18 |
A |
1377 |
1317 |
34.38 |
|
|
|
| 19 |
A |
1332 |
1275 |
29.15 |
|
|
|
| 20 |
A |
1281 |
1226 |
40.07 |
|
|
|
| 21 |
A |
1262 |
1207 |
5.61 |
|
|
|
| 22 |
A |
1147 |
1098 |
0.75 |
|
|
|
| 23 |
A |
1128 |
1079 |
3.27 |
|
|
|
| 24 |
A |
1089 |
1042 |
1.00 |
|
|
|
| 25 |
A |
1067 |
1021 |
2.02 |
|
|
|
| 26 |
A |
940 |
899 |
2.63 |
|
|
|
| 27 |
A |
894 |
855 |
1.11 |
|
|
|
| 28 |
A |
856 |
819 |
3.75 |
|
|
|
| 29 |
A |
755 |
722 |
7.79 |
|
|
|
| 30 |
A |
675 |
646 |
7.48 |
|
|
|
| 31 |
A |
617 |
590 |
15.56 |
|
|
|
| 32 |
A |
520 |
497 |
6.38 |
|
|
|
| 33 |
A |
425 |
407 |
2.02 |
|
|
|
| 34 |
A |
342 |
327 |
2.67 |
|
|
|
| 35 |
A |
248 |
237 |
0.34 |
|
|
|
| 36 |
A |
185 |
177 |
8.15 |
|
|
|
| 37 |
A |
173 |
166 |
238.98 |
|
|
|
| 38 |
A |
90 |
86 |
1.46 |
|
|
|
| 39 |
A |
39 |
38 |
4.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28904.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 27652.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.514 |
|
|
-0.367 |
| 2 |
H |
0.164 |
|
|
0.093 |
| 3 |
H |
0.172 |
|
|
0.087 |
| 4 |
H |
0.161 |
|
|
0.082 |
| 5 |
C |
-0.304 |
|
|
0.244 |
| 6 |
H |
0.169 |
|
|
-0.016 |
| 7 |
H |
0.196 |
|
|
-0.009 |
| 8 |
C |
-0.419 |
|
|
-0.349 |
| 9 |
H |
0.161 |
|
|
0.093 |
| 10 |
H |
0.192 |
|
|
0.097 |
| 11 |
N |
-0.792 |
|
|
-0.977 |
| 12 |
H |
0.359 |
|
|
0.426 |
| 13 |
H |
0.352 |
|
|
0.404 |
| 14 |
C |
0.612 |
|
|
0.734 |
| 15 |
O |
-0.510 |
|
|
-0.543 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.340 |
-3.539 |
0.026 |
3.555 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.324 |
-3.562 |
0.045 |
3.577 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.093 |
-6.237 |
-0.224 |
| y |
-6.237 |
-38.170 |
-0.292 |
| z |
-0.224 |
-0.292 |
-37.403 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.694 |
-6.237 |
-0.224 |
| y |
-6.237 |
-3.422 |
-0.292 |
| z |
-0.224 |
-0.292 |
-2.272 |
|
| Polar |
|---|
| 3z2-r2 | -4.544 |
|---|
| x2-y2 | 6.077 |
|---|
| xy | -6.237 |
|---|
| xz | -0.224 |
|---|
| yz | -0.292 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
216.946 |
| (<r2>)1/2 |
14.729 |