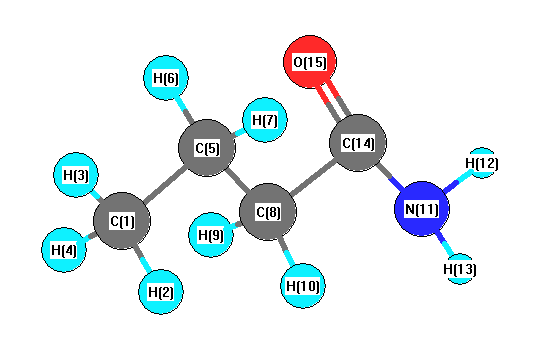
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3767 |
3580 |
35.75 |
|
|
|
| 2 |
A |
3635 |
3454 |
36.82 |
|
|
|
| 3 |
A |
3151 |
2994 |
36.26 |
|
|
|
| 4 |
A |
3149 |
2992 |
41.02 |
|
|
|
| 5 |
A |
3125 |
2970 |
4.10 |
|
|
|
| 6 |
A |
3095 |
2941 |
17.09 |
|
|
|
| 7 |
A |
3083 |
2930 |
22.63 |
|
|
|
| 8 |
A |
3063 |
2911 |
27.06 |
|
|
|
| 9 |
A |
3051 |
2899 |
12.37 |
|
|
|
| 10 |
A |
1842 |
1751 |
280.97 |
|
|
|
| 11 |
A |
1651 |
1569 |
127.71 |
|
|
|
| 12 |
A |
1534 |
1458 |
6.57 |
|
|
|
| 13 |
A |
1527 |
1451 |
8.17 |
|
|
|
| 14 |
A |
1518 |
1442 |
3.28 |
|
|
|
| 15 |
A |
1493 |
1419 |
8.28 |
|
|
|
| 16 |
A |
1446 |
1374 |
77.49 |
|
|
|
| 17 |
A |
1438 |
1367 |
7.85 |
|
|
|
| 18 |
A |
1382 |
1314 |
34.93 |
|
|
|
| 19 |
A |
1337 |
1270 |
24.71 |
|
|
|
| 20 |
A |
1283 |
1219 |
34.17 |
|
|
|
| 21 |
A |
1265 |
1202 |
3.65 |
|
|
|
| 22 |
A |
1155 |
1098 |
0.77 |
|
|
|
| 23 |
A |
1131 |
1075 |
2.96 |
|
|
|
| 24 |
A |
1099 |
1044 |
0.92 |
|
|
|
| 25 |
A |
1077 |
1023 |
1.85 |
|
|
|
| 26 |
A |
948 |
901 |
2.73 |
|
|
|
| 27 |
A |
896 |
852 |
1.20 |
|
|
|
| 28 |
A |
862 |
820 |
3.52 |
|
|
|
| 29 |
A |
756 |
719 |
8.41 |
|
|
|
| 30 |
A |
675 |
641 |
7.48 |
|
|
|
| 31 |
A |
621 |
590 |
16.33 |
|
|
|
| 32 |
A |
521 |
495 |
6.20 |
|
|
|
| 33 |
A |
428 |
406 |
1.94 |
|
|
|
| 34 |
A |
345 |
328 |
2.92 |
|
|
|
| 35 |
A |
248 |
236 |
0.42 |
|
|
|
| 36 |
A |
186 |
177 |
8.33 |
|
|
|
| 37 |
A |
182 |
173 |
241.69 |
|
|
|
| 38 |
A |
90 |
85 |
1.11 |
|
|
|
| 39 |
A |
36 |
34 |
4.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29045.1 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 27601.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.525 |
|
|
|
| 2 |
H |
0.169 |
|
|
|
| 3 |
H |
0.177 |
|
|
|
| 4 |
H |
0.166 |
|
|
|
| 5 |
C |
-0.316 |
|
|
|
| 6 |
H |
0.175 |
|
|
|
| 7 |
H |
0.200 |
|
|
|
| 8 |
C |
-0.433 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
| 10 |
H |
0.197 |
|
|
|
| 11 |
N |
-0.800 |
|
|
|
| 12 |
H |
0.362 |
|
|
|
| 13 |
H |
0.355 |
|
|
|
| 14 |
C |
0.613 |
|
|
|
| 15 |
O |
-0.508 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.356 |
-3.539 |
0.064 |
3.557 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.159 |
-6.186 |
-0.088 |
| y |
-6.186 |
-38.245 |
-0.329 |
| z |
-0.088 |
-0.329 |
-37.491 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.709 |
-6.186 |
-0.088 |
| y |
-6.186 |
-3.420 |
-0.329 |
| z |
-0.088 |
-0.329 |
-2.289 |
|
| Polar |
|---|
| 3z2-r2 | -4.579 |
|---|
| x2-y2 | 6.086 |
|---|
| xy | -6.186 |
|---|
| xz | -0.088 |
|---|
| yz | -0.329 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.977 |
-0.350 |
-0.172 |
| y |
-0.350 |
7.840 |
0.091 |
| z |
-0.172 |
0.091 |
5.719 |
<r2> (average value of r
2) Å
2
| <r2> |
216.046 |
| (<r2>)1/2 |
14.699 |