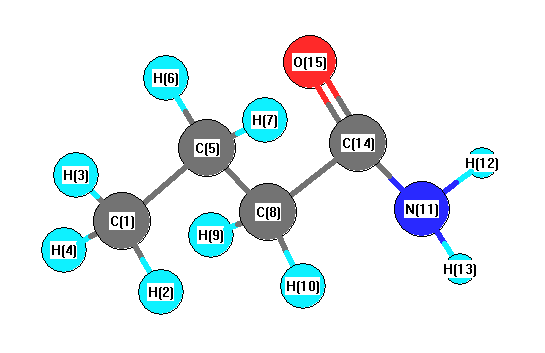
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3628 |
3568 |
20.11 |
|
|
|
| 2 |
A |
3498 |
3440 |
17.27 |
|
|
|
| 3 |
A |
3058 |
3008 |
34.40 |
|
|
|
| 4 |
A |
3055 |
3004 |
50.89 |
|
|
|
| 5 |
A |
3032 |
2982 |
0.91 |
|
|
|
| 6 |
A |
3000 |
2950 |
19.53 |
|
|
|
| 7 |
A |
2988 |
2939 |
28.22 |
|
|
|
| 8 |
A |
2976 |
2927 |
25.95 |
|
|
|
| 9 |
A |
2957 |
2908 |
13.23 |
|
|
|
| 10 |
A |
1756 |
1727 |
233.60 |
|
|
|
| 11 |
A |
1591 |
1564 |
99.56 |
|
|
|
| 12 |
A |
1490 |
1465 |
5.52 |
|
|
|
| 13 |
A |
1482 |
1458 |
7.37 |
|
|
|
| 14 |
A |
1473 |
1449 |
2.69 |
|
|
|
| 15 |
A |
1445 |
1421 |
5.77 |
|
|
|
| 16 |
A |
1391 |
1368 |
1.39 |
|
|
|
| 17 |
A |
1377 |
1354 |
68.74 |
|
|
|
| 18 |
A |
1333 |
1311 |
26.32 |
|
|
|
| 19 |
A |
1291 |
1270 |
33.84 |
|
|
|
| 20 |
A |
1240 |
1220 |
32.28 |
|
|
|
| 21 |
A |
1223 |
1202 |
6.70 |
|
|
|
| 22 |
A |
1113 |
1095 |
1.32 |
|
|
|
| 23 |
A |
1090 |
1072 |
2.90 |
|
|
|
| 24 |
A |
1057 |
1040 |
0.58 |
|
|
|
| 25 |
A |
1037 |
1020 |
2.25 |
|
|
|
| 26 |
A |
914 |
899 |
2.55 |
|
|
|
| 27 |
A |
866 |
852 |
0.87 |
|
|
|
| 28 |
A |
828 |
814 |
4.99 |
|
|
|
| 29 |
A |
735 |
723 |
6.79 |
|
|
|
| 30 |
A |
657 |
647 |
6.35 |
|
|
|
| 31 |
A |
601 |
591 |
16.30 |
|
|
|
| 32 |
A |
503 |
495 |
6.87 |
|
|
|
| 33 |
A |
412 |
405 |
1.72 |
|
|
|
| 34 |
A |
333 |
328 |
2.34 |
|
|
|
| 35 |
A |
245 |
241 |
1.33 |
|
|
|
| 36 |
A |
206 |
203 |
215.24 |
|
|
|
| 37 |
A |
182 |
179 |
8.49 |
|
|
|
| 38 |
A |
90 |
89 |
1.32 |
|
|
|
| 39 |
A |
42 |
42 |
5.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28097.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 27633.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.488 |
|
|
|
| 2 |
H |
0.155 |
|
|
|
| 3 |
H |
0.161 |
|
|
|
| 4 |
H |
0.152 |
|
|
|
| 5 |
C |
-0.276 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
H |
0.181 |
|
|
|
| 8 |
C |
-0.389 |
|
|
|
| 9 |
H |
0.146 |
|
|
|
| 10 |
H |
0.177 |
|
|
|
| 11 |
N |
-0.725 |
|
|
|
| 12 |
H |
0.336 |
|
|
|
| 13 |
H |
0.329 |
|
|
|
| 14 |
C |
0.555 |
|
|
|
| 15 |
O |
-0.471 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.414 |
-3.392 |
0.132 |
3.420 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.162 |
-6.076 |
0.047 |
| y |
-6.076 |
-38.217 |
-0.453 |
| z |
0.047 |
-0.453 |
-37.653 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.773 |
-6.076 |
0.047 |
| y |
-6.076 |
-3.309 |
-0.453 |
| z |
0.047 |
-0.453 |
-2.464 |
|
| Polar |
|---|
| 3z2-r2 | -4.927 |
|---|
| x2-y2 | 6.055 |
|---|
| xy | -6.076 |
|---|
| xz | 0.047 |
|---|
| yz | -0.453 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.652 |
-0.414 |
-0.226 |
| y |
-0.414 |
8.145 |
0.102 |
| z |
-0.226 |
0.102 |
5.877 |
<r2> (average value of r
2) Å
2
| <r2> |
218.945 |
| (<r2>)1/2 |
14.797 |