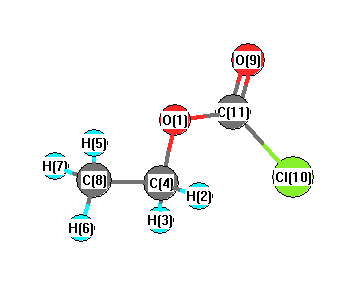
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3034 |
24.65 |
|
|
|
| 2 |
A |
3159 |
3022 |
13.91 |
|
|
|
| 3 |
A |
3145 |
3009 |
3.12 |
|
|
|
| 4 |
A |
3093 |
2959 |
11.62 |
|
|
|
| 5 |
A |
3078 |
2944 |
10.92 |
|
|
|
| 6 |
A |
1932 |
1848 |
493.93 |
|
|
|
| 7 |
A |
1537 |
1471 |
3.78 |
|
|
|
| 8 |
A |
1521 |
1455 |
5.91 |
|
|
|
| 9 |
A |
1509 |
1444 |
7.77 |
|
|
|
| 10 |
A |
1446 |
1384 |
15.15 |
|
|
|
| 11 |
A |
1409 |
1348 |
8.23 |
|
|
|
| 12 |
A |
1317 |
1260 |
0.71 |
|
|
|
| 13 |
A |
1227 |
1174 |
511.67 |
|
|
|
| 14 |
A |
1183 |
1132 |
4.43 |
|
|
|
| 15 |
A |
1150 |
1100 |
23.31 |
|
|
|
| 16 |
A |
1055 |
1010 |
69.25 |
|
|
|
| 17 |
A |
936 |
896 |
92.07 |
|
|
|
| 18 |
A |
829 |
793 |
1.29 |
|
|
|
| 19 |
A |
671 |
642 |
19.31 |
|
|
|
| 20 |
A |
653 |
624 |
90.85 |
|
|
|
| 21 |
A |
530 |
507 |
3.81 |
|
|
|
| 22 |
A |
450 |
430 |
11.48 |
|
|
|
| 23 |
A |
329 |
315 |
5.69 |
|
|
|
| 24 |
A |
261 |
250 |
0.49 |
|
|
|
| 25 |
A |
192 |
184 |
0.13 |
|
|
|
| 26 |
A |
113 |
108 |
0.68 |
|
|
|
| 27 |
A |
80 |
76 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17987.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 17208.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.399 |
|
|
-0.316 |
| 2 |
H |
0.202 |
|
|
0.007 |
| 3 |
H |
0.202 |
|
|
0.007 |
| 4 |
C |
-0.107 |
|
|
0.355 |
| 5 |
H |
0.198 |
|
|
0.134 |
| 6 |
H |
0.186 |
|
|
0.121 |
| 7 |
H |
0.198 |
|
|
0.134 |
| 8 |
C |
-0.531 |
|
|
-0.443 |
| 9 |
O |
-0.373 |
|
|
-0.391 |
| 10 |
Cl |
-0.054 |
|
|
-0.179 |
| 11 |
C |
0.477 |
|
|
0.571 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.434 |
-2.124 |
-0.000 |
4.038 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-3.454 |
-2.055 |
-0.001 |
4.019 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.176 |
-0.824 |
-0.000 |
| y |
-0.824 |
-44.987 |
-0.001 |
| z |
-0.000 |
-0.001 |
-40.397 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.516 |
-0.824 |
-0.000 |
| y |
-0.824 |
-4.200 |
-0.001 |
| z |
-0.000 |
-0.001 |
2.684 |
|
| Polar |
|---|
| 3z2-r2 | 5.368 |
|---|
| x2-y2 | 3.811 |
|---|
| xy | -0.824 |
|---|
| xz | -0.000 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
232.323 |
| (<r2>)1/2 |
15.242 |