return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (1,3-Cyclopentadiene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-193.468139
Energy at 298.15K-193.474354
HF Energy-192.789917
Nuclear repulsion energy156.138208
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3256 3101 3.80      
2 A1 3231 3077 11.73      
3 A1 3057 2911 12.76      
4 A1 1616 1538 1.30      
5 A1 1489 1418 6.88      
6 A1 1439 1370 8.42      
7 A1 1158 1103 0.09      
8 A1 1033 983 0.04      
9 A1 951 906 7.78      
10 A1 819 779 0.01      
11 A2 1161 1105 0.00      
12 A2 906 862 0.00      
13 A2 718 684 0.00      
14 A2 507 483 0.00      
15 B1 3091 2944 13.43      
16 B1 963 917 18.40      
17 B1 930 886 1.13      
18 B1 684 652 70.49      
19 B1 347 330 6.60      
20 B2 3248 3093 27.72      
21 B2 3222 3069 2.00      
22 B2 1682 1602 0.36      
23 B2 1355 1290 1.81      
24 B2 1317 1254 1.00      
25 B2 1143 1088 1.11      
26 B2 1000 953 8.39      
27 B2 821 782 4.19      

Unscaled Zero Point Vibrational Energy (zpe) 20571.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 19589.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.28020 0.27327 0.14200

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.882 0.000 1.882
H2 -0.882 0.000 1.882
H3 0.000 2.216 0.611
H4 0.000 -2.216 0.611
H5 0.000 1.354 -1.890
H6 0.000 -1.354 -1.890
C7 0.000 1.180 0.282
C8 0.000 -1.180 0.282
C9 0.000 0.000 1.222
C10 0.000 0.738 -0.994
C11 0.000 -0.738 -0.994

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 C7 C8 C9 C10 C11
H11.76422.70212.70214.10294.10292.17482.17481.10133.09673.0967
H21.76422.70212.70214.10294.10292.17482.17481.10133.09673.0967
H32.70212.70214.43162.64524.35881.08653.41212.29852.18133.3620
H42.70212.70214.43164.35882.64523.41211.08652.29853.36202.1813
H54.10294.10292.64524.35882.70842.17863.33773.39381.08722.2764
H64.10294.10294.35882.64522.70843.33772.17863.39382.27641.0872
C72.17482.17481.08653.41212.17863.33772.36081.50911.35002.3041
C82.17482.17483.41211.08653.33772.17862.36081.50912.30411.3500
C91.10131.10132.29852.29853.39383.39381.50911.50912.33562.3356
C103.09673.09672.18133.36201.08722.27641.35002.30412.33561.4769
C113.09673.09673.36202.18132.27641.08722.30411.35002.33561.4769

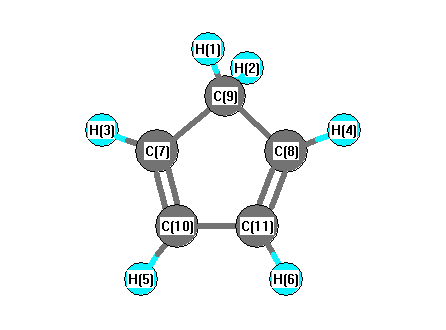
picture of 1,3-Cyclopentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C9 H2 106.450 H1 C9 C7 111.901
H1 C9 C8 111.901 H2 C9 C7 111.901
H2 C9 C8 111.901 H3 C7 C9 123.820
H3 C7 C10 126.750 H4 C8 C9 123.820
H4 C8 C11 126.750 H5 C10 C7 126.394
H5 C10 C11 124.497 H6 C11 C8 126.394
H6 C11 C10 124.497 C7 C9 C8 102.922
C7 C10 C11 109.109 C8 C11 C10 109.109
C9 C7 C10 109.430 C9 C8 C11 109.430
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability