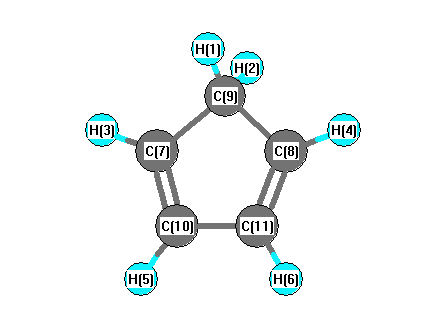
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3271 |
3111 |
2.48 |
|
|
|
| 2 |
A1 |
3246 |
3087 |
13.44 |
|
|
|
| 3 |
A1 |
3054 |
2904 |
8.39 |
|
|
|
| 4 |
A1 |
1597 |
1519 |
2.20 |
|
|
|
| 5 |
A1 |
1441 |
1370 |
4.05 |
|
|
|
| 6 |
A1 |
1424 |
1355 |
17.70 |
|
|
|
| 7 |
A1 |
1143 |
1087 |
0.17 |
|
|
|
| 8 |
A1 |
1040 |
989 |
0.02 |
|
|
|
| 9 |
A1 |
950 |
904 |
10.71 |
|
|
|
| 10 |
A1 |
815 |
775 |
0.04 |
|
|
|
| 11 |
A2 |
1134 |
1078 |
0.00 |
|
|
|
| 12 |
A2 |
961 |
913 |
0.00 |
|
|
|
| 13 |
A2 |
715 |
680 |
0.00 |
|
|
|
| 14 |
A2 |
523 |
498 |
0.00 |
|
|
|
| 15 |
B1 |
3086 |
2935 |
8.07 |
|
|
|
| 16 |
B1 |
955 |
908 |
0.20 |
|
|
|
| 17 |
B1 |
925 |
880 |
21.92 |
|
|
|
| 18 |
B1 |
689 |
655 |
63.78 |
|
|
|
| 19 |
B1 |
344 |
327 |
11.47 |
|
|
|
| 20 |
B2 |
3265 |
3105 |
22.11 |
|
|
|
| 21 |
B2 |
3235 |
3076 |
4.51 |
|
|
|
| 22 |
B2 |
1680 |
1598 |
0.47 |
|
|
|
| 23 |
B2 |
1340 |
1274 |
1.59 |
|
|
|
| 24 |
B2 |
1286 |
1223 |
1.56 |
|
|
|
| 25 |
B2 |
1125 |
1069 |
2.58 |
|
|
|
| 26 |
B2 |
993 |
944 |
12.23 |
|
|
|
| 27 |
B2 |
814 |
775 |
7.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20524.6 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 19518.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.198 |
|
|
|
| 2 |
H |
0.198 |
|
|
|
| 3 |
H |
0.163 |
|
|
|
| 4 |
H |
0.163 |
|
|
|
| 5 |
H |
0.160 |
|
|
|
| 6 |
H |
0.160 |
|
|
|
| 7 |
C |
-0.167 |
|
|
|
| 8 |
C |
-0.167 |
|
|
|
| 9 |
C |
-0.409 |
|
|
|
| 10 |
C |
-0.150 |
|
|
|
| 11 |
C |
-0.150 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.474 |
0.474 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.200 |
0.000 |
0.000 |
| y |
0.000 |
-27.964 |
0.000 |
| z |
0.000 |
0.000 |
-26.519 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.959 |
0.000 |
0.000 |
| y |
0.000 |
1.395 |
0.000 |
| z |
0.000 |
0.000 |
3.563 |
|
| Polar |
|---|
| 3z2-r2 | 7.127 |
|---|
| x2-y2 | -4.236 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.634 |
0.000 |
0.000 |
| y |
0.000 |
7.910 |
0.000 |
| z |
0.000 |
0.000 |
8.800 |
<r2> (average value of r
2) Å
2
| <r2> |
92.363 |
| (<r2>)1/2 |
9.611 |