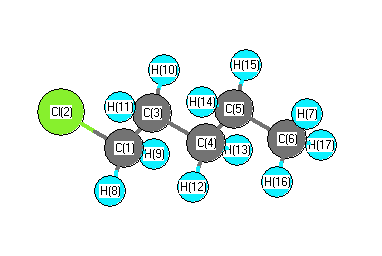
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3133 |
2997 |
35.75 |
|
|
|
| 2 |
A' |
3104 |
2970 |
26.03 |
|
|
|
| 3 |
A' |
3065 |
2932 |
23.35 |
|
|
|
| 4 |
A' |
3053 |
2921 |
36.29 |
|
|
|
| 5 |
A' |
3045 |
2913 |
30.75 |
|
|
|
| 6 |
A' |
3031 |
2900 |
9.48 |
|
|
|
| 7 |
A' |
1535 |
1469 |
7.29 |
|
|
|
| 8 |
A' |
1526 |
1460 |
0.31 |
|
|
|
| 9 |
A' |
1515 |
1449 |
2.03 |
|
|
|
| 10 |
A' |
1513 |
1447 |
0.28 |
|
|
|
| 11 |
A' |
1509 |
1444 |
0.22 |
|
|
|
| 12 |
A' |
1440 |
1378 |
3.67 |
|
|
|
| 13 |
A' |
1423 |
1361 |
0.14 |
|
|
|
| 14 |
A' |
1391 |
1331 |
20.36 |
|
|
|
| 15 |
A' |
1335 |
1277 |
12.87 |
|
|
|
| 16 |
A' |
1266 |
1211 |
7.57 |
|
|
|
| 17 |
A' |
1143 |
1094 |
2.53 |
|
|
|
| 18 |
A' |
1083 |
1036 |
0.30 |
|
|
|
| 19 |
A' |
1067 |
1021 |
7.66 |
|
|
|
| 20 |
A' |
1032 |
987 |
1.42 |
|
|
|
| 21 |
A' |
918 |
878 |
1.39 |
|
|
|
| 22 |
A' |
751 |
719 |
48.57 |
|
|
|
| 23 |
A' |
451 |
432 |
3.76 |
|
|
|
| 24 |
A' |
344 |
329 |
0.90 |
|
|
|
| 25 |
A' |
261 |
250 |
2.17 |
|
|
|
| 26 |
A' |
119 |
114 |
1.31 |
|
|
|
| 27 |
A" |
3166 |
3029 |
18.69 |
|
|
|
| 28 |
A" |
3127 |
2992 |
58.75 |
|
|
|
| 29 |
A" |
3112 |
2977 |
21.02 |
|
|
|
| 30 |
A" |
3086 |
2953 |
18.89 |
|
|
|
| 31 |
A" |
3063 |
2930 |
2.16 |
|
|
|
| 32 |
A" |
1527 |
1461 |
7.76 |
|
|
|
| 33 |
A" |
1347 |
1289 |
0.76 |
|
|
|
| 34 |
A" |
1340 |
1282 |
0.45 |
|
|
|
| 35 |
A" |
1303 |
1246 |
0.30 |
|
|
|
| 36 |
A" |
1235 |
1182 |
0.07 |
|
|
|
| 37 |
A" |
1125 |
1077 |
1.02 |
|
|
|
| 38 |
A" |
982 |
939 |
0.02 |
|
|
|
| 39 |
A" |
851 |
814 |
1.47 |
|
|
|
| 40 |
A" |
767 |
734 |
0.15 |
|
|
|
| 41 |
A" |
741 |
709 |
5.56 |
|
|
|
| 42 |
A" |
254 |
243 |
0.00 |
|
|
|
| 43 |
A" |
158 |
151 |
0.46 |
|
|
|
| 44 |
A" |
104 |
100 |
0.26 |
|
|
|
| 45 |
A" |
74 |
71 |
1.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33706.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 32247.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.427 |
|
|
-0.257 |
| 2 |
Cl |
-0.084 |
|
|
-0.179 |
| 3 |
C |
-0.306 |
|
|
0.111 |
| 4 |
C |
-0.310 |
|
|
-0.044 |
| 5 |
C |
-0.305 |
|
|
0.162 |
| 6 |
C |
-0.510 |
|
|
-0.316 |
| 7 |
H |
0.171 |
|
|
0.083 |
| 8 |
H |
0.214 |
|
|
0.144 |
| 9 |
H |
0.214 |
|
|
0.144 |
| 10 |
H |
0.182 |
|
|
0.011 |
| 11 |
H |
0.182 |
|
|
0.011 |
| 12 |
H |
0.160 |
|
|
0.012 |
| 13 |
H |
0.160 |
|
|
0.012 |
| 14 |
H |
0.163 |
|
|
-0.022 |
| 15 |
H |
0.163 |
|
|
-0.022 |
| 16 |
H |
0.167 |
|
|
0.075 |
| 17 |
H |
0.167 |
|
|
0.075 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.624 |
-1.846 |
0.000 |
2.459 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.619 |
-1.912 |
0.000 |
2.505 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.094 |
4.297 |
0.000 |
| y |
4.297 |
-48.681 |
0.000 |
| z |
0.000 |
0.000 |
-45.200 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.154 |
4.297 |
0.000 |
| y |
4.297 |
-1.033 |
0.000 |
| z |
0.000 |
0.000 |
4.187 |
|
| Polar |
|---|
| 3z2-r2 | 8.375 |
|---|
| x2-y2 | -1.414 |
|---|
| xy | 4.297 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
405.231 |
| (<r2>)1/2 |
20.130 |