return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H11Cl (Pentane, 1-chloro-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-656.932795
Energy at 298.15K-656.944901
HF Energy-656.674149
Nuclear repulsion energy282.441137
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3154 2993 36.54      
2 A' 3130 2971 24.80      
3 A' 3086 2930 23.01      
4 A' 3077 2920 32.17      
5 A' 3067 2911 29.01      
6 A' 3052 2897 11.01      
7 A' 1563 1483 5.17      
8 A' 1553 1474 0.08      
9 A' 1542 1463 1.66      
10 A' 1539 1461 0.79      
11 A' 1535 1457 0.16      
12 A' 1464 1389 2.37      
13 A' 1444 1371 0.14      
14 A' 1413 1341 21.67      
15 A' 1356 1287 12.69      
16 A' 1286 1220 9.73      
17 A' 1156 1098 1.99      
18 A' 1090 1035 0.22      
19 A' 1075 1020 6.50      
20 A' 1038 985 0.84      
21 A' 926 879 1.28      
22 A' 756 717 46.73      
23 A' 457 433 3.57      
24 A' 348 330 0.91      
25 A' 264 250 2.04      
26 A' 119 113 1.29      
27 A" 3191 3029 18.15      
28 A" 3148 2989 58.91      
29 A" 3132 2973 22.78      
30 A" 3106 2949 19.55      
31 A" 3083 2927 2.79      
32 A" 1552 1473 6.49      
33 A" 1365 1296 0.81      
34 A" 1358 1289 0.40      
35 A" 1323 1255 0.23      
36 A" 1258 1194 0.10      
37 A" 1142 1084 0.89      
38 A" 996 946 0.01      
39 A" 864 821 1.02      
40 A" 778 739 0.08      
41 A" 751 713 4.04      
42 A" 256 243 0.00      
43 A" 157 149 0.49      
44 A" 105 99 0.24      
45 A" 72 68 1.37      

Unscaled Zero Point Vibrational Energy (zpe) 34062.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 32331.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.50143 0.02617 0.02546

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.508 0.120 0.000
Cl2 -2.477 1.628 0.000
C3 0.000 0.385 0.000
C4 0.818 -0.924 0.000
C5 2.339 -0.682 0.000
C6 3.154 -1.990 0.000
H7 4.230 -1.797 0.000
H8 -1.795 -0.450 0.884
H9 -1.795 -0.450 -0.884
H10 0.252 0.977 -0.884
H11 0.252 0.977 0.884
H12 0.555 -1.517 0.883
H13 0.555 -1.517 -0.883
H14 2.601 -0.089 0.882
H15 2.601 -0.089 -0.882
H16 2.922 -2.587 0.885
H17 2.922 -2.587 -0.885

Atom - Atom Distances (Å)
  C1 Cl2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.79301.53112.54963.92955.11726.05011.09091.09092.14822.14822.77782.77784.20824.20825.26635.2663
Cl21.79302.77214.16825.34176.69357.53172.35972.35972.94252.94254.45754.45755.43335.43336.90686.9068
C31.53112.77211.54322.57053.94784.75972.16862.16861.09351.09352.16902.16902.78732.78734.26044.2604
C42.54964.16821.54321.53962.56763.52222.79952.79952.17102.17101.09591.09592.15762.15762.82402.8240
C53.92955.34172.57051.53961.54062.19554.23404.23402.80832.80832.15822.15821.09491.09492.17992.1799
C65.11726.69353.94782.56761.54061.09355.25835.25834.24264.24262.78542.78542.16692.16691.09301.0930
H76.05017.53174.75973.52222.19551.09356.23756.23754.92924.92923.79043.79042.51942.51941.76691.7669
H81.09092.35972.16862.79954.23405.25836.23751.76903.05902.49592.58123.12844.41174.75215.17855.4725
H91.09092.35972.16862.79954.23405.25836.23751.76902.49593.05903.12842.58124.75214.41175.47255.1785
H102.14822.94251.09352.17102.80834.24264.92923.05902.49591.76813.07142.51213.12582.57944.79134.4527
H112.14822.94251.09352.17102.80834.24264.92922.49593.05901.76812.51213.07142.57943.12584.45274.7913
H122.77784.45752.16901.09592.15822.78543.79042.58123.12843.07142.51211.76612.49573.05652.59733.1419
H132.77784.45752.16901.09592.15822.78543.79043.12842.58122.51213.07141.76613.05652.49573.14192.5973
H144.20825.43332.78732.15761.09492.16692.51944.41174.75213.12582.57942.49573.05651.76322.51913.0768
H154.20825.43332.78732.15761.09492.16692.51944.75214.41172.57943.12583.05652.49571.76323.07682.5191
H165.26636.90684.26042.82402.17991.09301.76695.17855.47254.79134.45272.59733.14192.51913.07681.7699
H175.26636.90684.26042.82402.17991.09301.76695.47255.17854.45274.79133.14192.59733.07682.51911.7699

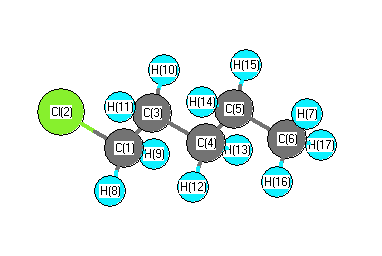
picture of Pentane, 1-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C4 112.059 C1 C3 H10 108.717
C1 C3 H11 108.717 Cl2 C1 C3 112.774
Cl2 C1 H8 107.296 Cl2 C1 H9 107.296
C3 C1 H8 110.477 C3 C1 H9 110.477
C3 C4 C5 112.990 C3 C4 H12 109.378
C3 C4 H13 109.378 C4 C3 H10 109.676
C4 C3 H11 109.676 C4 C5 C6 112.936
C4 C5 H14 108.792 C4 C5 H15 108.792
C5 C4 H12 108.783 C5 C4 H13 108.783
C5 C6 H7 111.794 C5 C6 H16 110.576
C5 C6 H17 110.576 C6 C5 H14 109.450
C6 C5 H15 109.450 H7 C6 H16 107.818
H7 C6 H17 107.818 H8 C1 H9 108.344
H10 C3 H11 107.895 H12 C4 H13 107.373
H14 C5 H15 107.258 H16 C6 H17 108.120
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.374      
2 Cl -0.104      
3 C -0.277      
4 C -0.282      
5 C -0.273      
6 C -0.461      
7 H 0.155      
8 H 0.199      
9 H 0.199      
10 H 0.167      
11 H 0.167      
12 H 0.144      
13 H 0.144      
14 H 0.147      
15 H 0.147      
16 H 0.151      
17 H 0.151      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.697 -1.953 0.000
y -1.953 9.643 0.000
z 0.000 0.000 7.764


<r2> (average value of r2) Å2
<r2> 405.753
(<r2>)1/2 20.143