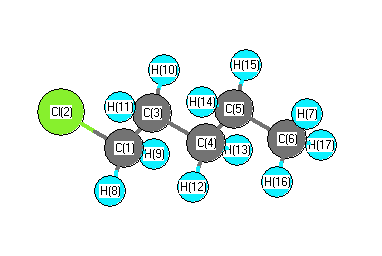
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3060 |
3009 |
33.26 |
|
|
|
| 2 |
A' |
3022 |
2972 |
26.80 |
|
|
|
| 3 |
A' |
2986 |
2937 |
23.70 |
|
|
|
| 4 |
A' |
2980 |
2931 |
34.18 |
|
|
|
| 5 |
A' |
2966 |
2918 |
29.69 |
|
|
|
| 6 |
A' |
2951 |
2903 |
9.59 |
|
|
|
| 7 |
A' |
1493 |
1468 |
6.23 |
|
|
|
| 8 |
A' |
1483 |
1459 |
0.29 |
|
|
|
| 9 |
A' |
1471 |
1446 |
1.57 |
|
|
|
| 10 |
A' |
1467 |
1443 |
0.36 |
|
|
|
| 11 |
A' |
1464 |
1440 |
0.16 |
|
|
|
| 12 |
A' |
1391 |
1368 |
2.67 |
|
|
|
| 13 |
A' |
1369 |
1346 |
0.09 |
|
|
|
| 14 |
A' |
1341 |
1319 |
21.74 |
|
|
|
| 15 |
A' |
1289 |
1268 |
12.12 |
|
|
|
| 16 |
A' |
1223 |
1203 |
8.91 |
|
|
|
| 17 |
A' |
1110 |
1091 |
2.17 |
|
|
|
| 18 |
A' |
1056 |
1038 |
0.63 |
|
|
|
| 19 |
A' |
1039 |
1022 |
8.96 |
|
|
|
| 20 |
A' |
1006 |
989 |
1.71 |
|
|
|
| 21 |
A' |
892 |
877 |
1.32 |
|
|
|
| 22 |
A' |
722 |
710 |
48.77 |
|
|
|
| 23 |
A' |
438 |
431 |
4.20 |
|
|
|
| 24 |
A' |
333 |
328 |
0.85 |
|
|
|
| 25 |
A' |
254 |
249 |
2.36 |
|
|
|
| 26 |
A' |
112 |
110 |
1.21 |
|
|
|
| 27 |
A" |
3082 |
3031 |
20.25 |
|
|
|
| 28 |
A" |
3052 |
3001 |
51.02 |
|
|
|
| 29 |
A" |
3031 |
2981 |
22.20 |
|
|
|
| 30 |
A" |
3007 |
2957 |
19.23 |
|
|
|
| 31 |
A" |
2982 |
2933 |
2.31 |
|
|
|
| 32 |
A" |
1482 |
1457 |
7.50 |
|
|
|
| 33 |
A" |
1306 |
1285 |
0.81 |
|
|
|
| 34 |
A" |
1297 |
1275 |
0.35 |
|
|
|
| 35 |
A" |
1263 |
1242 |
0.35 |
|
|
|
| 36 |
A" |
1197 |
1178 |
0.06 |
|
|
|
| 37 |
A" |
1087 |
1069 |
1.15 |
|
|
|
| 38 |
A" |
951 |
935 |
0.02 |
|
|
|
| 39 |
A" |
825 |
811 |
1.60 |
|
|
|
| 40 |
A" |
748 |
735 |
0.20 |
|
|
|
| 41 |
A" |
725 |
713 |
5.82 |
|
|
|
| 42 |
A" |
245 |
241 |
0.00 |
|
|
|
| 43 |
A" |
149 |
147 |
0.46 |
|
|
|
| 44 |
A" |
101 |
99 |
0.23 |
|
|
|
| 45 |
A" |
66 |
64 |
1.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32754.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 32214.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.411 |
|
|
|
| 2 |
Cl |
-0.080 |
|
|
|
| 3 |
C |
-0.277 |
|
|
|
| 4 |
C |
-0.284 |
|
|
|
| 5 |
C |
-0.280 |
|
|
|
| 6 |
C |
-0.484 |
|
|
|
| 7 |
H |
0.161 |
|
|
|
| 8 |
H |
0.202 |
|
|
|
| 9 |
H |
0.202 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
| 12 |
H |
0.148 |
|
|
|
| 13 |
H |
0.148 |
|
|
|
| 14 |
H |
0.151 |
|
|
|
| 15 |
H |
0.151 |
|
|
|
| 16 |
H |
0.158 |
|
|
|
| 17 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.634 |
-1.812 |
0.000 |
2.440 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.169 |
4.143 |
-0.003 |
| y |
4.143 |
-48.744 |
0.000 |
| z |
-0.003 |
0.000 |
-45.447 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.074 |
4.143 |
-0.003 |
| y |
4.143 |
-0.936 |
0.000 |
| z |
-0.003 |
0.000 |
4.010 |
|
| Polar |
|---|
| 3z2-r2 | 8.020 |
|---|
| x2-y2 | -1.425 |
|---|
| xy | 4.143 |
|---|
| xz | -0.003 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.451 |
-2.168 |
0.000 |
| y |
-2.168 |
10.044 |
-0.001 |
| z |
0.000 |
-0.001 |
7.994 |
<r2> (average value of r
2) Å
2
| <r2> |
409.023 |
| (<r2>)1/2 |
20.224 |