Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -322.511515 |
| Energy at 298.15K | -322.520506 |
| HF Energy | -321.739921 |
| Nuclear repulsion energy | 236.596076 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3258 |
3010 |
28.08 |
|
|
|
| 2 |
A' |
3192 |
2948 |
37.59 |
|
|
|
| 3 |
A' |
3186 |
2943 |
13.13 |
|
|
|
| 4 |
A' |
3172 |
2930 |
16.68 |
|
|
|
| 5 |
A' |
1892 |
1747 |
190.38 |
|
|
|
| 6 |
A' |
1627 |
1503 |
10.58 |
|
|
|
| 7 |
A' |
1611 |
1488 |
3.55 |
|
|
|
| 8 |
A' |
1597 |
1475 |
1.58 |
|
|
|
| 9 |
A' |
1530 |
1414 |
9.58 |
|
|
|
| 10 |
A' |
1522 |
1406 |
18.63 |
|
|
|
| 11 |
A' |
1423 |
1314 |
10.26 |
|
|
|
| 12 |
A' |
1243 |
1148 |
10.39 |
|
|
|
| 13 |
A' |
1170 |
1080 |
80.52 |
|
|
|
| 14 |
A' |
1117 |
1032 |
8.11 |
|
|
|
| 15 |
A' |
1058 |
977 |
537.06 |
|
|
|
| 16 |
A' |
970 |
896 |
16.24 |
|
|
|
| 17 |
A' |
742 |
686 |
0.18 |
|
|
|
| 18 |
A' |
429 |
397 |
1.48 |
|
|
|
| 19 |
A' |
376 |
347 |
0.06 |
|
|
|
| 20 |
A' |
167 |
154 |
0.64 |
|
|
|
| 21 |
A" |
3256 |
3008 |
76.99 |
|
|
|
| 22 |
A" |
3239 |
2992 |
0.06 |
|
|
|
| 23 |
A" |
3224 |
2978 |
1.70 |
|
|
|
| 24 |
A" |
1603 |
1481 |
7.90 |
|
|
|
| 25 |
A" |
1402 |
1295 |
0.01 |
|
|
|
| 26 |
A" |
1352 |
1248 |
0.43 |
|
|
|
| 27 |
A" |
1268 |
1171 |
1.39 |
|
|
|
| 28 |
A" |
953 |
880 |
1.46 |
|
|
|
| 29 |
A" |
804 |
742 |
1.49 |
|
|
|
| 30 |
A" |
249 |
230 |
0.02 |
|
|
|
| 31 |
A" |
215 |
198 |
0.78 |
|
|
|
| 32 |
A" |
108 |
100 |
2.24 |
|
|
|
| 33 |
A" |
21 |
19 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24485.2 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 22617.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.980 |
2.127 |
0.000 |
| C2 |
-1.509 |
0.679 |
0.000 |
| C3 |
0.000 |
0.583 |
0.000 |
| O4 |
0.333 |
-0.809 |
0.000 |
| N5 |
1.684 |
-0.977 |
0.000 |
| O6 |
1.966 |
-2.117 |
0.000 |
| H7 |
-3.066 |
2.179 |
0.000 |
| H8 |
-1.623 |
2.659 |
0.880 |
| H9 |
-1.623 |
2.659 |
-0.880 |
| H10 |
-1.892 |
0.155 |
-0.874 |
| H11 |
-1.892 |
0.155 |
0.874 |
| H12 |
0.426 |
1.054 |
0.883 |
| H13 |
0.426 |
1.054 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5231 | 2.5110 | 3.7376 | 4.8022 | 5.7949 | 1.0870 | 1.0888 | 1.0888 | 2.1586 | 2.1586 | 2.7785 | 2.7785 |
C2 | 1.5231 | | 1.5119 | 2.3675 | 3.5967 | 4.4596 | 2.1621 | 2.1704 | 2.1704 | 1.0886 | 1.0886 | 2.1600 | 2.1600 | C3 | 2.5110 | 1.5119 | | 1.4310 | 2.2955 | 3.3396 | 3.4564 | 2.7785 | 2.7785 | 2.1278 | 2.1278 | 1.0883 | 1.0883 | O4 | 3.7376 | 2.3675 | 1.4310 | | 1.3613 | 2.0921 | 4.5254 | 4.0778 | 4.0778 | 2.5780 | 2.5780 | 2.0642 | 2.0642 | N5 | 4.8022 | 3.5967 | 2.2955 | 1.3613 | | 1.1739 | 5.7028 | 4.9934 | 4.9934 | 3.8518 | 3.8518 | 2.5477 | 2.5477 | O6 | 5.7949 | 4.4596 | 3.3396 | 2.0921 | 1.1739 | | 6.6160 | 6.0386 | 6.0386 | 4.5620 | 4.5620 | 3.6344 | 3.6344 | H7 | 1.0870 | 2.1621 | 3.4564 | 4.5254 | 5.7028 | 6.6160 | | 1.7571 | 1.7571 | 2.4972 | 2.4972 | 3.7733 | 3.7733 | H8 | 1.0888 | 2.1704 | 2.7785 | 4.0778 | 4.9934 | 6.0386 | 1.7571 | | 1.7604 | 3.0691 | 2.5182 | 2.6026 | 3.1438 | H9 | 1.0888 | 2.1704 | 2.7785 | 4.0778 | 4.9934 | 6.0386 | 1.7571 | 1.7604 | | 2.5182 | 3.0691 | 3.1438 | 2.6026 | H10 | 2.1586 | 1.0886 | 2.1278 | 2.5780 | 3.8518 | 4.5620 | 2.4972 | 3.0691 | 2.5182 | | 1.7486 | 3.0450 | 2.4865 | H11 | 2.1586 | 1.0886 | 2.1278 | 2.5780 | 3.8518 | 4.5620 | 2.4972 | 2.5182 | 3.0691 | 1.7486 | | 2.4865 | 3.0450 | H12 | 2.7785 | 2.1600 | 1.0883 | 2.0642 | 2.5477 | 3.6344 | 3.7733 | 2.6026 | 3.1438 | 3.0450 | 2.4865 | | 1.7668 | H13 | 2.7785 | 2.1600 | 1.0883 | 2.0642 | 2.5477 | 3.6344 | 3.7733 | 3.1438 | 2.6026 | 2.4865 | 3.0450 | 1.7668 | |
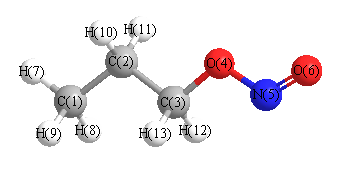 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.651 |
|
C1 |
C2 |
H10 |
110.372 |
| C1 |
C2 |
H11 |
110.372 |
|
C2 |
C1 |
H7 |
110.752 |
| C2 |
C1 |
H8 |
111.306 |
|
C2 |
C1 |
H9 |
111.306 |
| C2 |
C3 |
O4 |
107.091 |
|
C2 |
C3 |
H12 |
111.299 |
| C2 |
C3 |
H13 |
111.299 |
|
C3 |
C2 |
H10 |
108.721 |
| C3 |
C2 |
H11 |
108.721 |
|
C3 |
O4 |
N5 |
110.567 |
| O4 |
C3 |
H12 |
109.292 |
|
O4 |
C3 |
H13 |
109.292 |
| O4 |
N5 |
O6 |
111.002 |
|
H7 |
C1 |
H8 |
107.716 |
| H7 |
C1 |
H9 |
107.716 |
|
H8 |
C1 |
H9 |
107.883 |
| H10 |
C2 |
H11 |
106.863 |
|
H12 |
C3 |
H13 |
108.531 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -322.512425 |
| Energy at 298.15K | -322.521696 |
| HF Energy | -321.739749 |
| Nuclear repulsion energy | 243.904185 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3264 |
3015 |
30.70 |
|
|
|
| 2 |
A |
3260 |
3011 |
28.43 |
|
|
|
| 3 |
A |
3250 |
3002 |
30.75 |
|
|
|
| 4 |
A |
3220 |
2975 |
15.86 |
|
|
|
| 5 |
A |
3201 |
2956 |
22.54 |
|
|
|
| 6 |
A |
3176 |
2934 |
9.41 |
|
|
|
| 7 |
A |
3174 |
2932 |
28.99 |
|
|
|
| 8 |
A |
1895 |
1750 |
199.60 |
|
|
|
| 9 |
A |
1610 |
1487 |
9.25 |
|
|
|
| 10 |
A |
1608 |
1485 |
5.74 |
|
|
|
| 11 |
A |
1596 |
1474 |
3.70 |
|
|
|
| 12 |
A |
1586 |
1465 |
1.25 |
|
|
|
| 13 |
A |
1527 |
1411 |
5.83 |
|
|
|
| 14 |
A |
1507 |
1392 |
15.30 |
|
|
|
| 15 |
A |
1476 |
1364 |
1.68 |
|
|
|
| 16 |
A |
1389 |
1283 |
10.62 |
|
|
|
| 17 |
A |
1375 |
1270 |
15.65 |
|
|
|
| 18 |
A |
1274 |
1177 |
21.98 |
|
|
|
| 19 |
A |
1199 |
1107 |
47.10 |
|
|
|
| 20 |
A |
1171 |
1081 |
38.84 |
|
|
|
| 21 |
A |
1078 |
996 |
23.26 |
|
|
|
| 22 |
A |
1039 |
960 |
405.42 |
|
|
|
| 23 |
A |
964 |
890 |
72.69 |
|
|
|
| 24 |
A |
929 |
859 |
4.90 |
|
|
|
| 25 |
A |
840 |
776 |
1.71 |
|
|
|
| 26 |
A |
689 |
636 |
0.68 |
|
|
|
| 27 |
A |
489 |
452 |
2.57 |
|
|
|
| 28 |
A |
412 |
381 |
2.56 |
|
|
|
| 29 |
A |
299 |
277 |
0.42 |
|
|
|
| 30 |
A |
259 |
239 |
0.88 |
|
|
|
| 31 |
A |
182 |
169 |
0.55 |
|
|
|
| 32 |
A |
118 |
109 |
1.27 |
|
|
|
| 33 |
A |
49 |
46 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24551.9 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 22678.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.171 |
-0.913 |
0.184 |
| C2 |
-1.738 |
0.458 |
-0.318 |
| C3 |
-0.393 |
0.893 |
0.222 |
| O4 |
0.576 |
-0.034 |
-0.280 |
| N5 |
1.775 |
0.208 |
0.311 |
| O6 |
2.536 |
-0.559 |
-0.145 |
| H7 |
-3.148 |
-1.177 |
-0.214 |
| H8 |
-2.238 |
-0.927 |
1.270 |
| H9 |
-1.463 |
-1.679 |
-0.121 |
| H10 |
-1.696 |
0.466 |
-1.405 |
| H11 |
-2.466 |
1.213 |
-0.023 |
| H12 |
-0.129 |
1.892 |
-0.117 |
| H13 |
-0.372 |
0.871 |
1.309 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5227 | 2.5345 | 2.9212 | 4.1046 | 4.7316 | 1.0874 | 1.0889 | 1.0859 | 2.1570 | 2.1569 | 3.4823 | 2.7723 |
C2 | 1.5227 | | 1.5123 | 2.3655 | 3.5773 | 4.3960 | 2.1619 | 2.1655 | 2.1629 | 1.0886 | 1.0900 | 2.1640 | 2.1633 | C3 | 2.5345 | 1.5123 | | 1.4318 | 2.2757 | 3.2894 | 3.4733 | 2.7955 | 2.8062 | 2.1275 | 2.1115 | 1.0871 | 1.0875 | O4 | 2.9212 | 2.3655 | 1.4318 | | 1.3586 | 2.0333 | 3.8961 | 3.3346 | 2.6247 | 2.5843 | 3.2978 | 2.0572 | 2.0595 | N5 | 4.1046 | 3.5773 | 2.2757 | 1.3586 | | 1.1723 | 5.1415 | 4.2800 | 3.7731 | 3.8808 | 4.3714 | 2.5773 | 2.4589 | O6 | 4.7316 | 4.3960 | 3.2894 | 2.0333 | 1.1723 | | 5.7179 | 4.9930 | 4.1531 | 4.5328 | 5.3078 | 3.6203 | 3.5514 | H7 | 1.0874 | 2.1619 | 3.4733 | 3.8961 | 5.1415 | 5.7179 | | 1.7589 | 1.7602 | 2.4956 | 2.4934 | 4.3059 | 3.7710 | H8 | 1.0889 | 2.1655 | 2.7955 | 3.3346 | 4.2800 | 4.9930 | 1.7589 | | 1.7608 | 3.0648 | 2.5111 | 3.7836 | 2.5917 | H9 | 1.0859 | 2.1629 | 2.8062 | 2.6247 | 3.7731 | 4.1531 | 1.7602 | 1.7608 | | 2.5107 | 3.0625 | 3.8114 | 3.1201 | H10 | 2.1570 | 1.0886 | 2.1275 | 2.5843 | 3.8808 | 4.5328 | 2.4956 | 3.0648 | 2.5107 | | 1.7499 | 2.4795 | 3.0467 | H11 | 2.1569 | 1.0900 | 2.1115 | 3.2978 | 4.3714 | 5.3078 | 2.4934 | 2.5111 | 3.0625 | 1.7499 | | 2.4350 | 2.5050 | H12 | 3.4823 | 2.1640 | 1.0871 | 2.0572 | 2.5773 | 3.6203 | 4.3059 | 3.7836 | 3.8114 | 2.4795 | 2.4350 | | 1.7698 | H13 | 2.7723 | 2.1633 | 1.0875 | 2.0595 | 2.4589 | 3.5514 | 3.7710 | 2.5917 | 3.1201 | 3.0467 | 2.5050 | 1.7698 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.247 |
|
C1 |
C2 |
H10 |
110.273 |
| C1 |
C2 |
H11 |
110.186 |
|
C2 |
C1 |
H7 |
110.735 |
| C2 |
C1 |
H8 |
110.937 |
|
C2 |
C1 |
H9 |
110.913 |
| C2 |
C3 |
O4 |
106.897 |
|
C2 |
C3 |
H12 |
111.671 |
| C2 |
C3 |
H13 |
111.587 |
|
C3 |
C2 |
H10 |
108.669 |
| C3 |
C2 |
H11 |
107.353 |
|
C3 |
O4 |
N5 |
109.252 |
| O4 |
C3 |
H12 |
108.744 |
|
O4 |
C3 |
H13 |
108.905 |
| O4 |
N5 |
O6 |
106.677 |
|
H7 |
C1 |
H8 |
107.845 |
| H7 |
C1 |
H9 |
108.171 |
|
H8 |
C1 |
H9 |
108.121 |
| H10 |
C2 |
H11 |
106.871 |
|
H12 |
C3 |
H13 |
108.949 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
