Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -323.515233 |
| Energy at 298.15K | |
| HF Energy | -323.515233 |
| Nuclear repulsion energy | 234.250576 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3144 |
3008 |
26.53 |
|
|
|
| 2 |
A' |
3071 |
2938 |
39.88 |
|
|
|
| 3 |
A' |
3059 |
2927 |
30.66 |
|
|
|
| 4 |
A' |
3056 |
2924 |
3.14 |
|
|
|
| 5 |
A' |
1782 |
1705 |
215.39 |
|
|
|
| 6 |
A' |
1541 |
1474 |
12.66 |
|
|
|
| 7 |
A' |
1530 |
1464 |
1.52 |
|
|
|
| 8 |
A' |
1518 |
1453 |
2.14 |
|
|
|
| 9 |
A' |
1440 |
1377 |
4.22 |
|
|
|
| 10 |
A' |
1430 |
1368 |
19.70 |
|
|
|
| 11 |
A' |
1342 |
1284 |
11.98 |
|
|
|
| 12 |
A' |
1173 |
1122 |
5.30 |
|
|
|
| 13 |
A' |
1089 |
1042 |
27.23 |
|
|
|
| 14 |
A' |
1060 |
1014 |
4.07 |
|
|
|
| 15 |
A' |
944 |
903 |
180.46 |
|
|
|
| 16 |
A' |
882 |
843 |
342.82 |
|
|
|
| 17 |
A' |
676 |
647 |
24.87 |
|
|
|
| 18 |
A' |
392 |
375 |
0.09 |
|
|
|
| 19 |
A' |
353 |
337 |
0.87 |
|
|
|
| 20 |
A' |
155 |
148 |
0.44 |
|
|
|
| 21 |
A" |
3136 |
3001 |
67.02 |
|
|
|
| 22 |
A" |
3117 |
2982 |
6.59 |
|
|
|
| 23 |
A" |
3098 |
2964 |
3.52 |
|
|
|
| 24 |
A" |
1527 |
1461 |
8.60 |
|
|
|
| 25 |
A" |
1331 |
1274 |
0.00 |
|
|
|
| 26 |
A" |
1278 |
1222 |
0.27 |
|
|
|
| 27 |
A" |
1193 |
1141 |
0.56 |
|
|
|
| 28 |
A" |
902 |
863 |
1.47 |
|
|
|
| 29 |
A" |
766 |
733 |
2.83 |
|
|
|
| 30 |
A" |
240 |
230 |
0.02 |
|
|
|
| 31 |
A" |
216 |
207 |
0.77 |
|
|
|
| 32 |
A" |
99 |
95 |
1.48 |
|
|
|
| 33 |
A" |
66i |
63i |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23236.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22230.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.945 |
2.187 |
0.000 |
| C2 |
-1.511 |
0.723 |
0.000 |
| C3 |
0.000 |
0.581 |
0.000 |
| O4 |
0.303 |
-0.825 |
0.000 |
| N5 |
1.691 |
-1.022 |
0.000 |
| O6 |
1.950 |
-2.174 |
0.000 |
| H7 |
-3.037 |
2.269 |
0.000 |
| H8 |
-1.573 |
2.716 |
0.885 |
| H9 |
-1.573 |
2.716 |
-0.885 |
| H10 |
-1.914 |
0.206 |
-0.880 |
| H11 |
-1.914 |
0.206 |
0.880 |
| H12 |
0.442 |
1.047 |
0.889 |
| H13 |
0.442 |
1.047 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5274 | 2.5223 | 3.7589 | 4.8495 | 5.8475 | 1.0946 | 1.0965 | 1.0965 | 2.1674 | 2.1674 | 2.7911 | 2.7911 |
C2 | 1.5274 | | 1.5177 | 2.3849 | 3.6465 | 4.5134 | 2.1720 | 2.1821 | 2.1821 | 1.0968 | 1.0968 | 2.1706 | 2.1706 | C3 | 2.5223 | 1.5177 | | 1.4388 | 2.3300 | 3.3757 | 3.4740 | 2.7956 | 2.7956 | 2.1399 | 2.1399 | 1.0969 | 1.0969 | O4 | 3.7589 | 2.3849 | 1.4388 | | 1.4018 | 2.1287 | 4.5528 | 4.1042 | 4.1042 | 2.5993 | 2.5993 | 2.0772 | 2.0772 | N5 | 4.8495 | 3.6465 | 2.3300 | 1.4018 | | 1.1815 | 5.7601 | 5.0405 | 5.0405 | 3.9093 | 3.9093 | 2.5746 | 2.5746 | O6 | 5.8475 | 4.5134 | 3.3757 | 2.1287 | 1.1815 | | 6.6788 | 6.0917 | 6.0917 | 4.6233 | 4.6233 | 3.6660 | 3.6660 | H7 | 1.0946 | 2.1720 | 3.4740 | 4.5528 | 5.7601 | 6.6788 | | 1.7683 | 1.7683 | 2.5072 | 2.5072 | 3.7930 | 3.7930 | H8 | 1.0965 | 2.1821 | 2.7956 | 4.1042 | 5.0405 | 6.0917 | 1.7683 | | 1.7710 | 3.0870 | 2.5326 | 2.6167 | 3.1617 | H9 | 1.0965 | 2.1821 | 2.7956 | 4.1042 | 5.0405 | 6.0917 | 1.7683 | 1.7710 | | 2.5326 | 3.0870 | 3.1617 | 2.6167 | H10 | 2.1674 | 1.0968 | 2.1399 | 2.5993 | 3.9093 | 4.6233 | 2.5072 | 3.0870 | 2.5326 | | 1.7594 | 3.0642 | 2.5020 | H11 | 2.1674 | 1.0968 | 2.1399 | 2.5993 | 3.9093 | 4.6233 | 2.5072 | 2.5326 | 3.0870 | 1.7594 | | 2.5020 | 3.0642 | H12 | 2.7911 | 2.1706 | 1.0969 | 2.0772 | 2.5746 | 3.6660 | 3.7930 | 2.6167 | 3.1617 | 3.0642 | 2.5020 | | 1.7784 | H13 | 2.7911 | 2.1706 | 1.0969 | 2.0772 | 2.5746 | 3.6660 | 3.7930 | 3.1617 | 2.6167 | 2.5020 | 3.0642 | 1.7784 | |
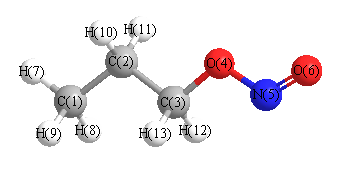 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.854 |
|
C1 |
C2 |
H10 |
110.283 |
| C1 |
C2 |
H11 |
110.283 |
|
C2 |
C1 |
H7 |
110.783 |
| C2 |
C1 |
H8 |
111.469 |
|
C2 |
C1 |
H9 |
111.469 |
| C2 |
C3 |
O4 |
107.512 |
|
C2 |
C3 |
H12 |
111.219 |
| C2 |
C3 |
H13 |
111.219 |
|
C3 |
C2 |
H10 |
108.801 |
| C3 |
C2 |
H11 |
108.801 |
|
C3 |
O4 |
N5 |
110.210 |
| O4 |
C3 |
H12 |
109.268 |
|
O4 |
C3 |
H13 |
109.268 |
| O4 |
N5 |
O6 |
110.688 |
|
H7 |
C1 |
H8 |
107.610 |
| H7 |
C1 |
H9 |
107.610 |
|
H8 |
C1 |
H9 |
107.715 |
| H10 |
C2 |
H11 |
106.662 |
|
H12 |
C3 |
H13 |
108.322 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.522 |
|
|
|
| 2 |
C |
-0.336 |
|
|
|
| 3 |
C |
-0.066 |
|
|
|
| 4 |
O |
-0.368 |
|
|
|
| 5 |
N |
0.293 |
|
|
|
| 6 |
O |
-0.251 |
|
|
|
| 7 |
H |
0.182 |
|
|
|
| 8 |
H |
0.173 |
|
|
|
| 9 |
H |
0.173 |
|
|
|
| 10 |
H |
0.184 |
|
|
|
| 11 |
H |
0.184 |
|
|
|
| 12 |
H |
0.176 |
|
|
|
| 13 |
H |
0.176 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.500 |
2.502 |
0.000 |
2.917 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.447 |
1.684 |
0.000 |
| y |
1.684 |
-38.971 |
0.000 |
| z |
0.000 |
0.000 |
-34.345 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.789 |
1.684 |
0.000 |
| y |
1.684 |
-3.075 |
0.000 |
| z |
0.000 |
0.000 |
3.864 |
|
| Polar |
|---|
| 3z2-r2 | 7.728 |
|---|
| x2-y2 | 1.524 |
|---|
| xy | 1.684 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.912 |
-2.130 |
0.000 |
| y |
-2.130 |
8.082 |
0.000 |
| z |
0.000 |
0.000 |
5.445 |
<r2> (average value of r
2) Å
2
| <r2> |
243.834 |
| (<r2>)1/2 |
15.615 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -323.517052 |
| Energy at 298.15K | -323.526074 |
| HF Energy | -323.517052 |
| Nuclear repulsion energy | 241.131552 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3148 |
3011 |
26.72 |
|
|
|
| 2 |
A |
3137 |
3001 |
37.39 |
|
|
|
| 3 |
A |
3122 |
2987 |
23.37 |
|
|
|
| 4 |
A |
3099 |
2965 |
14.75 |
|
|
|
| 5 |
A |
3065 |
2932 |
28.75 |
|
|
|
| 6 |
A |
3061 |
2929 |
25.04 |
|
|
|
| 7 |
A |
3056 |
2924 |
13.80 |
|
|
|
| 8 |
A |
1789 |
1712 |
235.29 |
|
|
|
| 9 |
A |
1532 |
1466 |
8.12 |
|
|
|
| 10 |
A |
1526 |
1460 |
8.82 |
|
|
|
| 11 |
A |
1510 |
1444 |
1.93 |
|
|
|
| 12 |
A |
1503 |
1438 |
2.81 |
|
|
|
| 13 |
A |
1440 |
1378 |
7.18 |
|
|
|
| 14 |
A |
1415 |
1353 |
12.42 |
|
|
|
| 15 |
A |
1394 |
1334 |
3.17 |
|
|
|
| 16 |
A |
1318 |
1261 |
3.02 |
|
|
|
| 17 |
A |
1301 |
1245 |
9.86 |
|
|
|
| 18 |
A |
1198 |
1146 |
8.26 |
|
|
|
| 19 |
A |
1128 |
1079 |
8.28 |
|
|
|
| 20 |
A |
1100 |
1053 |
24.23 |
|
|
|
| 21 |
A |
1018 |
974 |
8.07 |
|
|
|
| 22 |
A |
944 |
903 |
112.71 |
|
|
|
| 23 |
A |
887 |
848 |
3.29 |
|
|
|
| 24 |
A |
864 |
827 |
329.65 |
|
|
|
| 25 |
A |
789 |
754 |
16.03 |
|
|
|
| 26 |
A |
623 |
596 |
23.74 |
|
|
|
| 27 |
A |
457 |
437 |
5.29 |
|
|
|
| 28 |
A |
383 |
366 |
3.04 |
|
|
|
| 29 |
A |
284 |
272 |
0.74 |
|
|
|
| 30 |
A |
242 |
231 |
0.84 |
|
|
|
| 31 |
A |
186 |
178 |
0.44 |
|
|
|
| 32 |
A |
110 |
105 |
0.87 |
|
|
|
| 33 |
A |
59 |
57 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23342.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22332.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.200 |
-0.919 |
0.183 |
| C2 |
-1.759 |
0.458 |
-0.314 |
| C3 |
-0.405 |
0.893 |
0.221 |
| O4 |
0.579 |
-0.028 |
-0.289 |
| N5 |
1.800 |
0.206 |
0.316 |
| O6 |
2.580 |
-0.559 |
-0.144 |
| H7 |
-3.185 |
-1.180 |
-0.218 |
| H8 |
-2.267 |
-0.941 |
1.277 |
| H9 |
-1.491 |
-1.692 |
-0.128 |
| H10 |
-1.726 |
0.475 |
-1.410 |
| H11 |
-2.489 |
1.221 |
-0.010 |
| H12 |
-0.144 |
1.903 |
-0.114 |
| H13 |
-0.377 |
0.867 |
1.317 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5286 | 2.5506 | 2.9559 | 4.1572 | 4.8042 | 1.0951 | 1.0966 | 1.0940 | 2.1684 | 2.1674 | 3.5039 | 2.7927 |
C2 | 1.5286 | | 1.5194 | 2.3881 | 3.6235 | 4.4598 | 2.1744 | 2.1783 | 2.1750 | 1.0969 | 1.0979 | 2.1762 | 2.1763 | C3 | 2.5506 | 1.5194 | | 1.4413 | 2.3120 | 3.3396 | 3.4958 | 2.8182 | 2.8259 | 2.1402 | 2.1217 | 1.0957 | 1.0962 | O4 | 2.9559 | 2.3881 | 1.4413 | | 1.3832 | 2.0754 | 3.9370 | 3.3737 | 2.6610 | 2.6113 | 3.3235 | 2.0691 | 2.0724 | N5 | 4.1572 | 3.6235 | 2.3120 | 1.3832 | | 1.1854 | 5.2018 | 4.3329 | 3.8253 | 3.9350 | 4.4194 | 2.6164 | 2.4857 | O6 | 4.8042 | 4.4598 | 3.3396 | 2.0754 | 1.1854 | | 5.7986 | 5.0647 | 4.2257 | 4.6051 | 5.3735 | 3.6717 | 3.5934 | H7 | 1.0951 | 2.1744 | 3.4958 | 3.9370 | 5.2018 | 5.7986 | | 1.7706 | 1.7716 | 2.5080 | 2.5087 | 4.3321 | 3.7991 | H8 | 1.0966 | 2.1783 | 2.8182 | 3.3737 | 4.3329 | 5.0647 | 1.7706 | | 1.7720 | 3.0847 | 2.5256 | 3.8116 | 2.6155 | H9 | 1.0940 | 2.1750 | 2.8259 | 2.6610 | 3.8253 | 4.2257 | 1.7716 | 1.7720 | | 2.5281 | 3.0811 | 3.8393 | 3.1434 | H10 | 2.1684 | 1.0969 | 2.1402 | 2.6113 | 3.9350 | 4.6051 | 2.5080 | 3.0847 | 2.5281 | | 1.7602 | 2.4943 | 3.0674 | H11 | 2.1674 | 1.0979 | 2.1217 | 3.3235 | 4.4194 | 5.3735 | 2.5087 | 2.5256 | 3.0811 | 1.7602 | | 2.4442 | 2.5189 | H12 | 3.5039 | 2.1762 | 1.0957 | 2.0691 | 2.6164 | 3.6717 | 4.3321 | 3.8116 | 3.8393 | 2.4943 | 2.4442 | | 1.7819 | H13 | 2.7927 | 2.1763 | 1.0962 | 2.0724 | 2.4857 | 3.5934 | 3.7991 | 2.6155 | 3.1434 | 3.0674 | 2.5189 | 1.7819 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.605 |
|
C1 |
C2 |
H10 |
110.271 |
| C1 |
C2 |
H11 |
110.133 |
|
C2 |
C1 |
H7 |
110.859 |
| C2 |
C1 |
H8 |
111.080 |
|
C2 |
C1 |
H9 |
110.967 |
| C2 |
C3 |
O4 |
107.500 |
|
C2 |
C3 |
H12 |
111.617 |
| C2 |
C3 |
H13 |
111.594 |
|
C3 |
C2 |
H10 |
108.695 |
| C3 |
C2 |
H11 |
107.219 |
|
C3 |
O4 |
N5 |
109.862 |
| O4 |
C3 |
H12 |
108.519 |
|
O4 |
C3 |
H13 |
108.753 |
| O4 |
N5 |
O6 |
107.552 |
|
H7 |
C1 |
H8 |
107.772 |
| H7 |
C1 |
H9 |
108.052 |
|
H8 |
C1 |
H9 |
107.976 |
| H10 |
C2 |
H11 |
106.639 |
|
H12 |
C3 |
H13 |
108.769 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.518 |
|
|
|
| 2 |
C |
-0.326 |
|
|
|
| 3 |
C |
-0.092 |
|
|
|
| 4 |
O |
-0.348 |
|
|
|
| 5 |
N |
0.285 |
|
|
|
| 6 |
O |
-0.247 |
|
|
|
| 7 |
H |
0.176 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.185 |
|
|
|
| 10 |
H |
0.183 |
|
|
|
| 11 |
H |
0.170 |
|
|
|
| 12 |
H |
0.182 |
|
|
|
| 13 |
H |
0.180 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.436 |
-0.894 |
0.721 |
2.693 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.236 |
0.479 |
-0.611 |
| y |
0.479 |
-34.338 |
-0.788 |
| z |
-0.611 |
-0.788 |
-36.433 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.850 |
0.479 |
-0.611 |
| y |
0.479 |
2.996 |
-0.788 |
| z |
-0.611 |
-0.788 |
-0.145 |
|
| Polar |
|---|
| 3z2-r2 | -0.291 |
|---|
| x2-y2 | -3.898 |
|---|
| xy | 0.479 |
|---|
| xz | -0.611 |
|---|
| yz | -0.788 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.965 |
-1.123 |
0.004 |
| y |
-1.123 |
6.542 |
-0.016 |
| z |
0.004 |
-0.016 |
5.715 |
<r2> (average value of r
2) Å
2
| <r2> |
195.583 |
| (<r2>)1/2 |
13.985 |
