return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7ONO (Propyl nitrite)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-323.304495
Energy at 298.15K-323.313523
HF Energy-322.991517
Nuclear repulsion energy240.480693
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3170 3009 27.60      
2 A 3158 2998 39.32      
3 A 3148 2988 22.69      
4 A 3121 2963 15.39      
5 A 3090 2933 27.96      
6 A 3085 2928 21.46      
7 A 3078 2922 16.70      
8 A 1711 1624 204.81      
9 A 1558 1479 7.24      
10 A 1551 1472 6.64      
11 A 1536 1458 1.74      
12 A 1530 1452 2.22      
13 A 1464 1389 5.76      
14 A 1432 1359 16.83      
15 A 1415 1343 6.98      
16 A 1333 1265 1.43      
17 A 1314 1247 8.86      
18 A 1213 1152 7.46      
19 A 1140 1082 6.05      
20 A 1099 1043 24.67      
21 A 1015 963 14.14      
22 A 951 902 53.85      
23 A 892 847 3.21      
24 A 837 795 304.17      
25 A 791 750 84.99      
26 A 600 570 41.39      
27 A 455 431 11.40      
28 A 382 363 2.94      
29 A 286 271 1.32      
30 A 249 237 1.04      
31 A 186 177 0.40      
32 A 111 106 0.93      
33 A 61 58 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 23480.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 22287.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.25297 0.06754 0.05803

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.530 -2.101 0.104
N2 -1.527 0.160 0.427
O3 -2.501 0.504 -0.156
O4 -0.617 -0.510 -0.456
C5 1.801 -0.356 -0.174
H6 1.966 -0.583 -1.233
H7 2.625 -0.824 0.379
C8 0.510 -1.023 0.286
H9 0.327 -0.848 1.351
H10 2.772 1.587 -0.245
H11 1.645 1.400 1.105
H12 1.029 1.641 -0.535
C13 1.812 1.156 0.051

Atom - Atom Distances (Å)
  H1 N2 O3 O4 C5 H6 H7 C8 H9 H10 H11 H12 C13
H13.07334.00522.03992.17622.48042.46811.09331.77904.32943.80753.82883.5000
N23.07331.18661.43353.42123.93794.26662.35892.30314.57873.47223.10693.5044
O34.00521.18662.16054.38794.72235.32213.40463.47795.38394.42533.72844.3672
O42.03991.43352.16052.43952.69853.36201.44302.06623.99033.34622.70952.9882
C52.17623.42124.38792.43951.09491.09651.52472.17772.17312.17822.17201.5288
H62.48043.93794.72232.69851.09491.75742.14963.07132.51663.08212.51262.1667
H72.46814.26665.32213.36201.09651.75742.12652.49512.49452.53623.07562.1651
C81.09332.35893.40461.44301.52472.14962.12651.09483.49422.79802.83572.5491
H91.77902.30313.47792.06622.17773.07132.49511.09483.80162.61743.20112.8128
H104.32944.57875.38393.99032.17312.51662.49453.49423.80161.76811.76771.0928
H113.80753.47224.42533.34622.17823.08212.53622.79802.61741.76811.76841.0948
H123.82883.10693.72842.70952.17202.51263.07562.83573.20111.76771.76841.0918
C133.50003.50444.36722.98821.52882.16672.16512.54912.81281.09281.09481.0918

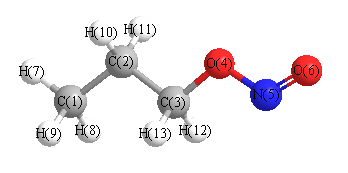
picture of Propyl nitrite state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 135.351 C1 C2 H10 65.510
C1 C2 H11 70.841 C2 C1 H7 100.127
C2 C1 H8 40.947 C2 C1 H9 48.080
C2 C3 O4 38.353 C2 C3 H12 50.262
C2 C3 H13 37.706 C3 C2 H10 127.491
C3 C2 H11 137.950 C3 O4 N5 145.002
O4 C3 H12 45.822 O4 C3 H13 38.293
O4 N5 O6 91.415 H7 C1 H8 59.180
H7 C1 H9 69.810 H8 C1 H9 35.632
H10 C2 H11 19.916 H12 C3 H13 12.597
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.170      
2 N 0.341      
3 O -0.259      
4 O -0.426      
5 C -0.290      
6 H 0.167      
7 H 0.152      
8 C -0.032      
9 H 0.167      
10 H 0.161      
11 H 0.153      
12 H 0.170      
13 C -0.472      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.971 -1.203 -0.019
y -1.203 6.495 0.004
z -0.019 0.004 5.665


<r2> (average value of r2) Å2
<r2> 195.479
(<r2>)1/2 13.981