Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -93.678689 |
| Energy at 298.15K | -93.680001 |
| HF Energy | -93.391940 |
| Nuclear repulsion energy | 27.952423 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3321 |
3163 |
3.96 |
|
|
|
| 2 |
A' |
3038 |
2893 |
58.32 |
|
|
|
| 3 |
A' |
1841 |
1753 |
22.12 |
|
|
|
| 4 |
A' |
1076 |
1025 |
135.85 |
|
|
|
| 5 |
A' |
908 |
864 |
110.48 |
|
|
|
| 6 |
A" |
928 |
884 |
7.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5555.7 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 5290.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.113 |
0.644 |
0.000 |
| N2 |
0.113 |
-0.596 |
0.000 |
| H3 |
-0.677 |
1.410 |
0.000 |
| H4 |
-0.787 |
-1.101 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2396 | 1.1006 | 1.9625 |
N2 | 1.2396 | | 2.1561 | 1.0314 | H3 | 1.1006 | 2.1561 | | 2.5133 | H4 | 1.9625 | 1.0314 | 2.5133 | |
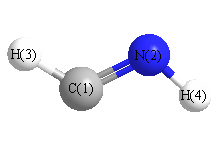 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
119.289 |
|
H3 |
C1 |
N2 |
134.156 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -93.687053 |
| Energy at 298.15K | -93.688410 |
| HF Energy | -93.397625 |
| Nuclear repulsion energy | 27.889604 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3448 |
3283 |
1.84 |
|
|
|
| 2 |
A' |
3091 |
2944 |
33.55 |
|
|
|
| 3 |
A' |
1798 |
1712 |
25.54 |
|
|
|
| 4 |
A' |
1253 |
1193 |
14.60 |
|
|
|
| 5 |
A' |
952 |
907 |
212.18 |
|
|
|
| 6 |
A" |
1009 |
961 |
107.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5775.5 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 5500.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.656 |
0.000 |
| N2 |
0.001 |
-0.594 |
0.000 |
| H3 |
0.909 |
1.273 |
0.000 |
| H4 |
-0.917 |
-1.052 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2496 | 1.0980 | 1.9389 |
N2 | 1.2496 | | 2.0759 | 1.0258 | H3 | 1.0980 | 2.0759 | | 2.9563 | H4 | 1.9389 | 1.0258 | 2.9563 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
116.543 |
|
H3 |
C1 |
N2 |
124.193 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
