return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5N (Cyclopropanecarbonitrile)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-209.377281
Energy at 298.15K-209.382711
HF Energy-208.794161
Nuclear repulsion energy148.266797
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3374 3117 5.65      
2 A' 3312 3059 0.62      
3 A' 3282 3032 3.06      
4 A' 2491 2301 23.03      
5 A' 1611 1488 0.45      
6 A' 1475 1362 0.61      
7 A' 1294 1195 1.52      
8 A' 1206 1114 1.09      
9 A' 1160 1072 4.34      
10 A' 1026 948 25.83      
11 A' 858 792 2.35      
12 A' 800 739 5.03      
13 A' 557 514 0.43      
14 A' 223 206 4.73      
15 A" 3362 3105 0.25      
16 A" 3275 3025 8.25      
17 A" 1561 1441 3.02      
18 A" 1273 1176 0.72      
19 A" 1194 1103 1.08      
20 A" 1173 1084 7.27      
21 A" 997 921 1.46      
22 A" 886 818 9.65      
23 A" 577 533 1.34      
24 A" 226 209 3.20      

Unscaled Zero Point Vibrational Energy (zpe) 18595.7 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 17176.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.53626 0.11554 0.10955

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -1.770 1.031 1.265
H2 -1.770 1.031 -1.265
H3 -1.253 -0.712 -1.255
H4 -1.253 -0.712 1.255
H5 0.283 1.664 0.000
C6 0.000 0.622 0.000
N7 2.009 -1.024 0.000
C8 1.121 -0.289 0.000
C9 -1.253 0.239 0.745
C10 -1.253 0.239 -0.745

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 C6 N7 C8 C9 C10
H12.52923.10671.81782.49232.21314.48333.42051.07932.2207
H22.52921.81783.10672.49232.21314.48333.42052.22071.0793
H33.10671.81782.50973.09472.21893.50912.71852.21401.0791
H41.81783.10672.50973.09472.21893.50912.71851.07912.2140
H52.49232.49233.09473.09471.07973.19442.12582.22302.2230
C62.21312.21312.21892.21891.07972.59711.44491.50661.5066
N74.48334.48333.50913.50913.19442.59711.15223.57593.5759
C83.42053.42052.71852.71852.12581.44491.15222.54332.5433
C91.07932.22072.21401.07912.22301.50663.57592.54331.4892
C102.22071.07931.07912.21402.22301.50663.57592.54331.4892

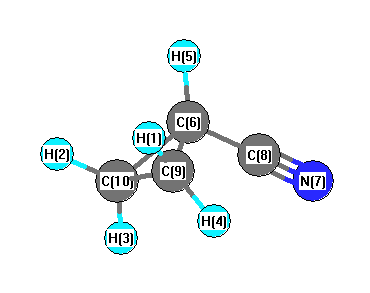
picture of Cyclopropanecarbonitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C9 H4 114.743 H1 C9 C6 116.733
H1 C9 C10 118.799 H2 C10 H3 114.743
H2 C10 C6 116.733 H2 C10 C9 118.799
H3 C10 C6 117.262 H3 C10 C9 118.220
H4 C9 C6 117.262 H4 C9 C10 118.220
H5 C6 C8 113.931 H5 C6 C9 117.579
H5 C6 C10 117.579 C6 C8 N7 179.521
C6 C9 C10 60.381 C6 C10 C9 60.381
C8 C6 C9 118.995 C8 C6 C10 118.995
C9 C6 C10 59.237
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability