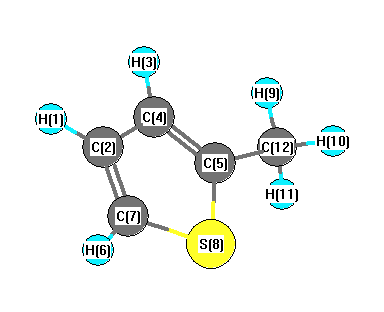
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3282 |
3140 |
1.08 |
|
|
|
| 2 |
A |
3239 |
3099 |
7.63 |
|
|
|
| 3 |
A |
3222 |
3082 |
8.50 |
|
|
|
| 4 |
A |
3145 |
3009 |
11.68 |
|
|
|
| 5 |
A |
3053 |
2921 |
31.53 |
|
|
|
| 6 |
A |
1618 |
1548 |
2.97 |
|
|
|
| 7 |
A |
1526 |
1460 |
2.91 |
|
|
|
| 8 |
A |
1514 |
1448 |
12.78 |
|
|
|
| 9 |
A |
1438 |
1376 |
0.16 |
|
|
|
| 10 |
A |
1402 |
1341 |
0.68 |
|
|
|
| 11 |
A |
1271 |
1216 |
11.20 |
|
|
|
| 12 |
A |
1197 |
1145 |
12.18 |
|
|
|
| 13 |
A |
1115 |
1066 |
2.08 |
|
|
|
| 14 |
A |
1078 |
1032 |
3.89 |
|
|
|
| 15 |
A |
1003 |
960 |
1.41 |
|
|
|
| 16 |
A |
866 |
829 |
12.90 |
|
|
|
| 17 |
A |
754 |
721 |
0.22 |
|
|
|
| 18 |
A |
676 |
647 |
2.17 |
|
|
|
| 19 |
A |
555 |
531 |
0.71 |
|
|
|
| 20 |
A |
304 |
291 |
0.86 |
|
|
|
| 21 |
A |
3114 |
2980 |
14.59 |
|
|
|
| 22 |
A |
1504 |
1439 |
8.87 |
|
|
|
| 23 |
A |
1067 |
1021 |
1.32 |
|
|
|
| 24 |
A |
905 |
866 |
0.42 |
|
|
|
| 25 |
A |
829 |
793 |
10.97 |
|
|
|
| 26 |
A |
703 |
672 |
65.10 |
|
|
|
| 27 |
A |
574 |
549 |
0.46 |
|
|
|
| 28 |
A |
475 |
454 |
4.91 |
|
|
|
| 29 |
A |
232 |
222 |
2.79 |
|
|
|
| 30 |
A |
103 |
98 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20881.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 19977.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.170 |
|
|
0.165 |
| 2 |
C |
-0.128 |
|
|
-0.173 |
| 3 |
H |
0.164 |
|
|
0.162 |
| 4 |
C |
-0.149 |
|
|
-0.209 |
| 5 |
C |
-0.144 |
|
|
0.107 |
| 6 |
H |
0.200 |
|
|
0.224 |
| 7 |
C |
-0.397 |
|
|
-0.275 |
| 8 |
S |
0.271 |
|
|
-0.011 |
| 9 |
H |
0.190 |
|
|
0.107 |
| 10 |
H |
0.193 |
|
|
0.127 |
| 11 |
H |
0.193 |
|
|
0.127 |
| 12 |
C |
-0.564 |
|
|
-0.350 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.504 |
0.527 |
0.000 |
0.729 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.506 |
0.522 |
0.000 |
0.727 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.570 |
0.750 |
0.000 |
| y |
0.750 |
-36.556 |
0.000 |
| z |
0.000 |
0.000 |
-45.630 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.522 |
0.750 |
0.000 |
| y |
0.750 |
6.545 |
0.000 |
| z |
0.000 |
0.000 |
-7.067 |
|
| Polar |
|---|
| 3z2-r2 | -14.133 |
|---|
| x2-y2 | -4.015 |
|---|
| xy | 0.750 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
177.072 |
| (<r2>)1/2 |
13.307 |