return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6S (Thiophene, 2-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-591.933278
Energy at 298.15K-591.939413
HF Energy-591.675867
Nuclear repulsion energy272.062048
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3296 3129 1.33      
2 A' 3255 3090 7.28      
3 A' 3238 3073 8.29      
4 A' 3163 3002 12.70      
5 A' 3077 2921 29.98      
6 A' 1613 1531 2.23      
7 A' 1545 1466 3.70      
8 A' 1515 1438 10.15      
9 A' 1462 1388 0.62      
10 A' 1410 1339 0.57      
11 A' 1286 1220 11.46      
12 A' 1204 1143 12.16      
13 A' 1125 1068 1.52      
14 A' 1086 1031 2.96      
15 A' 1014 962 1.55      
16 A' 869 824 12.16      
17 A' 754 716 0.39      
18 A' 673 639 2.44      
19 A' 557 529 0.83      
20 A' 305 290 0.76      
21 A" 3138 2979 15.95      
22 A" 1529 1451 7.52      
23 A" 1084 1029 0.61      
24 A" 885 840 0.28      
25 A" 823 781 11.83      
26 A" 700 664 60.27      
27 A" 565 536 0.89      
28 A" 470 447 4.23      
29 A" 230 218 2.51      
30 A" 100 95 0.15      

Unscaled Zero Point Vibrational Energy (zpe) 20984.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 19918.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.17497 0.10330 0.06575

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.399 -1.544 0.000
C2 -1.443 -1.033 0.000
H3 -2.145 1.078 0.000
C4 -1.303 0.394 0.000
C5 0.000 0.813 0.000
H6 -0.046 -2.740 0.000
C7 -0.238 -1.676 0.000
S8 1.087 -0.554 0.000
H9 -0.313 2.920 0.000
H10 1.137 2.424 0.884
H11 1.137 2.424 -0.884
C12 0.525 2.219 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 C5 H6 C7 S8 H9 H10 H11 C12
H11.08402.63442.22583.36292.63992.16513.62354.92775.38835.38834.7657
C21.08402.22481.43342.34282.20581.36562.57454.11184.40364.40363.8015
H32.63442.22481.08482.16114.35693.34953.62022.59793.65573.65572.9034
C42.22581.43341.08481.36903.37632.32732.57092.71403.29563.29562.5838
C53.36292.34282.16111.36903.55262.49971.74602.13092.16142.16141.5015
H62.63992.20584.35693.37633.55261.08132.46145.66655.37105.37104.9918
C72.16511.36563.34952.32732.49971.08131.73564.59674.41394.41393.9691
S83.62352.57453.62022.57091.74602.46141.73563.74583.10723.10722.8298
H94.92774.11182.59792.71402.13095.66654.59673.74581.76921.76921.0925
H105.38834.40363.65573.29562.16145.37104.41393.10721.76921.76731.0944
H115.38834.40363.65573.29562.16145.37104.41393.10721.76921.76731.0944
C124.76573.80152.90342.58381.50154.99183.96912.82981.09251.09441.0944

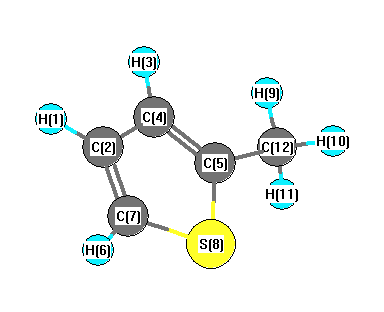
picture of Thiophene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 123.706 H1 C2 C7 123.815
C2 C4 H3 123.542 C2 C4 C5 113.418
C2 C7 H6 128.322 C2 C7 S8 111.675
H3 C4 C5 123.039 C4 C2 C7 112.479
C4 C5 S8 110.659 C4 C5 C12 128.291
C5 S8 C7 91.769 C5 C12 H9 109.453
C5 C12 H10 111.782 C5 C12 H11 111.782
H6 C7 S8 120.004 S8 C5 C12 121.050
H9 C12 H10 107.992 H9 C12 H11 107.992
H10 C12 H11 107.692
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.168      
2 C -0.126      
3 H 0.162      
4 C -0.152      
5 C -0.146      
6 H 0.195      
7 C -0.381      
8 S 0.256      
9 H 0.174      
10 H 0.178      
11 H 0.178      
12 C -0.505      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 11.009 0.387 0.000
y 0.387 11.431 0.000
z 0.000 0.000 4.905


<r2> (average value of r2) Å2
<r2> 177.830
(<r2>)1/2 13.335