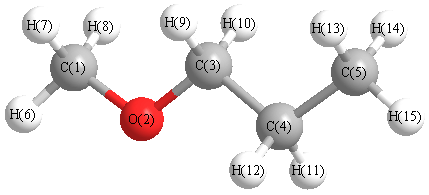
Vibrational Frequencies calculated at B2PLYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3175 |
3013 |
34.08 |
|
|
|
| 2 |
A' |
3158 |
2998 |
33.30 |
|
|
|
| 3 |
A' |
3090 |
2933 |
26.27 |
|
|
|
| 4 |
A' |
3078 |
2922 |
24.30 |
|
|
|
| 5 |
A' |
3024 |
2871 |
71.34 |
|
|
|
| 6 |
A' |
2993 |
2841 |
40.27 |
|
|
|
| 7 |
A' |
1581 |
1501 |
0.93 |
|
|
|
| 8 |
A' |
1560 |
1480 |
7.53 |
|
|
|
| 9 |
A' |
1555 |
1476 |
5.11 |
|
|
|
| 10 |
A' |
1544 |
1465 |
1.58 |
|
|
|
| 11 |
A' |
1522 |
1444 |
1.54 |
|
|
|
| 12 |
A' |
1465 |
1391 |
28.02 |
|
|
|
| 13 |
A' |
1462 |
1387 |
1.20 |
|
|
|
| 14 |
A' |
1367 |
1297 |
5.75 |
|
|
|
| 15 |
A' |
1256 |
1192 |
38.45 |
|
|
|
| 16 |
A' |
1187 |
1126 |
127.50 |
|
|
|
| 17 |
A' |
1152 |
1093 |
22.47 |
|
|
|
| 18 |
A' |
1078 |
1024 |
2.28 |
|
|
|
| 19 |
A' |
997 |
946 |
23.18 |
|
|
|
| 20 |
A' |
928 |
881 |
6.26 |
|
|
|
| 21 |
A' |
444 |
421 |
0.63 |
|
|
|
| 22 |
A' |
414 |
393 |
3.04 |
|
|
|
| 23 |
A' |
196 |
186 |
1.50 |
|
|
|
| 24 |
A" |
3152 |
2992 |
63.73 |
|
|
|
| 25 |
A" |
3127 |
2968 |
3.20 |
|
|
|
| 26 |
A" |
3073 |
2916 |
67.37 |
|
|
|
| 27 |
A" |
3024 |
2870 |
65.49 |
|
|
|
| 28 |
A" |
1553 |
1474 |
6.45 |
|
|
|
| 29 |
A" |
1537 |
1459 |
4.68 |
|
|
|
| 30 |
A" |
1346 |
1277 |
0.06 |
|
|
|
| 31 |
A" |
1299 |
1233 |
1.49 |
|
|
|
| 32 |
A" |
1230 |
1167 |
5.60 |
|
|
|
| 33 |
A" |
1196 |
1135 |
0.33 |
|
|
|
| 34 |
A" |
924 |
877 |
1.88 |
|
|
|
| 35 |
A" |
783 |
743 |
1.32 |
|
|
|
| 36 |
A" |
248 |
235 |
1.54 |
|
|
|
| 37 |
A" |
242 |
229 |
2.33 |
|
|
|
| 38 |
A" |
117 |
112 |
2.28 |
|
|
|
| 39 |
A" |
106 |
100 |
1.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30588.3 cm
-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29034.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.194 |
|
|
|
| 2 |
O |
-0.487 |
|
|
|
| 3 |
C |
-0.004 |
|
|
|
| 4 |
C |
-0.283 |
|
|
|
| 5 |
C |
-0.468 |
|
|
|
| 6 |
H |
0.164 |
|
|
|
| 7 |
H |
0.133 |
|
|
|
| 8 |
H |
0.133 |
|
|
|
| 9 |
H |
0.120 |
|
|
|
| 10 |
H |
0.120 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.155 |
|
|
|
| 13 |
H |
0.150 |
|
|
|
| 14 |
H |
0.150 |
|
|
|
| 15 |
H |
0.157 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.213 |
1.125 |
0.000 |
1.145 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.003 |
-2.219 |
0.000 |
| y |
-2.219 |
-33.409 |
0.000 |
| z |
0.000 |
0.000 |
-33.089 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.246 |
-2.219 |
0.000 |
| y |
-2.219 |
-1.363 |
0.000 |
| z |
0.000 |
0.000 |
-0.883 |
|
| Polar |
|---|
| 3z2-r2 | -1.766 |
|---|
| x2-y2 | 2.406 |
|---|
| xy | -2.219 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.303 |
-0.352 |
0.000 |
| y |
-0.352 |
6.440 |
0.000 |
| z |
0.000 |
0.000 |
6.296 |
<r2> (average value of r
2) Å
2
| <r2> |
181.086 |
| (<r2>)1/2 |
13.457 |