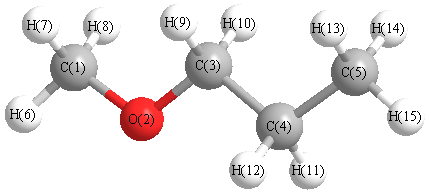
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3159 |
3004 |
33.14 |
|
|
|
| 2 |
A' |
3153 |
2999 |
30.18 |
|
|
|
| 3 |
A' |
3078 |
2928 |
27.85 |
|
|
|
| 4 |
A' |
3067 |
2916 |
23.88 |
|
|
|
| 5 |
A' |
3004 |
2857 |
75.69 |
|
|
|
| 6 |
A' |
2972 |
2827 |
42.25 |
|
|
|
| 7 |
A' |
1560 |
1483 |
1.50 |
|
|
|
| 8 |
A' |
1537 |
1461 |
11.28 |
|
|
|
| 9 |
A' |
1532 |
1457 |
4.74 |
|
|
|
| 10 |
A' |
1519 |
1445 |
1.64 |
|
|
|
| 11 |
A' |
1505 |
1432 |
0.55 |
|
|
|
| 12 |
A' |
1449 |
1378 |
25.19 |
|
|
|
| 13 |
A' |
1440 |
1369 |
3.35 |
|
|
|
| 14 |
A' |
1348 |
1282 |
3.74 |
|
|
|
| 15 |
A' |
1255 |
1193 |
74.63 |
|
|
|
| 16 |
A' |
1210 |
1151 |
115.50 |
|
|
|
| 17 |
A' |
1147 |
1091 |
6.09 |
|
|
|
| 18 |
A' |
1083 |
1030 |
3.47 |
|
|
|
| 19 |
A' |
1020 |
970 |
23.72 |
|
|
|
| 20 |
A' |
924 |
878 |
6.86 |
|
|
|
| 21 |
A' |
445 |
423 |
0.64 |
|
|
|
| 22 |
A' |
412 |
392 |
2.87 |
|
|
|
| 23 |
A' |
195 |
185 |
1.43 |
|
|
|
| 24 |
A" |
3144 |
2990 |
57.68 |
|
|
|
| 25 |
A" |
3118 |
2965 |
3.68 |
|
|
|
| 26 |
A" |
3052 |
2903 |
66.00 |
|
|
|
| 27 |
A" |
3002 |
2855 |
70.25 |
|
|
|
| 28 |
A" |
1530 |
1455 |
8.02 |
|
|
|
| 29 |
A" |
1515 |
1441 |
6.28 |
|
|
|
| 30 |
A" |
1330 |
1265 |
0.13 |
|
|
|
| 31 |
A" |
1285 |
1222 |
2.11 |
|
|
|
| 32 |
A" |
1216 |
1157 |
4.88 |
|
|
|
| 33 |
A" |
1187 |
1128 |
0.07 |
|
|
|
| 34 |
A" |
911 |
866 |
2.13 |
|
|
|
| 35 |
A" |
772 |
735 |
2.02 |
|
|
|
| 36 |
A" |
249 |
237 |
2.19 |
|
|
|
| 37 |
A" |
244 |
232 |
1.49 |
|
|
|
| 38 |
A" |
116 |
110 |
2.58 |
|
|
|
| 39 |
A" |
107 |
102 |
0.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30396.4 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 28907.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.256 |
|
|
|
| 2 |
O |
-0.462 |
|
|
|
| 3 |
C |
-0.045 |
|
|
|
| 4 |
C |
-0.330 |
|
|
|
| 5 |
C |
-0.525 |
|
|
|
| 6 |
H |
0.181 |
|
|
|
| 7 |
H |
0.148 |
|
|
|
| 8 |
H |
0.148 |
|
|
|
| 9 |
H |
0.137 |
|
|
|
| 10 |
H |
0.137 |
|
|
|
| 11 |
H |
0.176 |
|
|
|
| 12 |
H |
0.176 |
|
|
|
| 13 |
H |
0.169 |
|
|
|
| 14 |
H |
0.169 |
|
|
|
| 15 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.235 |
1.061 |
0.000 |
1.087 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.745 |
-2.173 |
0.000 |
| y |
-2.173 |
-33.048 |
0.000 |
| z |
0.000 |
0.000 |
-32.792 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.175 |
-2.173 |
0.000 |
| y |
-2.173 |
-1.280 |
0.000 |
| z |
0.000 |
0.000 |
-0.895 |
|
| Polar |
|---|
| 3z2-r2 | -1.790 |
|---|
| x2-y2 | 2.303 |
|---|
| xy | -2.173 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.381 |
-0.386 |
0.000 |
| y |
-0.386 |
6.527 |
0.000 |
| z |
0.000 |
0.000 |
6.374 |
<r2> (average value of r
2) Å
2
| <r2> |
179.763 |
| (<r2>)1/2 |
13.408 |