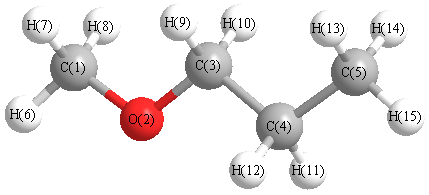
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3065 |
3015 |
32.71 |
|
|
|
| 2 |
A' |
3064 |
3013 |
29.78 |
|
|
|
| 3 |
A' |
2988 |
2939 |
34.13 |
|
|
|
| 4 |
A' |
2981 |
2932 |
18.88 |
|
|
|
| 5 |
A' |
2903 |
2856 |
84.34 |
|
|
|
| 6 |
A' |
2871 |
2824 |
44.86 |
|
|
|
| 7 |
A' |
1512 |
1487 |
1.15 |
|
|
|
| 8 |
A' |
1491 |
1467 |
8.07 |
|
|
|
| 9 |
A' |
1484 |
1460 |
5.85 |
|
|
|
| 10 |
A' |
1474 |
1450 |
2.76 |
|
|
|
| 11 |
A' |
1453 |
1429 |
0.98 |
|
|
|
| 12 |
A' |
1396 |
1373 |
23.85 |
|
|
|
| 13 |
A' |
1390 |
1367 |
2.20 |
|
|
|
| 14 |
A' |
1303 |
1282 |
4.56 |
|
|
|
| 15 |
A' |
1200 |
1180 |
31.74 |
|
|
|
| 16 |
A' |
1145 |
1126 |
141.34 |
|
|
|
| 17 |
A' |
1107 |
1089 |
10.69 |
|
|
|
| 18 |
A' |
1046 |
1029 |
2.19 |
|
|
|
| 19 |
A' |
970 |
954 |
22.27 |
|
|
|
| 20 |
A' |
894 |
879 |
7.35 |
|
|
|
| 21 |
A' |
431 |
423 |
0.58 |
|
|
|
| 22 |
A' |
398 |
391 |
2.66 |
|
|
|
| 23 |
A' |
187 |
184 |
1.27 |
|
|
|
| 24 |
A" |
3054 |
3003 |
55.04 |
|
|
|
| 25 |
A" |
3025 |
2975 |
5.95 |
|
|
|
| 26 |
A" |
2943 |
2894 |
68.44 |
|
|
|
| 27 |
A" |
2896 |
2848 |
76.89 |
|
|
|
| 28 |
A" |
1483 |
1459 |
7.55 |
|
|
|
| 29 |
A" |
1462 |
1438 |
5.61 |
|
|
|
| 30 |
A" |
1287 |
1266 |
0.05 |
|
|
|
| 31 |
A" |
1240 |
1220 |
1.59 |
|
|
|
| 32 |
A" |
1174 |
1155 |
3.95 |
|
|
|
| 33 |
A" |
1143 |
1124 |
0.06 |
|
|
|
| 34 |
A" |
885 |
871 |
2.07 |
|
|
|
| 35 |
A" |
754 |
742 |
2.40 |
|
|
|
| 36 |
A" |
246 |
242 |
2.94 |
|
|
|
| 37 |
A" |
234 |
230 |
0.05 |
|
|
|
| 38 |
A" |
114 |
113 |
2.33 |
|
|
|
| 39 |
A" |
105 |
103 |
0.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29399.6 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 28914.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.248 |
|
|
|
| 2 |
O |
-0.413 |
|
|
|
| 3 |
C |
-0.046 |
|
|
|
| 4 |
C |
-0.299 |
|
|
|
| 5 |
C |
-0.493 |
|
|
|
| 6 |
H |
0.168 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
H |
0.138 |
|
|
|
| 9 |
H |
0.124 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
| 12 |
H |
0.162 |
|
|
|
| 13 |
H |
0.158 |
|
|
|
| 14 |
H |
0.158 |
|
|
|
| 15 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.246 |
0.970 |
0.000 |
1.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.863 |
-2.041 |
0.000 |
| y |
-2.041 |
-33.194 |
0.000 |
| z |
0.000 |
0.000 |
-32.963 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.215 |
-2.041 |
0.000 |
| y |
-2.041 |
-1.281 |
0.000 |
| z |
0.000 |
0.000 |
-0.934 |
|
| Polar |
|---|
| 3z2-r2 | -1.868 |
|---|
| x2-y2 | 2.331 |
|---|
| xy | -2.041 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.873 |
-0.455 |
0.000 |
| y |
-0.455 |
6.751 |
0.000 |
| z |
0.000 |
0.000 |
6.538 |
<r2> (average value of r
2) Å
2
| <r2> |
182.256 |
| (<r2>)1/2 |
13.500 |