return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (1-Butene, 2-methyl-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-195.731204
Energy at 298.15K-195.741511
HF Energy-195.143499
Nuclear repulsion energy176.289986
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3348 3093 19.79      
2 A 3269 3020 9.04      
3 A 3247 3000 25.66      
4 A 3243 2995 32.45      
5 A 3237 2990 39.44      
6 A 3213 2968 26.56      
7 A 3198 2954 13.14      
8 A 3162 2921 25.05      
9 A 3151 2911 22.73      
10 A 3143 2903 27.18      
11 A 1834 1694 11.46      
12 A 1604 1482 2.52      
13 A 1598 1476 6.35      
14 A 1592 1471 9.83      
15 A 1585 1464 7.45      
16 A 1573 1453 4.43      
17 A 1540 1422 1.15      
18 A 1512 1396 3.64      
19 A 1506 1391 3.72      
20 A 1449 1338 1.55      
21 A 1388 1282 1.53      
22 A 1349 1246 1.35      
23 A 1186 1095 0.33      
24 A 1163 1074 5.25      
25 A 1116 1031 0.77      
26 A 1084 1001 1.09      
27 A 1058 978 0.69      
28 A 997 921 0.71      
29 A 980 905 46.87      
30 A 841 777 1.38      
31 A 816 754 1.07      
32 A 761 703 0.13      
33 A 561 518 5.58      
34 A 452 417 1.59      
35 A 414 383 0.62      
36 A 277 256 0.17      
37 A 233 215 0.34      
38 A 183 169 0.21      
39 A 64 59 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 31463.3 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 29062.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.24299 0.12572 0.09543

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.978 1.330 0.028
H2 0.381 1.872 -0.706
H3 2.026 1.443 -0.243
H4 0.820 1.819 0.991
C5 -1.901 0.085 -0.393
H6 -1.883 1.169 -0.487
H7 -2.890 -0.198 -0.036
H8 -1.765 -0.341 -1.386
C9 -0.819 -0.419 0.563
H10 -0.929 -1.496 0.695
H11 -0.972 0.034 1.546
C12 0.584 -0.120 0.097
C13 1.429 -1.090 -0.233
H14 1.147 -2.133 -0.180
H15 2.435 -0.882 -0.568

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 C13 H14 H15
C11.09041.08781.09153.16532.91204.16033.51002.56483.47422.79121.50492.47613.47332.7149
H21.09041.76161.75402.91552.38103.92923.15702.88123.87583.20682.15763.17734.11063.4379
H31.08781.76161.76604.15763.92615.18704.34263.49424.27153.76512.15342.60253.68232.3824
H41.09151.75401.76603.51133.14894.34684.12352.80773.76002.59052.14883.21494.13483.5121
C53.16532.91554.15763.51131.08871.08941.08941.52932.15122.15042.54133.53463.77494.4461
H62.91202.38103.92613.14891.08871.75731.76212.18173.06792.50052.84484.01744.49184.7814
H74.16033.92925.18704.34681.08941.75731.76372.16782.46312.49703.47824.41494.47905.3958
H83.51003.15704.34264.12351.08941.76211.76372.16782.52253.06032.78693.47693.62474.3129
C92.56482.88123.49422.80771.52932.18172.16782.16781.09001.09341.50832.47692.71093.4758
H103.47423.87584.27153.76002.15123.06792.46312.52251.09001.75062.13062.56612.34083.6456
H112.79123.20683.76512.59052.15042.50052.49703.06031.09341.75062.13213.19283.48734.1132
C121.50492.15762.15342.14882.54132.84483.47822.78691.50832.13062.13211.32792.10772.1091
C132.47613.17732.60253.21493.53464.01744.41493.47692.47692.56613.19281.32791.08111.0811
H143.47334.11063.68234.13483.77494.49184.47903.62472.71092.34083.48732.10771.08111.8374
H152.71493.43792.38243.51214.44614.78145.39584.31293.47583.64564.11322.10911.08111.8374

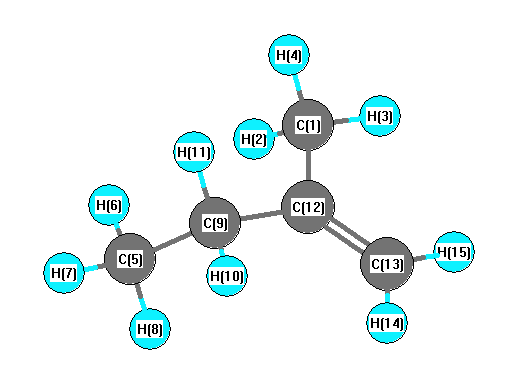
picture of 1-Butene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C12 C9 116.683 C1 C12 C13 121.751
H2 C1 H3 107.943 H2 C1 H4 107.003
H2 C1 C12 111.479 H3 C1 H4 108.252
H3 C1 C12 111.293 H4 C1 C12 110.699
C5 C9 H10 109.278 C5 C9 H11 109.016
C5 C9 C12 113.567 H6 C5 H7 107.568
H6 C5 H8 108.000 H6 C5 C9 111.778
H7 C5 H8 108.100 H7 C5 C9 110.629
H8 C5 C9 110.623 C9 C12 C13 121.565
H10 C9 H11 106.609 H10 C9 C12 109.110
H11 C9 C12 109.027 C12 C13 H14 121.739
C12 C13 H15 121.880 H14 C13 H15 116.382
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability