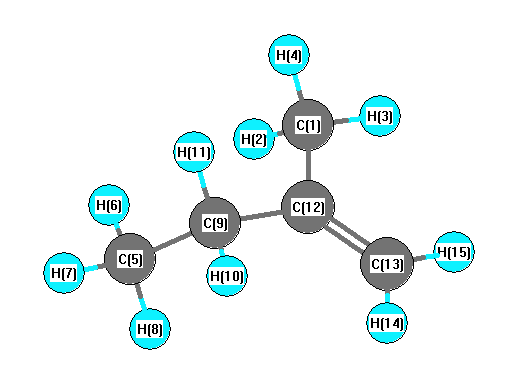
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3243 |
3102 |
20.92 |
|
|
|
| 2 |
A |
3163 |
3026 |
10.54 |
|
|
|
| 3 |
A |
3143 |
3007 |
25.96 |
|
|
|
| 4 |
A |
3140 |
3004 |
30.03 |
|
|
|
| 5 |
A |
3135 |
3000 |
39.32 |
|
|
|
| 6 |
A |
3099 |
2965 |
24.79 |
|
|
|
| 7 |
A |
3091 |
2957 |
14.58 |
|
|
|
| 8 |
A |
3060 |
2928 |
28.32 |
|
|
|
| 9 |
A |
3043 |
2911 |
26.90 |
|
|
|
| 10 |
A |
3032 |
2901 |
29.70 |
|
|
|
| 11 |
A |
1743 |
1668 |
14.54 |
|
|
|
| 12 |
A |
1534 |
1468 |
2.89 |
|
|
|
| 13 |
A |
1528 |
1462 |
6.26 |
|
|
|
| 14 |
A |
1521 |
1455 |
11.27 |
|
|
|
| 15 |
A |
1511 |
1446 |
10.71 |
|
|
|
| 16 |
A |
1499 |
1434 |
2.90 |
|
|
|
| 17 |
A |
1468 |
1404 |
0.37 |
|
|
|
| 18 |
A |
1434 |
1372 |
4.59 |
|
|
|
| 19 |
A |
1429 |
1367 |
3.30 |
|
|
|
| 20 |
A |
1369 |
1310 |
1.71 |
|
|
|
| 21 |
A |
1324 |
1267 |
2.08 |
|
|
|
| 22 |
A |
1285 |
1230 |
1.80 |
|
|
|
| 23 |
A |
1127 |
1079 |
0.40 |
|
|
|
| 24 |
A |
1106 |
1058 |
6.35 |
|
|
|
| 25 |
A |
1063 |
1017 |
1.35 |
|
|
|
| 26 |
A |
1036 |
992 |
1.11 |
|
|
|
| 27 |
A |
1009 |
965 |
1.16 |
|
|
|
| 28 |
A |
955 |
914 |
1.29 |
|
|
|
| 29 |
A |
920 |
881 |
45.70 |
|
|
|
| 30 |
A |
806 |
771 |
1.93 |
|
|
|
| 31 |
A |
784 |
751 |
1.49 |
|
|
|
| 32 |
A |
727 |
696 |
0.14 |
|
|
|
| 33 |
A |
536 |
513 |
5.91 |
|
|
|
| 34 |
A |
431 |
412 |
1.64 |
|
|
|
| 35 |
A |
397 |
380 |
0.67 |
|
|
|
| 36 |
A |
263 |
251 |
0.14 |
|
|
|
| 37 |
A |
224 |
215 |
0.45 |
|
|
|
| 38 |
A |
183 |
175 |
0.30 |
|
|
|
| 39 |
A |
68 |
65 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30215.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28907.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.574 |
|
|
-0.356 |
| 2 |
H |
0.179 |
|
|
0.107 |
| 3 |
H |
0.177 |
|
|
0.108 |
| 4 |
H |
0.174 |
|
|
0.101 |
| 5 |
C |
-0.510 |
|
|
-0.143 |
| 6 |
H |
0.168 |
|
|
0.041 |
| 7 |
H |
0.169 |
|
|
0.036 |
| 8 |
H |
0.175 |
|
|
0.049 |
| 9 |
C |
-0.372 |
|
|
0.005 |
| 10 |
H |
0.167 |
|
|
0.025 |
| 11 |
H |
0.164 |
|
|
0.024 |
| 12 |
C |
0.219 |
|
|
0.205 |
| 13 |
C |
-0.444 |
|
|
-0.606 |
| 14 |
H |
0.154 |
|
|
0.195 |
| 15 |
H |
0.154 |
|
|
0.210 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.345 |
0.360 |
0.101 |
0.508 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.317 |
0.373 |
0.096 |
0.499 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.438 |
0.548 |
-0.819 |
| y |
0.548 |
-31.897 |
0.148 |
| z |
-0.819 |
0.148 |
-33.911 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.466 |
0.548 |
-0.819 |
| y |
0.548 |
1.278 |
0.148 |
| z |
-0.819 |
0.148 |
-1.744 |
|
| Polar |
|---|
| 3z2-r2 | -3.487 |
|---|
| x2-y2 | -0.541 |
|---|
| xy | 0.548 |
|---|
| xz | -0.819 |
|---|
| yz | 0.148 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
143.388 |
| (<r2>)1/2 |
11.974 |