return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-235.032315
Energy at 298.15K-235.044632
HF Energy-234.175198
Nuclear repulsion energy243.147358
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3244 3089 21.60      
2 A' 3167 3016 11.35      
3 A' 3148 2998 28.50      
4 A' 3141 2991 27.65      
5 A' 3135 2986 73.42      
6 A' 3065 2919 21.93      
7 A' 3060 2914 9.81      
8 A' 3050 2905 19.65      
9 A' 1748 1664 8.63      
10 A' 1560 1486 5.04      
11 A' 1557 1483 8.70      
12 A' 1546 1472 8.35      
13 A' 1496 1425 2.22      
14 A' 1470 1400 3.77      
15 A' 1461 1392 6.00      
16 A' 1385 1319 3.22      
17 A' 1350 1286 0.41      
18 A' 1228 1169 2.09      
19 A' 1140 1085 9.46      
20 A' 1034 984 1.20      
21 A' 997 950 0.28      
22 A' 928 884 1.16      
23 A' 749 713 0.86      
24 A' 532 506 1.02      
25 A' 450 429 0.79      
26 A' 338 322 0.18      
27 A' 287 274 0.25      
28 A' 265 252 0.17      
29 A" 3139 2990 11.41      
30 A" 3132 2983 10.99      
31 A" 3122 2973 23.34      
32 A" 3058 2912 23.03      
33 A" 1551 1477 0.21      
34 A" 1539 1465 1.04      
35 A" 1530 1457 7.80      
36 A" 1449 1380 5.61      
37 A" 1389 1323 1.41      
38 A" 1165 1109 2.18      
39 A" 1093 1041 0.18      
40 A" 994 947 0.13      
41 A" 952 906 0.97      
42 A" 913 870 40.23      
43 A" 747 712 0.47      
44 A" 556 529 5.96      
45 A" 274 261 0.03      
46 A" 215 205 0.29      
47 A" 165 157 0.14      
48 A" 19 18 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 36765.4 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 35011.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.14711 0.08294 0.08198

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.957 -1.819 0.000
C2 -0.063 -0.818 0.000
C3 -0.518 0.633 0.000
C4 1.423 -1.096 0.000
C5 -0.063 1.378 1.264
C6 -0.063 1.378 -1.264
H7 -0.651 -2.864 0.000
H8 -2.029 -1.629 0.000
H9 -1.619 0.622 0.000
H10 1.616 -2.175 0.000
H11 1.913 -0.665 -0.883
H12 1.913 -0.665 0.883
H13 -0.494 2.388 1.289
H14 -0.381 0.847 2.170
H15 1.029 1.484 1.294
H16 -0.494 2.388 -1.289
H17 -0.381 0.847 -2.170
H18 1.029 1.484 -1.294

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34182.49082.48743.55193.55191.08931.08922.52912.59773.21683.21684.42383.48514.06524.42383.48514.0652
C21.34181.52071.51202.53372.53372.12882.12702.11992.15902.17002.17003.48192.75332.85763.48192.75332.8576
C32.49081.52072.59961.53601.53603.49972.72051.10063.52702.89422.89422.17742.18472.18912.17742.18472.1891
C42.48741.51202.59963.15063.15062.72533.49353.49361.09591.09811.09814.18003.42572.91294.18003.42572.9129
C53.55192.53371.53603.15062.52744.46533.80882.14224.12793.56222.86781.09801.09731.09772.77893.48892.7828
C63.55192.53371.53603.15062.52744.46533.80882.14224.12792.86783.56222.77893.48892.78281.09801.09731.0977
H71.08932.12883.49972.72534.46534.46531.85113.61822.36943.49123.49125.40994.30724.83755.40994.30724.8375
H81.08922.12702.72053.49353.80883.80881.85112.28843.68634.15374.15374.48933.68204.55194.48933.68204.5519
H92.52912.11991.10063.49362.14222.14223.61822.28844.27653.86153.86152.45852.50843.07052.45852.50843.0705
H102.59772.15903.52701.09594.12794.12792.36943.68634.27651.77401.77405.18944.22193.92485.18944.22193.9248
H113.21682.17002.89421.09813.56222.86783.49124.15373.86151.77401.76664.45344.10713.18423.90873.03342.3597
H123.21682.17002.89421.09812.86783.56223.49124.15373.86151.77401.76663.90873.03342.35974.45344.10713.1842
H134.42383.48192.17744.18001.09802.77895.40994.48932.45855.18944.45343.90871.77831.77102.57843.78843.1318
H143.48512.75332.18473.42571.09733.48894.30723.68202.50844.22194.10713.03341.77831.77783.78844.33973.7933
H154.06522.85762.18912.91291.09772.78284.83754.55193.07053.92483.18422.35971.77101.77783.13183.79332.5874
H164.42383.48192.17744.18002.77891.09805.40994.48932.45855.18943.90874.45342.57843.78843.13181.77831.7710
H173.48512.75332.18473.42573.48891.09734.30723.68202.50844.22193.03344.10713.78844.33973.79331.77831.7778
H184.06522.85762.18912.91292.78281.09774.83754.55193.07053.92482.35973.18423.13183.79332.58741.77101.7778

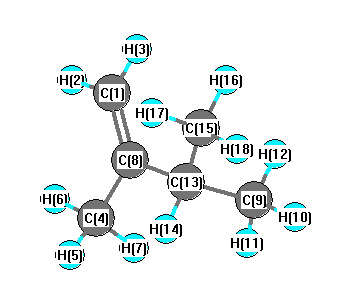
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.820 C1 C2 C4 121.174
C2 C1 H7 121.904 C2 C1 H8 121.736
C2 C3 C5 111.972 C2 C3 C6 111.972
C2 C3 H9 106.845 C2 C4 H10 110.745
C2 C4 H11 111.497 C2 C4 H12 111.497
C3 C2 C4 118.006 C3 C5 H13 110.411
C3 C5 H14 111.024 C3 C5 H15 111.354
C3 C6 H16 110.411 C3 C6 H17 111.024
C3 C6 H18 111.354 C5 C3 C6 110.716
C5 C3 H9 107.528 C6 C3 H9 107.528
H7 C1 H8 116.360 H10 C4 H11 107.911
H10 C4 H12 107.911 H11 C4 H12 107.099
H13 C5 H14 108.204 H13 C5 H15 107.533
H14 C5 H15 108.181 H16 C6 H17 108.204
H16 C6 H18 107.533 H17 C6 H18 108.181
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability