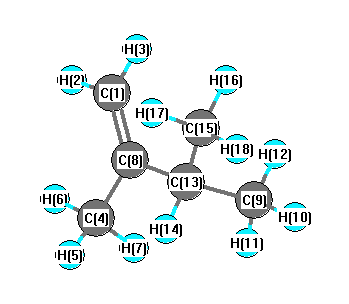
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3255 |
3095 |
20.21 |
|
|
|
| 2 |
A' |
3174 |
3018 |
10.99 |
|
|
|
| 3 |
A' |
3158 |
3004 |
22.87 |
|
|
|
| 4 |
A' |
3152 |
2997 |
24.92 |
|
|
|
| 5 |
A' |
3144 |
2990 |
71.79 |
|
|
|
| 6 |
A' |
3068 |
2918 |
32.13 |
|
|
|
| 7 |
A' |
3062 |
2912 |
0.27 |
|
|
|
| 8 |
A' |
3059 |
2909 |
25.31 |
|
|
|
| 9 |
A' |
1751 |
1666 |
14.78 |
|
|
|
| 10 |
A' |
1540 |
1465 |
9.52 |
|
|
|
| 11 |
A' |
1533 |
1458 |
6.56 |
|
|
|
| 12 |
A' |
1519 |
1444 |
11.47 |
|
|
|
| 13 |
A' |
1474 |
1402 |
1.17 |
|
|
|
| 14 |
A' |
1442 |
1371 |
3.75 |
|
|
|
| 15 |
A' |
1433 |
1363 |
8.52 |
|
|
|
| 16 |
A' |
1360 |
1294 |
3.94 |
|
|
|
| 17 |
A' |
1329 |
1264 |
0.36 |
|
|
|
| 18 |
A' |
1205 |
1146 |
2.46 |
|
|
|
| 19 |
A' |
1124 |
1069 |
13.20 |
|
|
|
| 20 |
A' |
1016 |
966 |
1.74 |
|
|
|
| 21 |
A' |
982 |
934 |
0.60 |
|
|
|
| 22 |
A' |
921 |
876 |
1.83 |
|
|
|
| 23 |
A' |
747 |
711 |
1.33 |
|
|
|
| 24 |
A' |
524 |
499 |
1.41 |
|
|
|
| 25 |
A' |
442 |
420 |
0.84 |
|
|
|
| 26 |
A' |
332 |
316 |
0.25 |
|
|
|
| 27 |
A' |
278 |
265 |
0.49 |
|
|
|
| 28 |
A' |
262 |
249 |
0.07 |
|
|
|
| 29 |
A" |
3150 |
2996 |
18.85 |
|
|
|
| 30 |
A" |
3139 |
2986 |
0.12 |
|
|
|
| 31 |
A" |
3120 |
2967 |
23.66 |
|
|
|
| 32 |
A" |
3064 |
2913 |
24.05 |
|
|
|
| 33 |
A" |
1524 |
1449 |
0.13 |
|
|
|
| 34 |
A" |
1515 |
1441 |
0.90 |
|
|
|
| 35 |
A" |
1501 |
1427 |
9.98 |
|
|
|
| 36 |
A" |
1419 |
1349 |
6.03 |
|
|
|
| 37 |
A" |
1359 |
1292 |
1.80 |
|
|
|
| 38 |
A" |
1147 |
1091 |
1.53 |
|
|
|
| 39 |
A" |
1078 |
1025 |
1.01 |
|
|
|
| 40 |
A" |
979 |
931 |
0.18 |
|
|
|
| 41 |
A" |
939 |
893 |
7.41 |
|
|
|
| 42 |
A" |
925 |
880 |
37.52 |
|
|
|
| 43 |
A" |
749 |
713 |
0.55 |
|
|
|
| 44 |
A" |
556 |
529 |
7.02 |
|
|
|
| 45 |
A" |
261 |
249 |
0.01 |
|
|
|
| 46 |
A" |
215 |
204 |
0.41 |
|
|
|
| 47 |
A" |
165 |
157 |
0.18 |
|
|
|
| 48 |
A" |
40 |
38 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36564.7 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 34773.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.452 |
|
|
|
| 2 |
C |
0.226 |
|
|
|
| 3 |
C |
-0.195 |
|
|
|
| 4 |
C |
-0.586 |
|
|
|
| 5 |
C |
-0.513 |
|
|
|
| 6 |
C |
-0.513 |
|
|
|
| 7 |
H |
0.155 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
| 10 |
H |
0.179 |
|
|
|
| 11 |
H |
0.180 |
|
|
|
| 12 |
H |
0.180 |
|
|
|
| 13 |
H |
0.169 |
|
|
|
| 14 |
H |
0.176 |
|
|
|
| 15 |
H |
0.166 |
|
|
|
| 16 |
H |
0.169 |
|
|
|
| 17 |
H |
0.176 |
|
|
|
| 18 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.277 |
0.379 |
0.000 |
0.469 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.457 |
-0.675 |
0.000 |
| y |
-0.675 |
-39.100 |
0.000 |
| z |
0.000 |
0.000 |
-40.982 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.585 |
-0.675 |
0.000 |
| y |
-0.675 |
0.620 |
0.000 |
| z |
0.000 |
0.000 |
-2.204 |
|
| Polar |
|---|
| 3z2-r2 | -4.409 |
|---|
| x2-y2 | 0.643 |
|---|
| xy | -0.675 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.611 |
0.799 |
0.000 |
| y |
0.799 |
10.918 |
0.000 |
| z |
0.000 |
0.000 |
7.573 |
<r2> (average value of r
2) Å
2
| <r2> |
190.089 |
| (<r2>)1/2 |
13.787 |