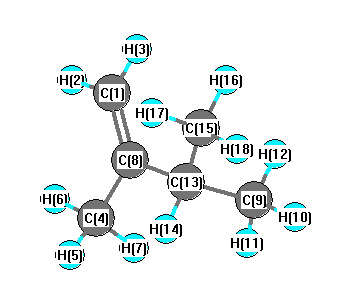
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3161 |
3109 |
22.21 |
|
|
|
| 2 |
A' |
3082 |
3031 |
11.81 |
|
|
|
| 3 |
A' |
3069 |
3018 |
23.53 |
|
|
|
| 4 |
A' |
3063 |
3012 |
23.91 |
|
|
|
| 5 |
A' |
3054 |
3004 |
70.31 |
|
|
|
| 6 |
A' |
2982 |
2933 |
31.29 |
|
|
|
| 7 |
A' |
2973 |
2924 |
9.92 |
|
|
|
| 8 |
A' |
2965 |
2916 |
19.49 |
|
|
|
| 9 |
A' |
1681 |
1653 |
13.63 |
|
|
|
| 10 |
A' |
1493 |
1469 |
6.82 |
|
|
|
| 11 |
A' |
1488 |
1463 |
7.80 |
|
|
|
| 12 |
A' |
1474 |
1450 |
10.68 |
|
|
|
| 13 |
A' |
1427 |
1404 |
0.61 |
|
|
|
| 14 |
A' |
1393 |
1370 |
2.74 |
|
|
|
| 15 |
A' |
1384 |
1361 |
7.30 |
|
|
|
| 16 |
A' |
1313 |
1291 |
4.18 |
|
|
|
| 17 |
A' |
1280 |
1259 |
0.51 |
|
|
|
| 18 |
A' |
1163 |
1144 |
2.57 |
|
|
|
| 19 |
A' |
1088 |
1071 |
14.10 |
|
|
|
| 20 |
A' |
987 |
970 |
1.81 |
|
|
|
| 21 |
A' |
949 |
934 |
0.89 |
|
|
|
| 22 |
A' |
889 |
874 |
1.88 |
|
|
|
| 23 |
A' |
722 |
710 |
1.47 |
|
|
|
| 24 |
A' |
508 |
500 |
1.44 |
|
|
|
| 25 |
A' |
430 |
423 |
0.86 |
|
|
|
| 26 |
A' |
321 |
315 |
0.22 |
|
|
|
| 27 |
A' |
273 |
268 |
0.41 |
|
|
|
| 28 |
A' |
250 |
246 |
0.17 |
|
|
|
| 29 |
A" |
3061 |
3011 |
19.84 |
|
|
|
| 30 |
A" |
3049 |
2999 |
0.01 |
|
|
|
| 31 |
A" |
3027 |
2977 |
23.46 |
|
|
|
| 32 |
A" |
2978 |
2929 |
24.50 |
|
|
|
| 33 |
A" |
1479 |
1455 |
0.25 |
|
|
|
| 34 |
A" |
1470 |
1445 |
0.63 |
|
|
|
| 35 |
A" |
1456 |
1432 |
8.93 |
|
|
|
| 36 |
A" |
1369 |
1346 |
4.26 |
|
|
|
| 37 |
A" |
1311 |
1290 |
2.93 |
|
|
|
| 38 |
A" |
1103 |
1085 |
1.57 |
|
|
|
| 39 |
A" |
1043 |
1025 |
1.17 |
|
|
|
| 40 |
A" |
946 |
931 |
0.13 |
|
|
|
| 41 |
A" |
909 |
894 |
2.28 |
|
|
|
| 42 |
A" |
876 |
862 |
38.93 |
|
|
|
| 43 |
A" |
721 |
709 |
0.45 |
|
|
|
| 44 |
A" |
535 |
526 |
6.50 |
|
|
|
| 45 |
A" |
255 |
250 |
0.01 |
|
|
|
| 46 |
A" |
207 |
203 |
0.42 |
|
|
|
| 47 |
A" |
174 |
172 |
0.21 |
|
|
|
| 48 |
A" |
41 |
40 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35435.6 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 34850.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.426 |
|
|
|
| 2 |
C |
0.257 |
|
|
|
| 3 |
C |
-0.169 |
|
|
|
| 4 |
C |
-0.567 |
|
|
|
| 5 |
C |
-0.484 |
|
|
|
| 6 |
C |
-0.484 |
|
|
|
| 7 |
H |
0.137 |
|
|
|
| 8 |
H |
0.139 |
|
|
|
| 9 |
H |
0.142 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
| 12 |
H |
0.169 |
|
|
|
| 13 |
H |
0.156 |
|
|
|
| 14 |
H |
0.163 |
|
|
|
| 15 |
H |
0.155 |
|
|
|
| 16 |
H |
0.156 |
|
|
|
| 17 |
H |
0.163 |
|
|
|
| 18 |
H |
0.155 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.300 |
0.397 |
0.000 |
0.498 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.605 |
-0.700 |
0.000 |
| y |
-0.700 |
-39.271 |
0.000 |
| z |
0.000 |
0.000 |
-40.996 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.528 |
-0.700 |
0.000 |
| y |
-0.700 |
0.530 |
0.000 |
| z |
0.000 |
0.000 |
-2.058 |
|
| Polar |
|---|
| 3z2-r2 | -4.116 |
|---|
| x2-y2 | 0.666 |
|---|
| xy | -0.700 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.898 |
0.753 |
0.000 |
| y |
0.753 |
11.296 |
0.000 |
| z |
0.000 |
0.000 |
7.805 |
<r2> (average value of r
2) Å
2
| <r2> |
192.523 |
| (<r2>)1/2 |
13.875 |