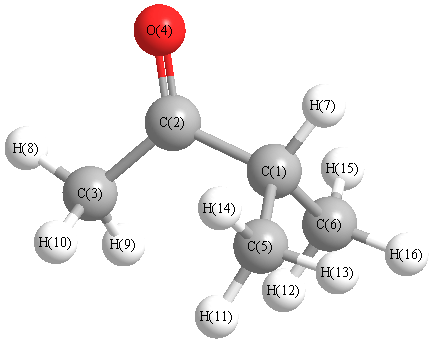
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3182 |
3044 |
9.26 |
|
|
|
| 2 |
A |
3143 |
3007 |
20.67 |
|
|
|
| 3 |
A |
3134 |
2999 |
56.95 |
|
|
|
| 4 |
A |
3133 |
2998 |
31.78 |
|
|
|
| 5 |
A |
3131 |
2996 |
0.28 |
|
|
|
| 6 |
A |
3128 |
2992 |
4.77 |
|
|
|
| 7 |
A |
3082 |
2948 |
4.97 |
|
|
|
| 8 |
A |
3063 |
2931 |
5.20 |
|
|
|
| 9 |
A |
3056 |
2924 |
15.12 |
|
|
|
| 10 |
A |
3054 |
2922 |
23.85 |
|
|
|
| 11 |
A |
1825 |
1746 |
162.05 |
|
|
|
| 12 |
A |
1536 |
1470 |
20.28 |
|
|
|
| 13 |
A |
1530 |
1464 |
6.95 |
|
|
|
| 14 |
A |
1518 |
1452 |
3.58 |
|
|
|
| 15 |
A |
1514 |
1448 |
0.85 |
|
|
|
| 16 |
A |
1497 |
1433 |
10.23 |
|
|
|
| 17 |
A |
1487 |
1422 |
11.00 |
|
|
|
| 18 |
A |
1439 |
1377 |
4.53 |
|
|
|
| 19 |
A |
1417 |
1356 |
4.88 |
|
|
|
| 20 |
A |
1404 |
1344 |
46.04 |
|
|
|
| 21 |
A |
1360 |
1301 |
1.70 |
|
|
|
| 22 |
A |
1340 |
1282 |
1.02 |
|
|
|
| 23 |
A |
1269 |
1214 |
49.63 |
|
|
|
| 24 |
A |
1196 |
1144 |
2.59 |
|
|
|
| 25 |
A |
1141 |
1092 |
0.94 |
|
|
|
| 26 |
A |
1101 |
1054 |
23.09 |
|
|
|
| 27 |
A |
1022 |
978 |
2.08 |
|
|
|
| 28 |
A |
976 |
934 |
1.40 |
|
|
|
| 29 |
A |
949 |
908 |
19.18 |
|
|
|
| 30 |
A |
940 |
899 |
1.53 |
|
|
|
| 31 |
A |
910 |
870 |
4.42 |
|
|
|
| 32 |
A |
728 |
697 |
5.29 |
|
|
|
| 33 |
A |
599 |
573 |
0.26 |
|
|
|
| 34 |
A |
526 |
503 |
8.16 |
|
|
|
| 35 |
A |
490 |
469 |
3.82 |
|
|
|
| 36 |
A |
332 |
317 |
1.41 |
|
|
|
| 37 |
A |
271 |
260 |
1.44 |
|
|
|
| 38 |
A |
252 |
241 |
0.37 |
|
|
|
| 39 |
A |
238 |
228 |
0.16 |
|
|
|
| 40 |
A |
209 |
200 |
0.34 |
|
|
|
| 41 |
A |
135 |
129 |
0.26 |
|
|
|
| 42 |
A |
47 |
45 |
3.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31150.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29802.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.444 |
|
|
-0.498 |
| 2 |
C |
0.469 |
|
|
0.549 |
| 3 |
H |
0.199 |
|
|
0.128 |
| 4 |
H |
0.205 |
|
|
0.138 |
| 5 |
H |
0.186 |
|
|
0.116 |
| 6 |
C |
-0.610 |
|
|
-0.465 |
| 7 |
H |
0.187 |
|
|
-0.013 |
| 8 |
C |
-0.208 |
|
|
0.269 |
| 9 |
H |
0.178 |
|
|
0.104 |
| 10 |
H |
0.189 |
|
|
0.113 |
| 11 |
H |
0.169 |
|
|
0.111 |
| 12 |
H |
0.167 |
|
|
0.110 |
| 13 |
H |
0.179 |
|
|
0.126 |
| 14 |
H |
0.178 |
|
|
0.129 |
| 15 |
C |
-0.520 |
|
|
-0.454 |
| 16 |
C |
-0.523 |
|
|
-0.466 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.931 |
1.634 |
1.291 |
2.840 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.966 |
1.655 |
1.324 |
2.890 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.239 |
3.393 |
2.728 |
| y |
3.393 |
-38.203 |
-1.350 |
| z |
2.728 |
-1.350 |
-37.974 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.150 |
3.393 |
2.728 |
| y |
3.393 |
0.903 |
-1.350 |
| z |
2.728 |
-1.350 |
1.247 |
|
| Polar |
|---|
| 3z2-r2 | 2.495 |
|---|
| x2-y2 | -2.035 |
|---|
| xy | 3.393 |
|---|
| xz | 2.728 |
|---|
| yz | -1.350 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
179.868 |
| (<r2>)1/2 |
13.411 |