return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2-Butanone, 3-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-271.457275
Energy at 298.15K-271.467853
HF Energy-271.182833
Nuclear repulsion energy244.152516
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3201 3038 9.76      
2 A 3163 3002 21.16      
3 A 3156 2996 59.31      
4 A 3154 2994 24.70      
5 A 3153 2992 5.67      
6 A 3148 2989 6.20      
7 A 3101 2944 5.45      
8 A 3087 2930 6.17      
9 A 3079 2923 11.30      
10 A 3077 2921 22.32      
11 A 1790 1699 129.89      
12 A 1560 1481 16.65      
13 A 1555 1476 5.90      
14 A 1544 1466 3.04      
15 A 1540 1462 0.93      
16 A 1524 1447 8.84      
17 A 1514 1437 10.59      
18 A 1464 1390 3.12      
19 A 1442 1369 3.43      
20 A 1430 1357 36.70      
21 A 1383 1312 2.03      
22 A 1362 1292 1.37      
23 A 1286 1220 47.76      
24 A 1215 1154 2.64      
25 A 1153 1095 0.99      
26 A 1118 1061 18.73      
27 A 1037 985 1.63      
28 A 991 941 0.92      
29 A 964 915 18.27      
30 A 954 906 1.37      
31 A 915 869 3.89      
32 A 732 695 4.49      
33 A 604 573 0.31      
34 A 529 502 7.54      
35 A 497 472 5.03      
36 A 337 320 1.40      
37 A 277 263 1.39      
38 A 261 247 0.29      
39 A 241 229 0.19      
40 A 214 203 0.46      
41 A 139 132 0.15      
42 A 48 46 3.06      

Unscaled Zero Point Vibrational Energy (zpe) 31468.5 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29869.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.15104 0.09061 0.07893

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 1.609 -0.317 -1.099
C2 0.819 -0.037 -0.215
H3 1.965 1.250 1.064
H4 1.949 -0.435 1.552
H5 0.532 0.572 1.883
C6 1.327 0.379 1.161
H7 -0.799 -0.423 -1.488
C8 -0.690 -0.108 -0.448
H9 -0.985 2.015 -0.856
H10 -0.793 -2.182 0.266
H11 -1.320 1.566 0.816
H12 -1.214 -0.978 1.500
H13 -2.357 -1.337 0.204
H14 -2.461 1.136 -0.450
C15 -1.293 -1.230 0.438
C16 -1.394 1.220 -0.220

Atom - Atom Distances (Å)
  O1 C2 H3 H4 H5 C6 H7 C8 H9 H10 H11 H12 H13 H14 C15 C16
O11.21822.69482.67493.29282.38112.44242.39893.49743.33363.97413.89404.29734.37063.40923.4870
C21.21822.14632.13422.20341.52402.09531.52882.80722.72602.86512.82153.45713.49202.51252.5460
H32.69482.14631.75441.78441.08474.11803.34393.60284.47483.31013.90685.11024.68004.14273.5972
H42.67492.13421.75441.76941.09654.09783.32694.51773.49563.90263.20944.60055.09163.51884.1300
H53.29282.20341.78441.76941.09103.75862.71833.44773.45762.35742.36623.84843.83712.94412.9250
C62.38111.52401.08471.09651.09103.48982.62513.47713.44242.92122.90024.17444.18543.15823.1654
H72.44242.09534.11804.09783.75863.48981.09232.52572.48443.08763.06722.47442.50382.14592.1592
C82.39891.52883.34393.32692.71832.62511.09232.18182.19632.18962.19682.17122.16431.55181.5205
H93.49742.80723.60284.51773.44773.47712.52572.18184.34901.76313.81573.77381.76503.50691.0969
H103.33362.72604.47483.49563.45763.44242.48442.19634.34903.82431.77481.77853.78261.08963.4893
H113.97412.86513.31013.90262.35742.92123.08762.18961.76313.82432.63593.14241.75782.82081.0945
H123.89402.82153.90683.20942.36622.90023.06722.19683.81571.77482.63591.76463.13491.09422.7969
H134.29733.45715.11024.60053.84844.17442.47442.17123.77381.77853.14241.76462.56051.09392.7652
H144.37063.49204.68005.09163.83714.18542.50382.16431.76503.78261.75783.13492.56052.78391.0946
C153.40922.51254.14273.51882.94413.15822.14591.55183.50691.08962.82081.09421.09392.78392.5389
C163.48702.54603.59724.13002.92503.16542.15921.52051.09693.48931.09452.79692.76521.09462.5389

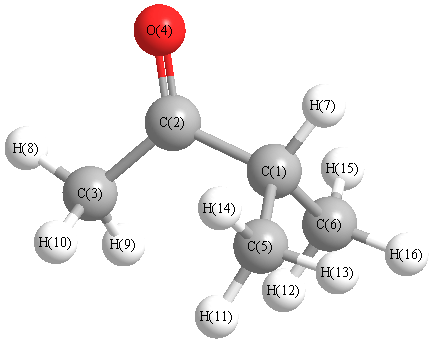
picture of 2-Butanone, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C6 120.118 O1 C2 C8 121.270
C2 C6 H3 109.576 C2 C6 H4 107.949
C2 C6 H5 113.802 C2 C8 H7 104.915
C2 C8 C15 109.284 C2 C8 C16 113.221
H3 C6 H4 107.096 H3 C6 H5 110.197
H4 C6 H5 107.967 C6 C2 C8 118.609
H7 C8 C15 107.217 H7 C8 C16 110.389
C8 C15 H10 111.300 C8 C15 H12 111.066
C8 C15 H13 109.071 C8 C16 H9 111.920
C8 C16 H11 112.700 C8 C16 H14 110.659
H9 C16 H11 107.136 H9 C16 H14 107.293
H10 C15 H12 108.721 H10 C15 H13 109.080
H11 C16 H14 106.831 H12 C15 H13 107.509
C15 C8 C16 111.455
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 O -0.488      
2 C 0.492      
3 H 0.184      
4 H 0.194      
5 H 0.172      
6 C -0.559      
7 H 0.176      
8 C -0.191      
9 H 0.163      
10 H 0.175      
11 H 0.154      
12 H 0.152      
13 H 0.164      
14 H 0.164      
15 C -0.475      
16 C -0.477      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.661 -0.138 -0.074
y -0.138 7.864 0.376
z -0.074 0.376 7.304


<r2> (average value of r2) Å2
<r2> 180.349
(<r2>)1/2 13.429