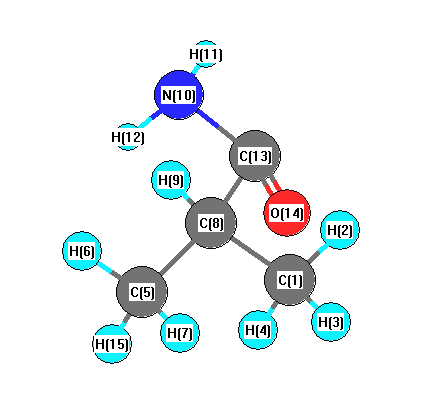
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3773 |
3585 |
36.31 |
|
|
|
| 2 |
A |
3638 |
3457 |
37.89 |
|
|
|
| 3 |
A |
3171 |
3014 |
25.14 |
|
|
|
| 4 |
A |
3168 |
3011 |
2.94 |
|
|
|
| 5 |
A |
3153 |
2996 |
34.91 |
|
|
|
| 6 |
A |
3151 |
2994 |
16.91 |
|
|
|
| 7 |
A |
3076 |
2923 |
13.46 |
|
|
|
| 8 |
A |
3073 |
2921 |
31.19 |
|
|
|
| 9 |
A |
3057 |
2905 |
35.55 |
|
|
|
| 10 |
A |
1838 |
1747 |
263.28 |
|
|
|
| 11 |
A |
1650 |
1568 |
129.57 |
|
|
|
| 12 |
A |
1543 |
1467 |
18.37 |
|
|
|
| 13 |
A |
1527 |
1451 |
11.51 |
|
|
|
| 14 |
A |
1519 |
1443 |
1.93 |
|
|
|
| 15 |
A |
1511 |
1436 |
0.79 |
|
|
|
| 16 |
A |
1447 |
1375 |
72.79 |
|
|
|
| 17 |
A |
1433 |
1362 |
5.36 |
|
|
|
| 18 |
A |
1417 |
1346 |
6.13 |
|
|
|
| 19 |
A |
1351 |
1284 |
1.04 |
|
|
|
| 20 |
A |
1305 |
1240 |
85.88 |
|
|
|
| 21 |
A |
1214 |
1154 |
2.94 |
|
|
|
| 22 |
A |
1150 |
1093 |
5.52 |
|
|
|
| 23 |
A |
1148 |
1091 |
0.00 |
|
|
|
| 24 |
A |
1074 |
1020 |
7.17 |
|
|
|
| 25 |
A |
992 |
943 |
0.31 |
|
|
|
| 26 |
A |
941 |
894 |
2.89 |
|
|
|
| 27 |
A |
941 |
894 |
3.73 |
|
|
|
| 28 |
A |
785 |
746 |
7.63 |
|
|
|
| 29 |
A |
773 |
735 |
3.77 |
|
|
|
| 30 |
A |
619 |
588 |
6.08 |
|
|
|
| 31 |
A |
608 |
577 |
15.12 |
|
|
|
| 32 |
A |
479 |
455 |
3.15 |
|
|
|
| 33 |
A |
327 |
311 |
0.95 |
|
|
|
| 34 |
A |
293 |
278 |
4.84 |
|
|
|
| 35 |
A |
247 |
235 |
6.02 |
|
|
|
| 36 |
A |
245 |
233 |
0.39 |
|
|
|
| 37 |
A |
218 |
207 |
1.46 |
|
|
|
| 38 |
A |
128 |
121 |
230.79 |
|
|
|
| 39 |
A |
25 |
24 |
8.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29003.0 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 27561.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.517 |
|
|
|
| 2 |
H |
0.174 |
|
|
|
| 3 |
H |
0.202 |
|
|
|
| 4 |
H |
0.171 |
|
|
|
| 5 |
C |
-0.517 |
|
|
|
| 6 |
H |
0.174 |
|
|
|
| 7 |
H |
0.202 |
|
|
|
| 8 |
C |
-0.235 |
|
|
|
| 9 |
H |
0.151 |
|
|
|
| 10 |
N |
-0.802 |
|
|
|
| 11 |
H |
0.363 |
|
|
|
| 12 |
H |
0.357 |
|
|
|
| 13 |
C |
0.625 |
|
|
|
| 14 |
O |
-0.519 |
|
|
|
| 15 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.333 |
0.000 |
-3.554 |
3.570 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.033 |
0.000 |
-4.713 |
| y |
0.000 |
-37.758 |
0.000 |
| z |
-4.713 |
0.000 |
-38.410 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.050 |
0.000 |
-4.713 |
| y |
0.000 |
-2.536 |
0.000 |
| z |
-4.713 |
0.000 |
-3.514 |
|
| Polar |
|---|
| 3z2-r2 | -7.028 |
|---|
| x2-y2 | 5.724 |
|---|
| xy | 0.000 |
|---|
| xz | -4.713 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.355 |
0.000 |
-0.314 |
| y |
0.000 |
6.389 |
0.000 |
| z |
-0.314 |
0.000 |
7.609 |
<r2> (average value of r
2) Å
2
| <r2> |
171.629 |
| (<r2>)1/2 |
13.101 |