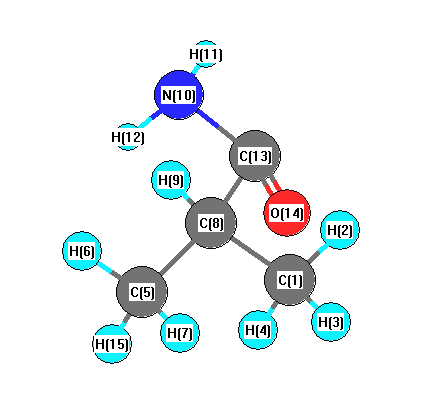
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3768 |
3583 |
35.16 |
|
|
|
| 2 |
A |
3633 |
3455 |
36.46 |
|
|
|
| 3 |
A |
3170 |
3015 |
26.50 |
|
|
|
| 4 |
A |
3167 |
3011 |
3.04 |
|
|
|
| 5 |
A |
3151 |
2997 |
36.37 |
|
|
|
| 6 |
A |
3149 |
2995 |
17.64 |
|
|
|
| 7 |
A |
3075 |
2925 |
13.83 |
|
|
|
| 8 |
A |
3072 |
2922 |
32.18 |
|
|
|
| 9 |
A |
3054 |
2904 |
36.81 |
|
|
|
| 10 |
A |
1835 |
1745 |
262.75 |
|
|
|
| 11 |
A |
1650 |
1569 |
128.24 |
|
|
|
| 12 |
A |
1544 |
1468 |
17.75 |
|
|
|
| 13 |
A |
1528 |
1453 |
11.39 |
|
|
|
| 14 |
A |
1519 |
1445 |
1.90 |
|
|
|
| 15 |
A |
1512 |
1438 |
0.71 |
|
|
|
| 16 |
A |
1446 |
1375 |
70.29 |
|
|
|
| 17 |
A |
1433 |
1362 |
6.92 |
|
|
|
| 18 |
A |
1417 |
1348 |
5.81 |
|
|
|
| 19 |
A |
1351 |
1285 |
1.16 |
|
|
|
| 20 |
A |
1304 |
1240 |
87.05 |
|
|
|
| 21 |
A |
1214 |
1155 |
2.90 |
|
|
|
| 22 |
A |
1149 |
1093 |
5.56 |
|
|
|
| 23 |
A |
1147 |
1090 |
0.00 |
|
|
|
| 24 |
A |
1073 |
1021 |
6.92 |
|
|
|
| 25 |
A |
992 |
943 |
0.29 |
|
|
|
| 26 |
A |
941 |
895 |
2.79 |
|
|
|
| 27 |
A |
940 |
894 |
3.70 |
|
|
|
| 28 |
A |
785 |
746 |
7.66 |
|
|
|
| 29 |
A |
772 |
735 |
3.75 |
|
|
|
| 30 |
A |
619 |
589 |
6.09 |
|
|
|
| 31 |
A |
607 |
578 |
15.17 |
|
|
|
| 32 |
A |
479 |
456 |
3.19 |
|
|
|
| 33 |
A |
328 |
312 |
0.96 |
|
|
|
| 34 |
A |
294 |
279 |
4.81 |
|
|
|
| 35 |
A |
249 |
236 |
6.05 |
|
|
|
| 36 |
A |
246 |
234 |
0.31 |
|
|
|
| 37 |
A |
219 |
208 |
1.31 |
|
|
|
| 38 |
A |
123 |
117 |
229.26 |
|
|
|
| 39 |
A |
25 |
24 |
9.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28989.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 27568.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.511 |
|
|
|
| 2 |
H |
0.172 |
|
|
|
| 3 |
H |
0.199 |
|
|
|
| 4 |
H |
0.169 |
|
|
|
| 5 |
C |
-0.511 |
|
|
|
| 6 |
H |
0.172 |
|
|
|
| 7 |
H |
0.199 |
|
|
|
| 8 |
C |
-0.225 |
|
|
|
| 9 |
H |
0.146 |
|
|
|
| 10 |
N |
-0.797 |
|
|
|
| 11 |
H |
0.361 |
|
|
|
| 12 |
H |
0.356 |
|
|
|
| 13 |
C |
0.622 |
|
|
|
| 14 |
O |
-0.519 |
|
|
|
| 15 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.337 |
0.000 |
-3.553 |
3.569 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.041 |
0.000 |
-4.714 |
| y |
0.000 |
-37.757 |
0.000 |
| z |
-4.714 |
0.000 |
-38.432 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.053 |
0.000 |
-4.714 |
| y |
0.000 |
-2.521 |
0.000 |
| z |
-4.714 |
0.000 |
-3.533 |
|
| Polar |
|---|
| 3z2-r2 | -7.065 |
|---|
| x2-y2 | 5.716 |
|---|
| xy | 0.000 |
|---|
| xz | -4.714 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.374 |
0.000 |
-0.315 |
| y |
0.000 |
6.396 |
0.000 |
| z |
-0.315 |
0.000 |
7.620 |
<r2> (average value of r
2) Å
2
| <r2> |
171.618 |
| (<r2>)1/2 |
13.100 |