return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9NO (Propanamide, 2-methyl-)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-286.938780
Energy at 298.15K-286.949263
HF Energy-286.044402
Nuclear repulsion energy247.914817
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3781 3549 38.25      
2 A 3654 3430 47.87      
3 A 3202 3005 21.14      
4 A 3197 3001 6.32      
5 A 3181 2985 31.00      
6 A 3178 2983 19.83      
7 A 3105 2915 11.07      
8 A 3103 2912 26.91      
9 A 3088 2898 30.02      
10 A 1874 1759 270.95      
11 A 1679 1576 133.66      
12 A 1574 1477 17.41      
13 A 1562 1466 10.35      
14 A 1552 1457 1.93      
15 A 1547 1452 1.00      
16 A 1484 1393 73.27      
17 A 1474 1384 4.64      
18 A 1456 1366 6.10      
19 A 1390 1305 0.88      
20 A 1333 1251 96.83      
21 A 1243 1167 4.02      
22 A 1179 1106 6.58      
23 A 1172 1100 0.13      
24 A 1105 1037 6.81      
25 A 1012 950 0.28      
26 A 962 903 2.24      
27 A 958 899 2.47      
28 A 795 746 7.71      
29 A 788 740 5.68      
30 A 635 596 13.94      
31 A 593 556 28.23      
32 A 489 459 3.65      
33 A 335 315 4.01      
34 A 299 281 8.60      
35 A 267 251 183.29      
36 A 252 237 4.38      
37 A 245 230 61.51      
38 A 216 203 17.28      
39 A 24 23 6.03      

Unscaled Zero Point Vibrational Energy (zpe) 29491.2 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 27680.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.16425 0.08824 0.08418

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.390 -1.209 -0.257
H2 -0.907 -2.047 -0.771
H3 -1.435 -1.444 0.809
H4 -2.409 -1.111 -0.643
C5 -1.311 1.275 0.221
H6 -0.762 2.209 0.057
H7 -1.366 1.094 1.298
C8 -0.613 0.094 -0.463
H9 -0.536 0.302 -1.539
N10 1.824 0.217 -0.722
H11 2.755 0.022 -0.384
H12 1.685 0.263 -1.718
C13 0.785 -0.055 0.125
O14 0.963 -0.354 1.295
H15 -2.325 1.398 -0.170

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 N10 H11 H12 C13 O14 H15
C11.09491.09291.09462.53153.48912.77951.53132.15763.54584.32543.70892.49142.94502.7714
H21.09491.77221.77433.49074.33763.78942.18302.49883.54694.22353.59852.76273.26013.7736
H31.09291.77221.78022.78503.78942.58592.15893.06103.96484.59664.36312.70662.67823.1350
H41.09461.77431.78022.76603.77153.11792.17092.51164.43685.29344.45053.45093.96212.5550
C52.53153.49072.78502.76601.09471.09331.53292.15483.43984.29733.70952.48412.99611.0946
H63.48914.33763.78943.77151.09471.77382.18242.49613.35584.16523.59552.74293.32821.7753
H72.77953.78942.58593.11791.09331.77382.16073.05973.87614.57874.37032.70642.74281.7796
C81.53132.18302.15892.17091.53292.18242.16071.09822.45313.36962.62391.52382.40272.1722
H92.15762.49883.06102.51162.15482.49613.05971.09822.49853.49912.22892.15423.27182.5050
N103.54583.54693.96484.43683.43983.35583.87612.45312.49851.00991.00711.36732.26554.3490
H114.32544.22354.59665.29344.29734.16524.57873.36963.49911.00991.72712.03622.48425.2680
H123.70893.59854.36314.45053.70953.59554.37032.62392.22891.00711.72712.07583.15904.4465
C132.49142.76272.70663.45092.48412.74292.70641.52382.15421.36732.03622.07581.22023.4458
O142.94503.26012.67823.96212.99613.32822.74282.40273.27182.26552.48423.15901.22024.0035
H152.77143.77363.13502.55501.09461.77531.77962.17222.50504.34905.26804.44653.44584.0035

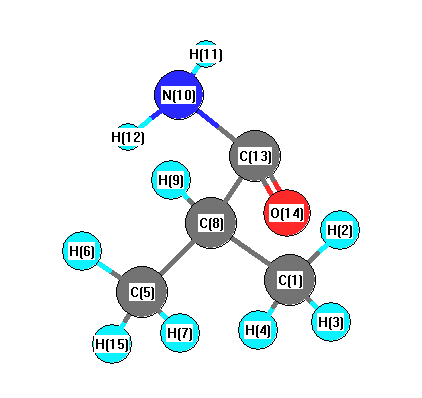
picture of Propanamide, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 C5 111.411 C1 C8 H9 109.166
C1 C8 C13 109.269 H2 C1 H3 108.196
H2 C1 H4 108.266 H2 C1 C8 111.370
H3 C1 H4 108.941 H3 C1 C8 109.576
H4 C1 C8 110.427 C5 C8 H9 108.841
C5 C8 C13 108.713 H6 C5 H7 108.327
H6 C5 C8 111.221 H6 C5 H15 108.377
H7 C5 C8 109.582 H7 C5 H15 108.857
C8 C5 H15 110.414 C8 C13 N10 115.996
C8 C13 O14 121.842 H9 C8 C13 109.416
N10 C13 O14 122.115 H11 N10 H12 117.808
H11 N10 C13 117.075 H12 N10 C13 121.169
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability