return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for H2NCH2COOH (Glycine)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-284.143516
Energy at 298.15K-284.150466
HF Energy-283.887432
Nuclear repulsion energy178.828357
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3699 3511 50.13      
2 A' 3506 3328 0.47      
3 A' 3098 2940 14.94      
4 A' 1843 1750 240.86      
5 A' 1732 1644 19.83      
6 A' 1508 1432 11.88      
7 A' 1439 1365 18.95      
8 A' 1340 1272 12.93      
9 A' 1194 1133 145.13      
10 A' 1156 1098 137.31      
11 A' 979 929 181.22      
12 A' 845 802 61.92      
13 A' 639 607 9.62      
14 A' 467 443 34.26      
15 A' 262 248 9.62      
16 A" 3588 3406 1.30      
17 A" 3140 2980 8.92      
18 A" 1418 1346 0.33      
19 A" 1207 1146 0.89      
20 A" 939 891 2.82      
21 A" 682 647 112.11      
22 A" 516 490 31.47      
23 A" 243 231 54.50      
24 A" 61 58 5.47      

Unscaled Zero Point Vibrational Energy (zpe) 17749.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 16848.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.34279 0.12928 0.09695

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.549 0.000
O2 1.184 0.819 0.000
O3 -0.973 1.493 0.000
C4 -0.582 -0.856 0.000
N5 0.398 -1.926 0.000
H6 -0.515 2.355 0.000
H7 -1.236 -0.950 0.873
H8 -1.236 -0.950 -0.873
H9 1.008 -1.815 0.807
H10 1.008 -1.815 -0.807

Atom - Atom Distances (Å)
  C1 O2 O3 C4 N5 H6 H7 H8 H9 H10
C11.21391.35601.52122.50681.87772.13062.13062.69362.6936
O21.21392.26012.43382.85472.29063.12193.12192.75972.7597
O31.35602.26012.38163.68380.97582.60842.60843.93953.9395
C41.52122.43382.38161.45093.21161.09471.09472.02462.0246
N52.50682.85473.68381.45094.37702.09342.09341.01751.0175
H61.87772.29060.97583.21164.37703.49393.49394.51194.5119
H72.13063.12192.60841.09472.09343.49391.74632.40542.9331
H82.13063.12192.60841.09472.09343.49391.74632.93312.4054
H92.69362.75973.93952.02461.01754.51192.40542.93311.6134
H102.69362.75973.93952.02461.01754.51192.93312.40541.6134

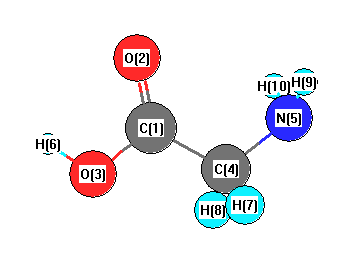
picture of Glycine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O3 H6 106.123 C1 C4 N5 114.993
C1 C4 H7 107.957 C1 C4 H8 107.957
O2 C1 O3 123.062 O2 C1 C4 125.326
O3 C1 C4 111.612 C4 N5 H9 108.944
C4 N5 H10 108.944 N5 C4 H7 109.853
N5 C4 H8 109.853 H7 C4 H8 105.807
H9 N5 H10 104.896
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.622      
2 O -0.509      
3 O -0.624      
4 C -0.232      
5 N -0.738      
6 H 0.437      
7 H 0.198      
8 H 0.198      
9 H 0.325      
10 H 0.325      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 5.391 -0.333 0.000
y -0.333 5.420 0.000
z 0.000 0.000 3.697


<r2> (average value of r2) Å2
<r2> 117.990
(<r2>)1/2 10.862