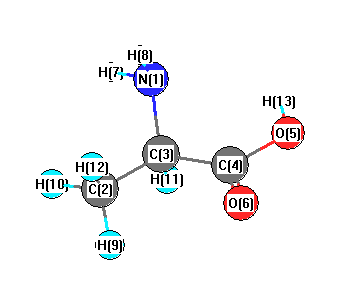
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3606 |
3449 |
6.64 |
|
|
|
| 2 |
A |
3510 |
3358 |
0.57 |
|
|
|
| 3 |
A |
3375 |
3229 |
282.52 |
|
|
|
| 4 |
A |
3172 |
3035 |
5.88 |
|
|
|
| 5 |
A |
3135 |
2999 |
26.68 |
|
|
|
| 6 |
A |
3066 |
2934 |
21.18 |
|
|
|
| 7 |
A |
3061 |
2929 |
7.61 |
|
|
|
| 8 |
A |
1891 |
1809 |
280.56 |
|
|
|
| 9 |
A |
1696 |
1623 |
38.36 |
|
|
|
| 10 |
A |
1523 |
1457 |
16.32 |
|
|
|
| 11 |
A |
1514 |
1449 |
3.92 |
|
|
|
| 12 |
A |
1474 |
1410 |
436.81 |
|
|
|
| 13 |
A |
1427 |
1365 |
11.09 |
|
|
|
| 14 |
A |
1406 |
1345 |
4.55 |
|
|
|
| 15 |
A |
1332 |
1274 |
9.66 |
|
|
|
| 16 |
A |
1259 |
1205 |
13.28 |
|
|
|
| 17 |
A |
1234 |
1181 |
8.62 |
|
|
|
| 18 |
A |
1166 |
1115 |
9.09 |
|
|
|
| 19 |
A |
1111 |
1063 |
47.17 |
|
|
|
| 20 |
A |
1028 |
984 |
1.42 |
|
|
|
| 21 |
A |
955 |
913 |
66.83 |
|
|
|
| 22 |
A |
920 |
880 |
62.28 |
|
|
|
| 23 |
A |
886 |
847 |
108.85 |
|
|
|
| 24 |
A |
812 |
777 |
7.88 |
|
|
|
| 25 |
A |
739 |
707 |
8.20 |
|
|
|
| 26 |
A |
568 |
544 |
4.74 |
|
|
|
| 27 |
A |
533 |
510 |
0.93 |
|
|
|
| 28 |
A |
410 |
393 |
5.57 |
|
|
|
| 29 |
A |
369 |
353 |
17.87 |
|
|
|
| 30 |
A |
312 |
298 |
17.17 |
|
|
|
| 31 |
A |
274 |
263 |
13.13 |
|
|
|
| 32 |
A |
225 |
216 |
0.29 |
|
|
|
| 33 |
A |
77 |
74 |
1.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24033.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22993.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.803 |
|
|
|
| 2 |
C |
-0.517 |
|
|
|
| 3 |
C |
-0.140 |
|
|
|
| 4 |
C |
0.628 |
|
|
|
| 5 |
O |
-0.593 |
|
|
|
| 6 |
O |
-0.471 |
|
|
|
| 7 |
H |
0.341 |
|
|
|
| 8 |
H |
0.344 |
|
|
|
| 9 |
H |
0.215 |
|
|
|
| 10 |
H |
0.174 |
|
|
|
| 11 |
H |
0.209 |
|
|
|
| 12 |
H |
0.167 |
|
|
|
| 13 |
H |
0.448 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.013 |
2.156 |
0.289 |
5.464 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.404 |
-0.004 |
-0.363 |
| y |
-0.004 |
-36.893 |
-0.100 |
| z |
-0.363 |
-0.100 |
-33.056 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.430 |
-0.004 |
-0.363 |
| y |
-0.004 |
-1.163 |
-0.100 |
| z |
-0.363 |
-0.100 |
4.593 |
|
| Polar |
|---|
| 3z2-r2 | 9.185 |
|---|
| x2-y2 | -1.511 |
|---|
| xy | -0.004 |
|---|
| xz | -0.363 |
|---|
| yz | -0.100 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.786 |
0.291 |
0.115 |
| y |
0.291 |
7.232 |
-0.091 |
| z |
0.115 |
-0.091 |
5.109 |
<r2> (average value of r
2) Å
2
| <r2> |
162.470 |
| (<r2>)1/2 |
12.746 |